

Vibrational Spectroscopy
Vibrational spectroscopy infers quantized nuclear motion from transitions between molecular vibrational states. A measured peak is not itself a bond vibration or a force constant. It is a feature produced by a transition operator, a pair of molecular states, rotational and environmental structure, populations, line broadening, and an instrument.
The harmonic approximation supplies initial state labels. Symmetry constrains the transition moments, while anharmonicity changes fundamental frequencies and produces overtones, combinations, and resonant mixing. Extracting a force field requires distinguishing those effects from rotation, temperature, and the sample environment.
Required background. The quantum harmonic oscillator provides energies and coordinate matrix elements; spectroscopic selection rules explains the operator and symmetry tests used for infrared activity.
What the Spectral Coordinate Means
Section titled “What the Spectral Coordinate Means”Energy, frequency, and vacuum wavenumber
Section titled “Energy, frequency, and vacuum wavenumber”For an isolated transition from lower state to upper state ,
Infrared spectra are commonly plotted against vacuum wavenumber in . Ordinary frequency and angular frequency obey
The same Greek letter is often used without a tilde for a wavenumber in spectroscopic tables. A trustworthy calculation follows the units, not the glyph. In this page:
- is an angular frequency in ;
- is an ordinary frequency in ;
- is an observed transition wavenumber; and
- is a conventional harmonic spectroscopic constant in .
Thus and are not interchangeable.
Line, band, and band origin
Section titled “Line, band, and band origin”A line corresponds to one resolved transition between specified quantum states. A vibrational band is a family of transitions sharing a vibrational-state change. In a gas, one vibrational band can contain many rotational lines. Its band origin is the transition between the two vibrational term values before rotational energy is added, within the stated effective Hamiltonian.
These distinctions matter because the largest peak in a rotationally structured band need not occur at its origin. In liquids, matrices, and solids, unresolved rotational structure and environmental interactions can instead produce one broad feature. Its maximum is then an operational peak position, not automatically an isolated-molecule band origin.
Absorbance is not a transition moment
Section titled “Absorbance is not a transition moment”For a homogeneous sample, absorbance is often reported as
where is transmittance after the relevant reference and baseline procedures. Peak height depends on concentration, path length, line shape, resolution, saturation, and overlap. Integrated intensity is usually closer to a state-to-state line strength, but it still requires declared units, temperature, abundance, and normalization conventions.
Harmonic Oscillator Approximation
Section titled “Harmonic Oscillator Approximation”One stable coordinate
Section titled “One stable coordinate”Let measure displacement from a stable equilibrium along one molecular coordinate. Expanding the potential gives
For positive curvature , keeping only the quadratic term produces
and energy levels
Adjacent levels are equally spaced. The transition wavenumber for any step is therefore
The approximation is local: it uses curvature at one minimum. A flat minimum with requires higher terms and does not define a positive-frequency harmonic oscillator. It does not encode dissociation, asymmetric level spacing, large-amplitude motion, or coupling to another electronic surface.
Many harmonic modes
Section titled “Many harmonic modes”Near a nonlinear equilibrium, subtracting three translation and three rotation displacement directions leaves harmonic vibration directions. At a linear equilibrium only two independent rotations displace the nuclei, leaving . This is the count of local harmonic coordinates, including both bending directions; it is not a global dimension formula for all molecular shapes. At quadratic order, a suitable mass-weighted transformation decouples them:
where
The harmonic term value is
with occupation vector . The zero-point term remains even when every .
What the harmonic model predicts
Section titled “What the harmonic model predicts”The model predicts:
- a reproducible number of vibrational coordinates;
- approximate frequency scales from a mass-weighted quadratic force field;
- collective displacement patterns called normal modes;
- isotope shifts through the masses; and
- simple zeroth-order state labels such as .
It does not by itself predict an observed infrared intensity. That also requires the dipole moment as a function of nuclear coordinates. Nor does a harmonic frequency equal the measured fundamental once anharmonicity, rovibrational structure, environment, and model error matter.
Normal Modes as Spectral Labels
Section titled “Normal Modes as Spectral Labels”Collective motion, not an isolated bond
Section titled “Collective motion, not an isolated bond”A normal coordinate is a collective mass-weighted displacement pattern. Even a mode described informally as a “C–O stretch” generally moves several atoms. The label identifies the dominant geometric character, not a literal one-bond coordinate.
At quadratic order, the Cartesian Hessian and mass matrix obey the generalized eigenproblem
Equivalently, the eigenvalues of
are . The full derivation, removal of translations and rotations, coordinate conventions, and Wilson formulation are developed in Normal Modes of Polyatomics.
Symmetry and degeneracy
Section titled “Symmetry and degeneracy”Normal modes can be labeled by irreducible representations of the equilibrium symmetry group that preserves the isotope masses as well as the geometry. Symmetry serves three separate purposes:
- It classifies the displacement subspace.
- It determines whether a transition moment must vanish in an idealized symmetry limit.
- It identifies which states can interact when anharmonic couplings are included.
A degenerate frequency corresponds to a multidimensional mode subspace. The individual vectors chosen inside that subspace are basis-dependent; the subspace and its symmetry are physical. Treating two arbitrary degenerate eigenvectors as uniquely oriented molecular motions overinterprets a numerical normal-mode calculation.
Two compact examples
Section titled “Two compact examples”Water is nonlinear and has
normal modes: a symmetric stretch, a bend, and an antisymmetric stretch. Each can be infrared active because its displacement symmetry is compatible with at least one Cartesian component of the molecular dipole.
Carbon dioxide is linear and has
vibrational coordinates but only three distinct harmonic frequencies: the bend is doubly degenerate. In the centrosymmetric equilibrium structure, the symmetric stretch is infrared inactive at first electric-dipole order, while the bend and antisymmetric stretch are active. The conclusion comes from the transition operator and symmetry, not from whether the atoms visibly move.
Infrared Activity
Section titled “Infrared Activity”Transition dipole criterion
Section titled “Transition dipole criterion”In the electric-dipole approximation, absorption of light polarized along is governed by
A transition is electric-dipole allowed only if this matrix element is nonzero. Near equilibrium, expand each space-fixed or molecule-fixed dipole component in normal coordinates:
For a harmonic fundamental , the leading criterion is
for at least one component . In point-group language, the normal mode must transform like , , or for the relevant component to be symmetry allowed.
Why a permanent dipole is neither necessary nor sufficient
Section titled “Why a permanent dipole is neither necessary nor sufficient”Pure rotational electric-dipole spectroscopy requires a permanent molecule-fixed dipole in the simplest treatment. Vibrational infrared activity instead requires a change of dipole along the normal coordinate.
- A molecule can have no equilibrium dipole and still possess infrared-active modes. The antisymmetric stretch of carbon dioxide is the standard example.
- A polar molecule can possess a mode whose dipole derivative vanishes by symmetry. The molecule is polar, but that particular fundamental is infrared inactive at leading order.
The equilibrium value and the derivative answer different questions.
Harmonic selection rule
Section titled “Harmonic selection rule”If the dipole expansion is truncated after its linear term and the vibrational states are harmonic, then
For absorption from a cold vibrational ground state, this gives the familiar fundamental transition
This is an approximation-dependent rule, not an exact prohibition against overtones. Higher dipole derivatives, anharmonic wavefunctions, state mixing, magnetic-dipole or electric-quadrupole operators, and symmetry breaking can all make nominally forbidden features observable.
Intensity and population
Section titled “Intensity and population”For an isolated transition between individual nondegenerate substates, the absorption coefficient integrated over frequency has the dependence
Here are substate number densities. For unresolved levels with degeneracies and statistically populated substates, the corresponding population difference is , using a lower-level-averaged line strength. In LTE this becomes . The measured integrated area can also contain rotational line-strength factors, nuclear-spin weights, isotopic abundance, a partition function, and the chosen line-intensity normalization. A weak band may reflect a small transition moment, a sparsely populated lower state, dilution by isotope abundance, or experimental limitations. “Weak” is not synonymous with “forbidden.”
Anharmonic Corrections
Section titled “Anharmonic Corrections”Nonquadratic potential and dipole surfaces
Section titled “Nonquadratic potential and dipole surfaces”Two independent approximations are often conflated:
- Mechanical harmonicity truncates the potential after quadratic terms.
- Electrical linearity truncates the dipole surface after terms linear in the normal coordinates.
Relaxing the first changes energies and wavefunctions. Relaxing the second changes the transition operator. Consequently, overtone intensity can arise from anharmonic state mixing, nonlinear dipole dependence, or both.
For several modes, a local potential expansion has the schematic form
The cubic and quartic constants shift levels and couple harmonic basis states. Their numerical values depend on the coordinate normalization and electronic structure model.
Diatomic term values
Section titled “Diatomic term values”For a semirigid diatomic, the leading spectroscopic expansion is
With , adjacent intervals contract:
The fundamental is therefore
not . Using a measured fundamental as the harmonic wavenumber biases a curvature-derived force constant. Vibrations of Diatomics develops the full convention and its relation to dissociation models.
Polyatomic term-value expansion
Section titled “Polyatomic term-value expansion”For nondegenerate modes of a semirigid polyatomic molecule away from strong resonances, a common second-order form is obtained by defining :
The sign of an is not fixed in general. With this convention, the fundamental of mode is approximately
Degenerate mode groups require a compatible occupation convention and vibrational angular-momentum terms; this simple nondegenerate formula should not be applied to them unchanged. The expression is useful only with its Hamiltonian convention and resonance treatment stated. Near-degenerate states can make ordinary nondegenerate perturbation theory unstable or misleading.
Resonance and intensity borrowing
Section titled “Resonance and intensity borrowing”Suppose two same-symmetry zeroth-order states and are close in energy and coupled by . In their subspace,
The mixed energies are
When is comparable to , neither observed level retains a pure zeroth-order identity. If is bright and is dark, mixing redistributes transition strength between the eigenstates. This intensity borrowing is a physical effect, not a violation of symmetry: the exact eigenstates, rather than the uncoupled labels, enter the transition matrix element.
Fermi resonance commonly describes coupling between a fundamental and a nearby overtone or combination state of the same symmetry. Coriolis and Darling–Dennison resonances represent different coupling structures. A fit must identify which interaction is included rather than absorbing every displacement into unrelated “observed frequencies.”
Where a local anharmonic expansion fails
Section titled “Where a local anharmonic expansion fails”Low-order expansions around one equilibrium are most reliable for semirigid molecules and low vibrational excitation. They can fail for:
- torsions that cross several equivalent minima;
- inversion or pseudorotation with a low barrier;
- strongly hydrogen-bonded or floppy complexes;
- levels near dissociation;
- conical intersections or other electronic-state interactions; and
- solids with strong temperature-dependent or collective anharmonicity.
Variational nuclear-motion calculations, hindered-rotor models, explicit multisurface dynamics, or periodic lattice methods may then be the appropriate description.
Overtones, Combinations, and Hot Bands
Section titled “Overtones, Combinations, and Hot Bands”Definitions
Section titled “Definitions”Starting from the vibrational ground state:
- a fundamental has one quantum in one mode, such as ;
- an overtone has two or more quanta in one mode, such as ;
- a combination band excites more than one mode, such as .
A difference band raises one mode while lowering an already occupied mode. It cannot start from the complete vibrational ground state; thermal excitation or other preparation must populate the mode that loses a quantum.
A hot band begins in a thermally populated excited vibrational state. For example, is a hot-band transition even though . “Overtone” describes the final excitation pattern relative to the ground state; “hot band” describes the lower-state population.
Harmonic locations and real band origins
Section titled “Harmonic locations and real band origins”The harmonic model places a first overtone near and a two-mode combination near . Real band origins are shifted by diagonal and cross-anharmonicities, and resonances can split or mix them.
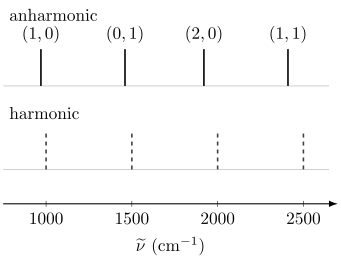
A hypothetical two-mode assignment from , with final-state labels . Solid sticks are anharmonic band origins; gray dashed sticks are the corresponding harmonic estimates. The displacement is not a universal red shift: its sign and size depend on the force field and state coupling. Stick heights are schematic and not intensities.
Worked two-mode assignment
Section titled “Worked two-mode assignment”Consider the term-value model
with all constants in :
Subtracting the ground-state term value gives
The overtone is not : here
whereas its calculated band origin is . In this model, : the cross-mode term cancels in this difference. By contrast, the combination origin differs from the sum of fundamentals by . These differences isolate distinct anharmonic contributions.
Why overtones can appear
Section titled “Why overtones can appear”In the doubly idealized harmonic-plus-linear-dipole model, vanishes because changes the oscillator quantum number by one. Two mechanisms relax that result:
- A quadratic dipole term contains , whose harmonic matrix element connects and .
- An anharmonic potential mixes harmonic states, so the exact vibrational eigenstates no longer carry pure labels.
Mechanical anharmonicity does not automatically open an overtone: an even potential with a strictly odd linear dipole still has a zero amplitude by parity. Only symmetry-compatible mixing or operator terms can supply the strength. An overtone can also borrow strength from a nearby bright fundamental of the same exact symmetry. Its observability therefore contains information about both the dipole surface and vibrational coupling.
Temperature and hot bands
Section titled “Temperature and hot bands”For nondegenerate harmonic levels in thermal equilibrium,
Writing gives
Low-frequency modes can therefore generate visible hot bands at ordinary temperatures, while high-frequency bond stretches may remain overwhelmingly in . Degeneracy and rotational populations must be included when a quantitative intensity ratio is required.
Connecting Spectra to Force Constants
Section titled “Connecting Spectra to Force Constants”Diatomic curvature from a harmonic wavenumber
Section titled “Diatomic curvature from a harmonic wavenumber”For a diatomic coordinate with reduced mass and harmonic wavenumber ,
so the quadratic force constant is
The units must be consistent. If is in and in , their product is in . For the hypothetical values
one obtains
This is the curvature at equilibrium in the specified coordinate. It is not a dissociation energy and need not predict highly excited levels.
Why a fundamental is not enough by itself
Section titled “Why a fundamental is not enough by itself”If only the fundamental is observed, then the diatomic relation
contains at least two unknown constants. A harmonic estimate may replace by the fundamental, but the resulting force constant is explicitly an effective approximation. Hot bands, overtones, isotope data, or a physically constrained potential can separate curvature from anharmonicity.
Polyatomic inverse problem
Section titled “Polyatomic inverse problem”For a polyatomic molecule, observed harmonic frequencies constrain the eigenvalues of the mass-weighted Hessian. Eigenvalues alone do not reconstruct the entire matrix. Different force fields can share the same spectrum, especially when only a subset of modes is observed.
A useful force-field determination may combine:
- infrared and Raman band positions and intensities;
- isotope substitutions, which change while approximately preserving the Born–Oppenheimer force field;
- rotational constants and symmetry assignments;
- mode displacement vectors from electronic-structure calculations;
- overtone, combination, and resonance data; and
- an internal-coordinate force-field model with stated constraints.
Even then, individual “bond force constants” depend on the chosen internal coordinates and coupling terms. Measured transition frequencies constrain term-value differences. Harmonic normal-mode frequencies are curvature-derived model quantities, and their decomposition into local stretching, bending, and interaction constants is also model-dependent.
Isotope substitution as a controlled perturbation
Section titled “Isotope substitution as a controlled perturbation”Within the Born–Oppenheimer approximation, isotopic substitution changes nuclear masses much more strongly than it changes the electronic potential. For an isolated diatomic harmonic mode,
For a polyatomic molecule, one must solve the full mass-weighted eigenproblem; there is generally no single reduced mass attached to a collective mode. Isotopic shifts are powerful because they test mode assignments and constrain eigenvectors as well as frequencies. Deviations from simple mass scaling can also expose vibrational mixing, nonadiabatic corrections, or inconsistent assignments.
Harmonic calculations and scale factors
Section titled “Harmonic calculations and scale factors”An electronic-structure frequency calculation usually diagonalizes a mass-weighted Hessian at an optimized geometry. Its output is a set of harmonic wavenumbers. Experiment commonly reports fundamentals. Their difference combines at least two effects:
- the physical anharmonic correction; and
- error in the electronic structure, basis set, geometry, and numerical Hessian.
Empirical scale factors can reduce average discrepancies for a declared method, basis, molecular class, and target property. They do not turn a harmonic calculation into an anharmonic Hamiltonian, resolve resonances, or provide a molecule-specific uncertainty automatically. A scale factor fitted to fundamentals should not be reused silently for zero-point energies or transition intensities.
Reading Experimental Spectra
Section titled “Reading Experimental Spectra”Gas, liquid, and solid samples answer different questions
Section titled “Gas, liquid, and solid samples answer different questions”| Environment | Typical spectral consequences | Interpretation caution |
|---|---|---|
| Dilute gas | Rotational fine structure may be resolved; isolated-molecule quantum numbers are useful | Doppler, pressure, hyperfine, and unresolved rovibrational structure still matter |
| Liquid or solution | Rotation is strongly interrupted; bands broaden and solvent shifts appear | Peak positions and widths depend on concentration, solvent, temperature, and hydrogen bonding |
| Matrix or molecular solid | Site splitting, crystal fields, phonon coupling, and collective modes may appear | Free-molecule point-group labels can be reduced by the environment |
| Periodic solid | Zone-center optical phonons and crystal momentum replace an isolated-molecule mode picture | A molecular normal-mode assignment may be only an approximate local description |
Comparing a gas-phase computed harmonic wavenumber directly with a broad solution-phase peak can be useful for orientation, but it is not a like-for-like test of an isolated molecular Hamiltonian.
High-resolution vibrational bands are rovibrational
Section titled “High-resolution vibrational bands are rovibrational”Molecules normally rotate while vibrating. A gas-phase infrared transition therefore connects rovibrational states, not pure vibrational states. Branch structure can reveal the band origin and vibration-dependent rotational constants, while unresolved rotation can shift the apparent envelope maximum. The P/Q/R hierarchy and Coriolis complications are developed in Rovibrational Coupling.
Peak position, area, and width carry different information
Section titled “Peak position, area, and width carry different information”- Position constrains an energy difference after shifts and unresolved structure are modeled.
- Integrated area constrains a population-weighted transition strength under a declared normalization.
- Width and shape constrain lifetimes, collisions, inhomogeneity, instrumental response, and unresolved components.
These observables should not be interchanged. A broader band can have a lower peak while retaining the same integrated area. Line Shapes and Broadening develops that distinction.
Databases and line lists
Section titled “Databases and line lists”A line list may tabulate wavenumbers, lower-state energies, Einstein coefficients, statistical weights, pressure-broadening parameters, and temperature-dependent intensities. Before using one, check:
- isotopologue and isotopic-abundance convention;
- vacuum versus medium wavenumber;
- reference temperature and partition-function convention;
- whether positions are measured, fitted, or predicted;
- uncertainty and quantum-number completeness; and
- whether the file contains discrete lines or a measured cross section.
For example, HITRAN line intensity incorporates the lower-state population, stimulated-emission factor, partition function, and a terrestrial isotopic abundance convention. It is not simply . The NIST Chemistry WebBook provides evaluated and compiled vibrational data for many species, but each species page must still be read for data provenance and phase.
Exercises
Section titled “Exercises”Exercise 1: Fundamental and hot-band spacing
Section titled “Exercise 1: Fundamental and hot-band spacing”A diatomic molecule has
Using the two-term anharmonic model, determine and . Predict the overtone position.
Solution
The adjacent intervals are
Their difference gives
so
The ground-state overtone is
It is below twice the fundamental, illustrating why an overtone is not located by simple multiplication.
Exercise 2: Bias from using a fundamental as a curvature
Section titled “Exercise 2: Bias from using a fundamental as a curvature”For the diatomic in Exercise 1, compare the force constant inferred by treating as the harmonic frequency with the one inferred from . The reduced mass is the same in both calculations.
Solution
Since ,
The fundamental-based estimate is about too small. Improving the peak-center precision does not remove this model bias; separating the curvature and anharmonic correction does.
Exercise 3: Isotope substitution and a degenerate bend
Section titled “Exercise 3: Isotope substitution and a degenerate bend”Replace one oxygen in linear by . Does the harmonic coordinate count change? Must the two bend frequencies split? What happens to the inversion-based infrared activity argument?
Solution
There are still four harmonic vibrational coordinates near the linear equilibrium. Axial symmetry still makes the two perpendicular bends degenerate, so the three generic distinct frequencies remain two stretches and one doubly degenerate bend. Inversion now exchanges unequal isotope masses and is not a symmetry of the full nuclear Hamiltonian. The stretches can mix their former symmetric/antisymmetric character, so both can carry infrared strength. Loss of inversion does not imply loss of axial symmetry.
Exercise 4: Infrared activity without a permanent dipole
Section titled “Exercise 4: Infrared activity without a permanent dipole”Carbon dioxide has no equilibrium dipole moment. Why can its antisymmetric stretch absorb infrared light while its symmetric stretch is inactive at first electric-dipole order?
Solution
Infrared activity depends on the transition dipole, whose leading vibrational term is controlled by
During the antisymmetric stretch, the two bond contributions cease to cancel instantaneously, so a dipole component changes along the mode and the derivative is nonzero. During the symmetric stretch, inversion symmetry is preserved and the electric-dipole derivative vanishes in the ideal molecule. The equilibrium value does not decide either result by itself.
Exercise 5: Mechanical and electrical anharmonicity
Section titled “Exercise 5: Mechanical and electrical anharmonicity”For a one-dimensional mode, suppose the potential is exactly harmonic but the dipole function contains a quadratic term. Can the transition appear? Conversely, can it appear with a linear dipole function and an anharmonic potential?
Solution
It can in both cases, provided the resulting matrix element is not forbidden by an exact symmetry.
With harmonic states and
the matrix element
is nonzero, so the quadratic dipole term gives electrical-anharmonic overtone intensity.
With a linear dipole but an anharmonic potential, the exact eigenstates are mixtures of harmonic number states. The matrix element of between the mixed ground and second excited states can then be nonzero. This is mechanical anharmonicity, but a symmetric quartic perturbation preserves parity and cannot open this transition with a strictly linear dipole. An asymmetric potential can do so. An observed overtone may contain both electrical and mechanical contributions.
Exercise 6: Isolate diagonal and cross-mode anharmonicity
Section titled “Exercise 6: Isolate diagonal and cross-mode anharmonicity”Suppose assigned band origins are , , , and for , , , and from . Use the stated quadratic term-value expansion to extract and without knowing the harmonic frequencies.
Solution
Subtracting differences of the same term values cancels the unknown harmonic frequencies and the spectator zero-point terms:
These are effective constants only within the stated expansion and assignments. Strong resonant mixing must be treated before interpreting the differences as uncoupled anharmonic coefficients.
Exercise 7: Thermal population of a low-frequency mode
Section titled “Exercise 7: Thermal population of a low-frequency mode”Estimate at for a nondegenerate mode with . Use . What does the result suggest about hot bands?
Solution
The Boltzmann ratio is
About nine percent as many molecules occupy as before rotational and degeneracy factors are included. A hot band may therefore be observable if its transition moment and spectral separation are favorable. The ratio would be much smaller for a high-frequency stretching mode at the same temperature.
Exercise 8: Force-field identifiability
Section titled “Exercise 8: Force-field identifiability”A nonlinear four-atom molecule has six accurately measured fundamental band origins. Do those six numbers uniquely determine its full Cartesian Hessian? Give three kinds of additional information that can make a force-field fit more informative.
Solution
No. The six fundamentals are anharmonic transition energies, not even the six harmonic Hessian eigenvalues without correction. Eigenvalues alone also do not determine all matrix elements or eigenvectors of the mass-weighted Hessian. The inverse problem remains model-dependent.
Useful additional constraints include any three of:
- isotope shifts from several substitutions;
- infrared and Raman symmetry assignments and polarization data;
- calculated normal-mode eigenvectors and a computed quadratic force field;
- overtone and combination-band term values that constrain anharmonic constants;
- rotational constants or structural constraints; and
- an internal-coordinate force-field model with physically justified sparsity or symmetry relations.
The fitted force constants must be reported with their coordinate convention and model uncertainty.
The vibrational-spectra lab implements CO Hessians and an HCl Morse-DVR calculation, with transition moments and convergence tests. Infrared spectroscopy and Raman spectroscopy develop the distinct measurement operators.
References
Section titled “References”- V. Barone, “Anharmonic Vibrational Properties by a Fully Automated Second-Order Perturbative Approach,” Journal of Chemical Physics 122, 014108 (2005), doi:10.1063/1.1824881.
- P. F. Bernath, Spectra of Atoms and Molecules, 5th ed., Oxford University Press, 2025, doi:10.1093/oso/9780197754498.001.0001.
- P. R. Bunker and P. Jensen, Molecular Symmetry and Spectroscopy, 2nd ed., NRC Research Press, 1998.
- J. L. Dunham, “The Energy Levels of a Rotating Vibrator,” Physical Review 41, 721–731 (1932), doi:10.1103/PhysRev.41.721.
- G. Herzberg, Molecular Spectra and Molecular Structure. I. Spectra of Diatomic Molecules, 2nd ed., Van Nostrand, 1950.
- G. Herzberg, Molecular Spectra and Molecular Structure. II. Infrared and Raman Spectra of Polyatomic Molecules, Van Nostrand, 1945.
- HITRANonline, Units and Definitions and Quantum Notation, accessed 2026-10-14.
- International Union of Pure and Applied Chemistry, “normal mode”, Compendium of Chemical Terminology, 5th ed., online version 5.0.0, 2025, doi:10.1351/goldbook.08648.
- K. K. Irikura, R. D. Johnson III, and R. N. Kacker, “Uncertainties in Scaling Factors for Ab Initio Vibrational Frequencies,” Journal of Physical Chemistry A 109, 8430–8437 (2005), doi:10.1021/jp052793n.
- P. M. Morse, “Diatomic Molecules According to the Wave Mechanics. II. Vibrational Levels,” Physical Review 34, 57–64 (1929), doi:10.1103/PhysRev.34.57.
- NIST Chemistry WebBook, SRD 69, Vibrational and electronic energy levels, accessed 2026-10-14.
- NIST Computational Chemistry Comparison and Benchmark Database, Vibrational Frequency Scaling Factors, accessed 2026-10-14.
- D. Papoušek and M. R. Aliev, Molecular Vibrational-Rotational Spectra, Elsevier, 1982.
- E. B. Wilson Jr., J. C. Decius, and P. C. Cross, Molecular Vibrations: The Theory of Infrared and Raman Vibrational Spectra, McGraw–Hill, 1955; Dover reprint, 1980.