

Oscillator Strengths
An oscillator strength is a dimensionless, energy-weighted measure of an electric-dipole transition. For absorption from a nondegenerate state to a higher state , the standard isotropic definition in SI units is
Here is the electric-dipole operator, is the electron mass, and is the elementary charge. The factor multiplying the matrix element normalizes the quantum transition to the response of a classical electron oscillator. It does not make a probability: an oscillator strength has no observation time, is not restricted to the interval , and by itself is neither a rate nor a peak cross section.
Oscillator strengths are valuable because they connect several descriptions of the same electric-dipole line:
- a transition dipole or reduced line strength from structure theory;
- an integrated absorption cross section;
- an Einstein coefficient after frequency and degeneracy factors are supplied;
- exact and approximate sum rules over complete sets of final states.
Those connections are convention-sensitive. A quoted value is incomplete unless the states, degeneracy average, polarization average, transition energy, and line grouping are known.
Required background. Transition rates explains preparation and channel sums; sum rules and completeness derives the commutator method. Here is the lower level, the upper level, and unqualified oscillator strengths are E1 absorption values in vacuum SI units. Signed reverse strengths are introduced separately below.
Dipole Matrix Elements
Section titled “Dipole Matrix Elements”Interaction with an electric field
Section titled “Interaction with an electric field”In the long-wavelength electric-dipole approximation, the interaction with a classical field is
For electrons with positions measured from a fixed origin,
For a molecule, the full dipole operator also contains the nuclear charges:
The transition dipole is
For two orthogonal exact states of a fixed-charge system, translating the coordinate origin adds a constant multiple of and therefore does not change the transition dipole. Permanent dipoles of charged systems require more care because the corresponding diagonal overlap is one.
The transition matrix element contains the structure physics. Symmetry can make it vanish, configuration mixing can redistribute it, and vibronic or spin–orbit coupling can lend intensity to a transition that is forbidden in a simpler model.
Polarization-resolved coupling
Section titled “Polarization-resolved coupling”For a unit polarization vector , the field probes
The corresponding directional oscillator strength is
For an isotropic ensemble or an average over three mutually orthogonal polarizations,
The average over is a mathematical three-direction average; a beam at one fixed propagation direction has only two transverse polarization modes. An isotropic sample supplies the required orientation average. The factor is not a dynamical suppression. It must not be inserted again when a tabulated quantity is already isotropically averaged.
For oriented molecules, aligned samples, Zeeman-resolved atoms, or polarization-selective detection, the component strengths are the physically relevant quantities. An isotropic value then discards information needed to predict the experiment.
Line strength and reduced matrix elements
Section titled “Line strength and reduced matrix elements”A line strength is the squared transition matrix element before the transition-energy factor is applied. For an atomic E1 transition between angular-momentum levels,
Here and collect all other labels. With the usual reduced-matrix-element convention and an unpolarized initial level,
The factor averages over initial magnetic substates. The reduced line strength already represents the appropriate sums over magnetic substates and spherical components. Adding an extra magnetic-substate sum to this formula double counts degeneracy.
For hyperfine-resolved data, if the stated line connects definite levels. For term-to-term or multiplet strengths, the adopted line grouping and statistical weights must be stated separately.
Length and velocity forms
Section titled “Length and velocity forms”For a nonrelativistic Hamiltonian with canonical kinetic energy and local coordinate-dependent potentials, define
The commutator relation
gives, between exact eigenstates,
Consequently the length and velocity forms agree:
These equations are written for nondegenerate states. Initial-state averages or reduced-matrix-element conventions add the corresponding degeneracy factors.
Approximate wavefunctions generally give different length- and velocity-form values. Their disagreement is a useful convergence diagnostic, but it is not automatically a statistical uncertainty. Nonlocal pseudopotentials, relativistic Hamiltonians, incomplete response terms, and inconsistent operators can modify the simple commutator relation, so apparent agreement is not by itself proof of accuracy.
Oscillator Strength Definition
Section titled “Oscillator Strength Definition”State-to-state absorption strength
Section titled “State-to-state absorption strength”Using
the isotropic state-to-state definition can be written in either dipole or coordinate form:
The sign of the electronic charge disappears after taking the modulus squared. The energy factor remains essential: two transitions with the same dipole matrix element but different frequencies do not have the same oscillator strength.
Dimensional analysis provides a quick check:
Atomic-unit form
Section titled “Atomic-unit form”In atomic units,
For a level-to-level E1 transition whose reduced line strength is in and whose energy separation is in hartrees,
For nondegenerate states, set and interpret as the squared vector transition dipole in atomic units. A common error is to insert an energy in electronvolts into this atomic-unit formula.
Reverse transitions and signed strengths
Section titled “Reverse transitions and signed strengths”The absorption oscillator strength is normally quoted as a positive number for . If the same algebraic definition is extended to the reverse transition without replacing the energy difference by its absolute value, then
For level-averaged strengths,
The negative sign is useful in sum rules because downward transitions from an excited initial state contribute negative spectral weight. It does not mean that an emission probability or rate is negative. Databases commonly list positive absorption values even when the corresponding coefficient describes downward emission.
Converting f, gf, log(gf), S, and A
Section titled “Converting f, gf, log(gf), S, and A”Several quantities that appear side by side in line lists are related but not interchangeable:
- is dimensionless and usually averages over the lower level’s substates.
- removes that lower-level average and is abbreviated .
- means in standard atomic line lists.
- is a squared matrix element and carries dipole-squared units unless atomic units are declared.
- is a spontaneous-emission rate in inverse seconds and averages over the upper level’s substates.
For an E1 line in vacuum,
Equivalently, with vacuum wavelength ,
These conversions assume the same E1 line, the same level grouping, and consistent degeneracy conventions. They do not apply unchanged in a dielectric medium. Magnetic-dipole, electric-quadrupole, and higher-multipole transitions have their own line strengths and rate formulas; NIST generally does not assign oscillator strengths to those forbidden-transition multipoles.
Einstein Coefficients gives the detailed-balance derivation of the and relations, their spectral-energy-density conventions, and their blackbody connection.
Why an oscillator strength can exceed one
Section titled “Why an oscillator strength can exceed one”An individual value is not a Bernoulli probability. In a many-electron system, collective or strongly allowed transitions can carry oscillator strength larger than one. What is constrained under the assumptions of the Thomas–Reiche–Kuhn rule is the complete signed sum, not each term separately.
Even for a one-electron system, excited-state upward strengths can exceed one because negative downward contributions must also be included in the signed sum.
Sum Rules
Section titled “Sum Rules”The Thomas–Reiche–Kuhn sum
Section titled “The Thomas–Reiche–Kuhn sum”For nonrelativistic electrons with fixed nuclei, canonical electron kinetic energy, and local coordinate-dependent interactions, the electronic-dipole oscillator strengths from an exact stationary state obey
The symbol is shorthand for a complete spectral resolution:
The continuum density and continuum-state normalization must be used consistently. A list of discrete bound lines generally does not exhaust the rule.
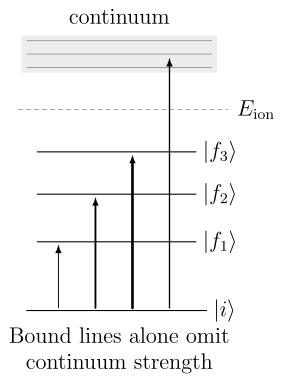
Schematic distribution of oscillator strength among bound lines and a continuum. Arrow widths represent relative strengths only. The exact Thomas–Reiche–Kuhn sum requires a complete set of final channels and a single polarization, degeneracy, and sign convention.
For one Cartesian component,
Averaging the three Cartesian component rules gives the isotropic form. For cross components, the underlying identity is a symmetrized sum:
The real part cannot generally be omitted for a complex stationary state, such as an oriented angular-momentum state. For , the product is already a real squared modulus.
For the stated Hamiltonian, the double commutator is
The full derivation and the general completeness method are given in Sum Rules and Completeness Tricks.
Conditions and qualifications
Section titled “Conditions and qualifications”The simple right-hand side assumes:
- nonrelativistic canonical electron kinetic energy;
- a consistent electric-dipole operator;
- local coordinate-dependent potentials;
- exact eigenstates of the same Hamiltonian;
- a complete discrete-plus-continuum final-state set;
- signed energy differences for an excited initial state;
- all required polarizations and unresolved degeneracies treated according to one convention.
If the nuclei move and the operator is the full molecular dipole, the right-hand side changes. For internal transitions of a translationally invariant nonrelativistic system of particles with masses and charges , the same electron-normalized definition gives
This follows by using the internal dipole in the double commutator. The final term removes center-of-mass acceleration for a charged system. It vanishes for a neutral molecule, but the nuclear terms remain. Zhou, Zhu, and Yan derive the generalized rule; the usual electronic value is the fixed-nucleus limit.
Relativistic corrections, nonlocal effective potentials, projected few-level models, and energy-dependent effective Hamiltonians can change the commutator or move strength outside the modeled space. A failed partial sum does not identify which assumption is responsible.
For an excited initial state, the signed rule is
Summing only positive upward strengths from an excited state need not yield and can exceed it.
Harmonic-oscillator saturation
Section titled “Harmonic-oscillator saturation”The one-dimensional harmonic oscillator gives a transparent example. For a single electron,
From the ground state, only has a nonzero dipole matrix element:
The directional oscillator strength is therefore
One line saturates the one-electron, one-component sum rule. Real atoms and molecules distribute dipole strength across many channels; the electronic and full molecular sums must use the respective assumptions just stated.
Partial sums as diagnostics
Section titled “Partial sums as diagnostics”Define a cumulative sum through an energy cutoff:
For a ground state, is nondecreasing when all included channels are ordinary upward E1 transitions. It approaches only after the continuum and sufficiently high-energy states are included. Its behavior can diagnose:
- missing diffuse or continuum basis functions;
- omitted symmetry or ionization channels;
- inconsistent degeneracy factors;
- accidental mixing of length- and velocity-form data;
- double-counted multiplet components;
- a truncated active-electron model whose effective sum is not the physical electron count.
Agreement with the final sum is necessary but not sufficient for accurate individual lines. Large positive and negative errors can cancel, and a basis can reproduce a global moment while misplacing strength spectrally.
Relation to many-body response
Section titled “Relation to many-body response”The oscillator-strength rule is an energy-weighted moment of a dipole response. In extended or interacting systems, analogous constraints appear as -sum rules for density and current response. Collective modes can carry substantial spectral weight while the exact total remains fixed by commutators and conservation laws. The response-function formulation is developed in Many-Body Sum Rules.
Relation to Absorption Strength
Section titled “Relation to Absorption Strength”Integrated cross section
Section titled “Integrated cross section”Consider an isolated narrow E1 line of targets in vacuum under weak illumination, with the initial-state and isotropic averages used above. Frequency-dependent prefactors are evaluated at the line center; extending the line profile to negative frequencies is the usual near-resonance approximation. Let be normalized in ordinary frequency:
The absorption cross section for an isolated line can be written
Its integrated area is
Thus measures integrated absorption strength. The peak cross section also depends on the line width and profile. Broadening can lower the peak while leaving the area unchanged when the line-strength sum is conserved.
The rate per target is related to photon flux by
This equation separates an intrinsic line quantity from illumination. A large oscillator strength does not imply a large observed rate if the source has little spectral photon flux at the transition frequency.
Ordinary frequency versus angular frequency
Section titled “Ordinary frequency versus angular frequency”If instead
then
and
The two areas differ by because . Using an angular-frequency line profile with an ordinary-frequency prefactor is one of the most common conversion errors.
Wavelength-integrated area
Section titled “Wavelength-integrated area”Cross-section area is not invariant under a change of horizontal coordinate. Since
a narrow line centered at satisfies
Therefore
The approximation replaces by across the line. For broad bands, apply the Jacobian point by point rather than using the narrow-line form.
Column density and optical depth
Section titled “Column density and optical depth”For a column density in the absorbing lower state,
In the optically thin limit,
Only in that limit is integrated fractional absorption directly proportional to . Saturation, unresolved blends, stimulated emission, state populations, radiative transfer, and instrumental response must be modeled before an observed line area is interpreted as an intrinsic oscillator strength.
Absorption strength is not emission intensity
Section titled “Absorption strength is not emission intensity”The same E1 matrix element enters absorption and spontaneous emission, but a measured emission intensity also depends on the upper-state population:
Population kinetics, branching fractions, collisional quenching, optical depth, and collection efficiency can all alter the observed emission. Consequently an intensity ratio is not generally an oscillator-strength ratio. The conversion through is intrinsic; the population and apparatus factors are not.
Atomic and Molecular Usage
Section titled “Atomic and Molecular Usage”Atomic level data
Section titled “Atomic level data”Atomic databases usually report absorption oscillator strengths between levels. For an initial level with total angular momentum ,
The quoted is normally averaged over the lower magnetic substates and summed over upper substates and the three dipole components, with the isotropic factor already included. The weighted quantity
is convenient because it removes the initial-level average. Atomic abundance work often tabulates
The NIST spectral-line documentation distinguishes , , , , and relative intensity. Match the level grouping and the energy used to convert before comparing values.
Configuration interaction can shift oscillator strength between nearby levels even when the summed strength of the group is comparatively stable. Near an avoided crossing, level labels based on a dominant configuration can swap while the physical transition amplitudes vary continuously.
Multiplets and unresolved structure
Section titled “Multiplets and unresolved structure”Suppose several fine-structure lines belong to one LS-coupled multiplet. A multiplet strength is obtained by summing consistently resolved line strengths:
Because oscillator strength includes a transition-energy factor and an initial degeneracy average, blindly summing unweighted line values need not reproduce the convention used for a term-to-term value. If the line frequencies are nearly equal, a weighted multiplet relation may be an excellent approximation, but the definition should still be stated.
The Wigner–Eckart Theorem and angular-momentum recoupling coefficients determine how a reduced strength is partitioned among Zeeman, hyperfine, or fine-structure components.
Molecular electronic transitions
Section titled “Molecular electronic transitions”For molecular states, the dipole operator depends on electronic and nuclear coordinates. Within a Born–Oppenheimer representation, a vibronic transition moment takes the form
where
An electronic oscillator strength at fixed geometry uses and the electronic energy gap at that geometry. A vibronic band oscillator strength uses the nuclear-motion integral and the vibronic transition energy. They are not the same number.
In the Condon approximation,
so
The squared vibrational overlap is the Franck–Condon factor. It distributes electronic intensity among vibronic bands, but the oscillator strengths also carry their individual transition-energy factors. The band strengths are therefore only proportional to Franck–Condon factors when those frequency differences and other couplings can be neglected.
If the electronic transition moment varies with normal coordinate,
Herzberg–Teller terms can create or redistribute vibronic intensity. A transition with a vanishing Condon moment can then acquire nonzero band strength through vibronic coupling.
Rotational and orientation factors
Section titled “Rotational and orientation factors”Within a vibronic band, rotational line strengths are partitioned by Hönl–London factors and by the symmetry of the transition dipole component. Recovering a band strength requires the appropriate sum over all allowed rotational lines and branches, with magnetic-substate normalization and lower-state populations treated consistently. One branch alone need not exhaust the strength.
For gas-phase isotropic samples, orientation averaging produces the familiar factor. For crystals, surfaces, aligned molecules, and polarization resolved spectroscopy, tensor components should be retained. Reporting only one scalar oscillator strength can hide dichroism and selection-rule information.
Continuum and photoabsorption
Section titled “Continuum and photoabsorption”Above an ionization or dissociation threshold, the final states are energy-normalized and oscillator strength becomes a density. One may write
provided the continuum states and channel measure are normalized so that
The continuum contribution to the sum rule is
Different continuum normalizations move density factors between the matrix element and the integration measure. The physical integrated strength is unchanged when the conversion is done consistently.
Because
replacing a calculated transition energy by an observed one changes and by different powers. The adjusted energy must be reported rather than silently folded into a quoted strength.
Exercises
Section titled “Exercises”Exercise 1: Convert a transition dipole to an oscillator strength
Section titled “Exercise 1: Convert a transition dipole to an oscillator strength”A nondegenerate transition has energy
and isotropic line strength
Find . Use .
Solution
Convert the energy to atomic units:
For ,
The result is dimensionless. Inserting directly into the atomic-unit formula would overestimate the value by the hartree-to-electronvolt conversion factor.
Exercise 2: Polarization and isotropic averaging
Section titled “Exercise 2: Polarization and isotropic averaging”A transition dipole is
Compare the oscillator strengths for light polarized along , light polarized along , and an isotropic polarization average.
Solution
The directional definitions give
and
Because the component also vanishes,
The isotropic value is smaller because it averages over orientations that do not all couple to the -directed transition dipole.
Exercise 3: Finite nuclear mass
Section titled “Exercise 3: Finite nuclear mass”For a neutral hydrogen atom with nuclear mass , show that the full internal dipole sum, normalized with , is . Recover the result directly from the relative-coordinate Hamiltonian.
Solution
The full-particle rule gives because . In relative coordinates, and , where . The double commutator therefore gives . Defining oscillator strengths with instead of would change their numerical normalization; it would not change the measured absorption.
Exercise 4: Reduced line strength, gf, and log(gf)
Section titled “Exercise 4: Reduced line strength, gf, and log(gf)”An atomic E1 line has
Calculate , , and .
Solution
The lower-level statistical weight is
Then
The weighted oscillator strength is
and
Exercise 5: An excited-state strength budget
Section titled “Exercise 5: An excited-state strength budget”For a one-electron excited state, the sum of all included downward oscillator strengths is , and the included upward bound-state sum is . Assuming the exact Thomas–Reiche–Kuhn rule, how much signed strength remains in omitted upward channels?
Solution
For one electron,
Let be the omitted contribution. Then
so
The upward positive strength totals , exceeding one because the downward contribution is negative. There is no conflict with the signed sum rule.
Exercise 6: Change the spectral coordinate
Section titled “Exercise 6: Change the spectral coordinate”Starting from
derive the narrow-line wavelength-integrated area.
Solution
Since ,
Across a narrow line, replace in the Jacobian by . Reversing the integration limits removes the minus sign:
Therefore
For a broad band, cannot be replaced by one central value.
Exercise 7: Interpret gauge disagreement
Section titled “Exercise 7: Interpret gauge disagreement”A calculation gives
What can and cannot be concluded from these numbers?
Solution
For exact eigenstates and consistent operators under the Hamiltonian assumed in the length–velocity derivation, the two values should agree. Their difference is therefore evidence that the approximate states, basis, response treatment, Hamiltonian, or transition operators are not mutually consistent.
The discrepancy does not show which value is closer to the exact result, and it is not a statistical error bar. Both forms can share a systematic error. One should study basis and correlation convergence, verify commutator compatibility for nonlocal or relativistic terms, compare transition energies, and seek independent benchmark data.
Exercise 8: Vibronic intensity partition
Section titled “Exercise 8: Vibronic intensity partition”In the Condon approximation, an electronic transition has
Three resolved vibronic bands have Franck–Condon factors , , and . Neglect variation in transition frequency. Estimate the three band oscillator strengths and the omitted strength.
Solution
Under the stated approximation,
Thus
The listed Franck–Condon factors sum to , so the omitted vibrational levels carry approximately
Exact band oscillator strengths also contain their individual transition energies and may include coordinate-dependent transition moments, rotation, nonadiabatic mixing, and other intensity-borrowing mechanisms.
The oscillator-strength reference collects conversions for quick use after the state and averaging conventions have been chosen.
References
Section titled “References”- P. F. Bernath, Spectra of Atoms and Molecules, 5th ed., Oxford University Press, 2025, doi:10.1093/oso/9780197754498.001.0001.
- H. A. Bethe and E. E. Salpeter, Quantum Mechanics of One- and Two-Electron Atoms, Springer, 1957.
- B. H. Bransden and C. J. Joachain, Physics of Atoms and Molecules, 2nd ed., Pearson, 2003.
- W. Demtröder, Laser Spectroscopy 1: Basic Principles, 5th ed., Springer, 2014, doi:10.1007/978-3-642-53859-9.
- R. C. Hilborn, “Einstein coefficients, cross sections, values, dipole moments, and all that,” American Journal of Physics 50, 982–986 (1982), doi:10.1119/1.12937; revised version, arXiv:physics/0202029; erratum, doi:10.1119/1.13515.
- International Union of Pure and Applied Chemistry, “Oscillator strength”, Compendium of Chemical Terminology, 5th ed., online version 5.0.0, 2025, doi:10.1351/goldbook.O04339.
- A. Kramida, “Spectral Lines: Selection Rules, Intensities, Transition Probabilities, Values, and Line Strengths”, in Atomic Spectroscopy: An Introduction, National Institute of Standards and Technology, accessed 2026-10-14.
- A. Kramida, Yu. Ralchenko, J. Reader, and NIST ASD Team, NIST Atomic Spectra Database, Standard Reference Database 78, doi:10.18434/T4W30F.
- W. Kuhn, “Über die Gesamtstärke der von einem Zustande ausgehenden Absorptionslinien,” Zeitschrift für Physik 33, 408–412 (1925), doi:10.1007/BF01328322.
- F. Reiche and W. Thomas, “Über die Zahl der Dispersionselektronen, die einem stationären Zustand zugeordnet sind,” Zeitschrift für Physik 34, 510–525 (1925), doi:10.1007/BF01328494.
- I. I. Sobelman, Atomic Spectra and Radiative Transitions, 2nd ed., Springer, 1992, doi:10.1007/978-3-642-76907-8.
- W. Thomas, “Über die Zahl der Dispersionselektronen, die einem stationären Zustande zugeordnet sind. (Vorläufige Mitteilung),” Naturwissenschaften 13, 627 (1925), doi:10.1007/BF01558908.
- B.-L. Zhou, J.-M. Zhu, and Z.-C. Yan, “Generalized Thomas–Reiche–Kuhn sum rule,” Physical Review A 73, 014501 (2006), doi:10.1103/PhysRevA.73.014501.