

Raman Spectroscopy
Raman spectroscopy measures inelastic scattering in which incident and scattered photons differ in energy because the material changes its internal state. In ordinary molecular Raman spectroscopy, the dominant off-resonant signal is controlled by how the electric polarizability changes during a rotation or vibration. This is a different coupling from direct infrared absorption, which is controlled by the electric dipole moment.
This page develops spontaneous molecular Raman scattering: its energy shifts, transition amplitudes, polarization, and population dependence. Surface-enhanced and coherent Raman methods require additional electromagnetic or nonlinear response models.
Required background. Vibrational spectroscopy supplies normal modes and anharmonic assignments; selection rules explains operator symmetry and angular-momentum constraints. Compare the dipole-derivative coupling in infrared spectroscopy.
Raman Shift and Energy Conservation
Section titled “Raman Shift and Energy Conservation”Incident, scattered, and material frequencies
Section titled “Incident, scattered, and material frequencies”Let the incident laser have angular frequency and the detected photon have angular frequency . Neglect center-of-mass recoil and external energy exchange. If the material begins in and ends in , energy conservation gives
Define the material transition frequency by
Then
Three cases follow:
| Process | Material change | Scattered photon |
|---|---|---|
| Rayleigh | ||
| Stokes Raman | ||
| Anti-Stokes Raman |
The labels describe the net initial and final states. They do not assert that the system occupies a stationary level at the laser photon energy during the scattering event.
Wavenumber-shift convention
Section titled “Wavenumber-shift convention”Spectra are commonly plotted against the Raman wavenumber shift
where the wavenumbers refer to vacuum. With this IUPAC convention,
Thus Stokes shifts are positive and anti-Stokes shifts are negative. Some instruments plot the anti-Stokes side against a positive shift magnitude, so the axis definition must accompany the data.
For a fixed material transition, the Raman shift is approximately independent of laser frequency even though the detected wavelength is not. Since
changing the laser changes the scattered wavelength and the optical response of every filter, grating, and detector in the experiment.
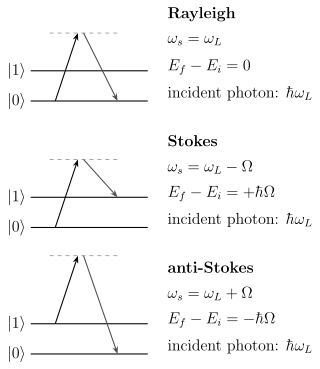
Energy bookkeeping for three scattering channels. The gray dashed intermediate guides represent off-resonant energy denominators, not stationary molecular eigenstates. In Stokes scattering the material gains ; in anti-Stokes scattering it loses .
Raman shift is not fluorescence wavelength
Section titled “Raman shift is not fluorescence wavelength”An elementary diagnostic is to repeat a measurement with another excitation wavelength:
- an ordinary Raman feature follows the laser while retaining approximately the same Raman shift;
- an ordinary fluorescence band is more naturally organized around an emission energy set by real electronic states; and
- resonance Raman, reabsorption, energy transfer, hot luminescence, and dispersive backgrounds can complicate this contrast.
Laser-shift testing is evidence, not a universal classifier. Time dependence, polarization, linewidth, power scaling, and sample chemistry provide independent checks.
Polarizability and Induced Dipoles
Section titled “Polarizability and Induced Dipoles”Linear optical response
Section titled “Linear optical response”For a nearly monochromatic field, the frequency-domain linear response is . In the off-resonant, slowly varying geometry approximation this is represented by
The dynamic polarizability tensor depends on the nuclear geometry and the optical frequency. Far from absorption resonances in a transparent, nonmagnetic system it can often be treated as approximately real and symmetric. Near resonance or in an absorbing sample it is generally complex and strongly frequency dependent; the response in time then requires a causal convolution rather than instantaneous multiplication.
For normal coordinates near equilibrium, fix , suppress it temporarily in the coefficient labels, and expand
where
is the first-order Raman tensor of mode in the off-resonant Placzek approximation.
Classical sideband derivation
Section titled “Classical sideband derivation”Take one linearly polarized field and one classical normal coordinate,
Keeping one scalar polarizability component,
the induced dipole becomes
An oscillating dipole radiates. The first term produces elastic Rayleigh scattering; the other two produce frequency-shifted sidebands. This argument gets the frequencies and the leading derivative rule right:
It does not by itself explain spontaneous scattering from a vibrational ground state, the Bose population factors, single-photon statistics, or the detailed interference among electronic intermediate states. Those require a quantum description.
A permanent dipole is not required
Section titled “A permanent dipole is not required”Raman activity asks whether the electron cloud’s deformability changes along the motion. A molecule can therefore have:
- no permanent dipole but a nonzero polarizability anisotropy;
- an infrared-inactive vibration with a nonzero polarizability derivative; or
- a permanent dipole while a particular polarizability derivative vanishes.
Homonuclear diatomics such as illustrate the first case. Their permanent-dipole pure rotational absorption is absent, but their anisotropic polarizability permits rotational Raman scattering.
Quantum Scattering Amplitude
Section titled “Quantum Scattering Amplitude”Transition polarizability
Section titled “Transition polarizability”In the electric-dipole approximation, the scattering amplitude for incident polarization and detected polarization has the form
Write electric-dipole matrix elements as
One common Kramers–Heisenberg–Dirac convention introduces
and writes
The two sums are the two time orderings of the dipole interactions. Here is an energy width, so a lifetime-limited level has . This is a phenomenological damping prescription; a consistent resonant open-system treatment also specifies initial/final widths and dephasing. Overall signs and damping conventions vary across sources, but the physical content does not: amplitudes from all intermediate states and both orderings add coherently before the modulus is squared.
The intermediate label runs over actual eigenstates used in the resolution of identity. A “virtual state” in a level diagram is shorthand for this off-shell sum and its energy denominators; it is not an additional stationary molecular level with a measurable lifetime.
Differential cross-section structure
Section titled “Differential cross-section structure”Define an energy cross section as scattered power per solid angle divided by incident irradiance. Suppressing unit-system constants, it has the structure
A photon-number cross section instead divides the scattered photon rate by incident photon flux. The two are related by
The displayed fourth-power formula uses . Photon counts and radiant power must not be compared without this conversion. The familiar fourth-power factor is only one part of the laser dependence. The transition polarizability itself is dynamic, and collection optics and detector responsivity change with . Raw intensities acquired with different lasers are therefore not made comparable by a wavelength-to-the- fourth correction alone.
Placzek limit
Section titled “Placzek limit”Far from electronic resonance, the electronic response can vary smoothly with the nuclear coordinates. Under Born–Oppenheimer and Placzek approximations, the vibronic transition polarizability reduces schematically to
Use dimensional mass-weighted coordinates, with mode Hamiltonian . For a harmonic fundamental,
The leading Raman-activity condition is therefore
for at least one tensor component admitted by the measurement geometry. Higher polarizability derivatives, anharmonic state mixing, vibronic coupling, and resonance effects produce overtone and combination intensity beyond this leading rule.
Stokes and Anti-Stokes Scattering
Section titled “Stokes and Anti-Stokes Scattering”Harmonic-oscillator matrix elements
Section titled “Harmonic-oscillator matrix elements”For one normal mode,
The adjacent-level matrix elements are
Stokes scattering creates one vibrational quantum and carries the factor . Anti-Stokes scattering removes one and requires an initial excitation, giving the factor .
Thermal occupation and intensity ratio
Section titled “Thermal occupation and intensity ratio”At thermal equilibrium the mean harmonic occupation is
For line-integrated radiant powers, with the same collection geometry and a slowly varying transition polarizability,
Hence
For response-corrected photon count rates, division by each photon energy replaces the fourth power by a third power. The excitation is the same for the two sides. This distinction persists even for a perfectly calibrated detector; calibration must identify whether it returns photons or energy.
In wavenumber units, use
For a mode, 532 nm excitation, and , the Boltzmann factor is about and the fourth-power factor is about , giving
This explains why room-temperature vibrational Raman spectra are commonly recorded on the Stokes side.
Raman thermometry requires calibration
Section titled “Raman thermometry requires calibration”Solving the ratio for temperature gives
That expression is valid only after accounting for:
- wavelength-dependent throughput and detector response;
- different Rayleigh-filter transmission on the two sides;
- polarization acceptance and collection geometry;
- background and baseline subtraction;
- reabsorption or self-absorption;
- resonance-dependent tensor changes;
- overlapping bands and line-shape integration; and
- nonequilibrium mode populations or laser heating.
A Stokes/anti-Stokes temperature is a mode-population temperature under a model. Agreement among several modes, laser powers, and independent thermometers is stronger evidence for thermal equilibrium.
Raman Selection Rules
Section titled “Raman Selection Rules”Tensor representation test
Section titled “Tensor representation test”In the ordinary nonresonant symmetric-tensor limit, the first-order Raman tensor transforms as the symmetric second-rank Cartesian products. A general resonant transition tensor need not obey this restriction. The matrix-element test can be written as
From a nondegenerate totally symmetric vibrational ground state, a fundamental is Raman active when
where the right side is spanned by
This is the character-table version of . It determines which tensor elements may survive, not how large they are.
Raman tensors and laboratory polarization
Section titled “Raman tensors and laboratory polarization”For a fixed molecular or crystal orientation, the idealized mode intensity is
As an example, in the standard convention with as the twofold axis, representative symmetric Raman tensors are
The tensor has symmetric and entries. Rotating the sample rotates these tensors into the laboratory frame. A missing line in one polarization geometry can therefore be a geometric zero rather than a mode that is inactive in every geometry.
Isotropic samples and depolarization
Section titled “Isotropic samples and depolarization”For a real symmetric mode tensor, define its isotropic invariant
and anisotropy invariant
In the standard ideal 90-degree experiment with linearly polarized excitation polarized perpendicular to the scattering plane, and analyzed scattered polarization parallel or perpendicular to that incident electric vector,
The depolarization ratio is
For ordinary nonresonant Raman scattering in an isotropic sample, for a totally symmetric vibration and for a nontotally symmetric vibration under the ideal assumptions. A totally symmetric band need not have ; anisotropy can make it partially depolarized.
Computational chemistry programs often report the orientationally averaged Raman activity
is not a raw experimental peak height. Converting it to a simulated spectrum requires excitation frequency, scattered-frequency factors, population, the oscillator matrix-element factor, line shape, and a stated photon-versus-energy and differential-versus-integrated convention.
Infrared–Raman complementarity
Section titled “Infrared–Raman complementarity”The leading activity tests are
| Feature | Infrared absorption | Ordinary Raman scattering |
|---|---|---|
| Leading material operator | Dipole derivative | Polarizability derivative |
| Optical process | One incident photon absorbed | Incident photon scattered into another mode |
| Tensor character | Vector | Symmetric rank and parts |
| Homonuclear diatomic rotation | Permanent-dipole E1 inactive | Can be active through anisotropy |
| Common experimental challenge | Optical propagation and saturation | Weak signal, Rayleigh rejection, fluorescence |
In a centrosymmetric molecule, the dipole operator is ungerade while the ordinary polarizability is gerade. Therefore a first-order fundamental cannot be both electric-dipole infrared active and ordinary Raman active. This mutual-exclusion rule requires exact inversion symmetry and the stated leading operators. It does not say every mode must appear in one spectrum, and it can change when an isotope pattern, surface, or field actually breaks inversion, or when another multipole participates. Resonance alone cannot defeat this parity rule: two electric-dipole interactions have even parity even when their intermediate-state denominator is small.
For linear , the gerade symmetric stretch is Raman active and infrared inactive at leading order. The ungerade bend and antisymmetric stretch are infrared active and ordinary Raman inactive. Water lacks inversion symmetry, so its fundamentals can be active in both methods.
Harmonic and anharmonic rules
Section titled “Harmonic and anharmonic rules”With a linear polarizability expansion and harmonic states,
is the leading one-mode rule. A quadratic derivative can produce overtones and combinations through matrix elements such as
and
Anharmonic eigenstate mixing can also make a linear polarizability term contribute to nominal overtones. Calling every feature “forbidden” hides the actual approximation that suppresses it.
Rotational Raman Spectroscopy
Section titled “Rotational Raman Spectroscopy”Polarizability anisotropy
Section titled “Polarizability anisotropy”For a linear molecule in a nondegenerate electronic state, neglect electronic spin structure and define body-fixed components parallel and perpendicular to the molecular axis. Its polarizability anisotropy is
If , reorientation does not modulate the induced dipole and shifted pure rotational Raman scattering vanishes in the leading model. If , the rank- polarizability channel permits
For pure rotational Raman scattering, belongs to the elastic channel; shifted lines use .
Rigid-rotor line positions
Section titled “Rigid-rotor line positions”For term values
the Stokes transition has shift
The first line, from , occurs at , and adjacent allowed initial values give a spacing of
The anti-Stokes transition has shift magnitude
Centrifugal distortion makes the spacings nonuniform. Nuclear-spin statistics can suppress alternating initial levels or change their statistical weights. Populations, degeneracies, Placzek–Teller factors, and the factor determine line strengths.
Nitrogen example
Section titled “Nitrogen example”Taking for a rigid-rotor estimate of , the first two Stokes shifts are
Their separation is . A real spectrum must also include centrifugal distortion and the nuclear-spin statistical pattern of the isotopologue.
Vibration–rotation Raman branches
Section titled “Vibration–rotation Raman branches”For a -to- vibration–rotation band of the same simple linear-rotor model, the scalar and rank-2 channels give
giving O, Q, and S branches in a common convention:
| Branch | Rotational change |
|---|---|
| O | |
| Q | |
| S |
If the lower and upper vibrational states have constants and , the simple branch shifts are
Electronic symmetry, vibrational angular momentum, polarization, and nuclear statistics can refine this simple branch picture. The detailed molecular Hamiltonian and Coriolis structure remain in Rovibrational Coupling.
Resonance Raman Spectroscopy
Section titled “Resonance Raman Spectroscopy”Near an electronic transition
Section titled “Near an electronic transition”When approaches a real electronic or vibronic transition, one or more denominators
become small. The scattering can be enhanced by many orders of magnitude, but the off-resonant approximation also becomes less reliable. The measured tensor can become complex, excitation-frequency dependent, and sensitive to individual intermediate states.
Resonance Raman spectroscopy is therefore selective rather than merely stronger. Modes coupled to the resonant chromophore or electronic excitation can be enhanced far more than spectator modes.
Franck–Condon and vibronic mechanisms
Section titled “Franck–Condon and vibronic mechanisms”In Albrecht’s organization of molecular resonance Raman amplitudes:
- the A term is associated with Franck–Condon structure and displacement of an allowed excited-state potential surface;
- the B and C terms involve vibronic, or Herzberg–Teller, coupling and can lend intensity through electronically excited states; and
- the terms interfere at the amplitude level.
This classification is a controlled way to discuss mode-specific enhancement. It is not equivalent to saying that every bond in the absorbing chromophore becomes equally intense.
Experimental tradeoffs
Section titled “Experimental tradeoffs”Moving toward resonance can improve sensitivity while introducing:
- fluorescence or phosphorescence backgrounds;
- absorption, self-absorption, and limited penetration depth;
- photochemistry, bleaching, or local heating;
- saturation and nonlinear power dependence;
- excitation-profile changes across a tunable laser scan; and
- altered polarization and selection-rule behavior.
A resonance Raman claim should report the excitation wavelength and bandwidth, sample absorption spectrum, power density, exposure history, and evidence that the sample remained chemically and thermally stable.
Boundaries of the ordinary theory
Section titled “Boundaries of the ordinary theory”Several named methods are not just high-signal versions of spontaneous Raman:
- surface-enhanced Raman scattering combines strongly nonuniform local electromagnetic fields with possible molecule–surface electronic effects;
- coherent anti-Stokes Raman scattering is a nonlinear four-wave-mixing process with phase matching and a nonresonant background;
- stimulated Raman scattering measures pump loss or Stokes gain through a nonlinear susceptibility; and
- Raman optical activity includes chiral electric-dipole–magnetic-dipole and electric-dipole–electric-quadrupole interference.
Their observables, power laws, tensors, and calibration requirements need process-specific treatment. The ordinary polarizability derivative is the baseline, not a universal replacement for those amplitudes.
What a Raman Instrument Measures
Section titled “What a Raman Instrument Measures”Minimal instrument chain
Section titled “Minimal instrument chain”A dispersive Raman measurement commonly contains:
- a narrowband laser and cleanup filter;
- illumination and collection optics;
- a sample geometry, often backscattering in microscopy;
- a notch, edge, or multi-stage filter that rejects the Rayleigh line;
- a spectrograph or interferometer; and
- a wavelength-resolved detector.
The useful Raman signal can be many orders of magnitude weaker than elastic scattering. Rayleigh rejection, stray-light control, detector dynamic range, and optical cleanliness are therefore part of the physics measurement, not mere presentation details.
A measurement model
Section titled “A measurement model”A schematic count model is
followed by convolution with the instrument line shape. Here is integration time, is incident power in the linear regime, is an effective number of scatterers, collects geometry and wavelength- dependent response, is a molecular line-area factor, and is a normalized line profile. When uses the energy cross section above, must include conversion from radiant power to counts, including the photon-energy factor.
This model is deliberately schematic. Confocal sampling, absorption, refraction, orientation, roughness, multiple scattering, and finite depth can make and spatially and spectrally dependent.
Calibration and response
Section titled “Calibration and response”A reproducible Raman spectrum records at least:
- excitation vacuum wavelength or frequency and its uncertainty;
- laser power at the sample, spot size, polarization, and exposure time;
- sampling geometry, objective numerical aperture, and confocal settings;
- sample phase, substrate, orientation, temperature, and atmosphere;
- Raman-shift calibration and reference material;
- spectral resolution and instrument line shape;
- cosmic-ray removal, baseline method, and smoothing or denoising;
- detector and optical relative-intensity correction;
- polarization response and analyzer convention; and
- whether plotted values are counts, count rate, corrected intensity, differential cross section, or normalized units.
Wavenumber calibration and relative-intensity calibration are distinct. Correct peak positions do not prove that relative band areas are unbiased. NIST Raman intensity standards, for example, characterize a wavelength- dependent response for specified excitation conditions rather than supplying a universal correction for every laser.
Choosing the laser
Section titled “Choosing the laser”Shorter wavelengths often increase the nonresonant scattering signal through the scattered-frequency factor and improve diffraction-limited spatial resolution. Longer wavelengths can reduce fluorescence and alter penetration, but may face weaker scattering and different detector noise. Resonance, absorption, sample heating, photochemistry, and the instrument’s response can dominate this simple tradeoff.
The scientifically useful laser is the one that preserves the sample and supports the intended inference, not automatically the wavelength giving the largest raw peak.
Exercises
Section titled “Exercises”Exercise 1: Quadratic sidebands
Section titled “Exercise 1: Quadratic sidebands”Take , , and . Find the induced-dipole frequencies and their amplitudes. Does a missing first derivative exclude all Raman scattering?
Solution
Since , the carrier amplitude is , and each sideband at has amplitude . The missing first derivative suppresses the leading fundamental, but not the quadratic overtone channel. Classical amplitudes alone do not give the quantum ground-state Stokes/anti-Stokes populations.
Exercise 2: Fundamental and overtone mechanisms
Section titled “Exercise 2: Fundamental and overtone mechanisms”For one harmonic coordinate, take
Which term first drives , and which first drives ? State one mechanism that can produce overtone intensity even if .
Solution
Because ,
The linear polarizability term therefore drives the harmonic fundamental. Since contains ,
so drives the overtone through electrical anharmonicity of the polarizability surface.
If the vibrational potential is anharmonic, exact eigenstates mix harmonic number states and can be nonzero. Near electronic resonance, explicit vibronic structure can also enhance overtone pathways. Exact symmetry still applies: an even potential and a strictly odd linear term leave the matrix element zero.
Exercise 3: Convert Raman shift to wavelength
Section titled “Exercise 3: Convert Raman shift to wavelength”A 532.0 nm laser excites a band at . Find the Stokes scattered wavelength. Also find the anti-Stokes wavelength for the corresponding feature.
Solution
The laser wavenumber is
For Stokes scattering,
so
For anti-Stokes scattering,
giving
The shift magnitude is the same, but wavelength is nonlinear in wavenumber.
Exercise 4: Photon counts and thermometry
Section titled “Exercise 4: Photon counts and thermometry”For the mode and 532 nm laser in the worked example, the true temperature is . A corrected detector reports photon counts with equal collection efficiency. Find the anti-Stokes/Stokes count ratio. What temperature results if it is mistaken for a power ratio?
Solution
Write and . The count ratio is , whereas the power ratio is about . Using the fourth-power thermometry formula on the count ratio gives
The error survives perfect relative quantum-efficiency correction. One must also convert photon number into radiant energy, or use the third-power formula.
Exercise 5: Depolarization ratio
Section titled “Exercise 5: Depolarization ratio”For an isotropic sample in the ideal 90-degree geometry, a mode has in arbitrary units and in the corresponding squared units. Calculate and the orientationally averaged Raman activity .
Solution
The parallel and perpendicular factors are
Therefore
The Raman activity is
Since , the band is polarized in this convention. That observation is compatible with a totally symmetric vibration, but a complete assignment still requires the point group, tensor geometry, and other bands.
Exercise 6: Carbon dioxide and mutual exclusion
Section titled “Exercise 6: Carbon dioxide and mutual exclusion”In centrosymmetric linear , classify the symmetric stretch and antisymmetric stretch as leading-order infrared or Raman active. Explain the result with inversion parity.
Solution
The vibrational ground state is gerade. The electric dipole operator is ungerade, so an electric-dipole infrared fundamental from that ground state must be ungerade. The polarizability tensor is gerade, so an ordinary Raman fundamental must be gerade.
The symmetric stretch is therefore Raman active and electric-dipole infrared inactive at leading order. The antisymmetric stretch is infrared active and ordinary Raman inactive.
This is mutual exclusion under exact inversion symmetry and the leading operators. It does not rule out weak intensity from isotopic symmetry breaking, inversion-breaking environments or fields, or higher multipoles. Electronic resonance by itself preserves the two-dipole even-parity rule.
Exercise 7: Which rotational lines survive?
Section titled “Exercise 7: Which rotational lines survive?”A linear rigid rotor has , but nuclear statistics allow only even in the electronic and vibrational state under study. Find the first three Stokes rotational Raman shifts and their spacing.
Solution
The initial states are , giving , , and : , , and . Their spacing is , not , because consecutive allowed initial states differ by two. Line positions alone must be interpreted together with isotope and spin statistics.
Exercise 8: Enhancement and interference
Section titled “Exercise 8: Enhancement and interference”Two intermediate states give a scalar Raman amplitude , where , , and is real. Compare at and . Why does adding the two separate intensities fail near the midpoint?
Solution
At the first resonance, . At the midpoint,
The separate intensities would instead sum to . They miss destructive interference between indistinguishable intermediate paths. The relative signs of dipole products and energy denominators matter.
For solid-state Raman tensors and collective excitations, see Raman and optical spectroscopy. For coherent population transfer through a Raman coupling, see STIRAP.
References
Section titled “References”- A. C. Albrecht, “On the Theory of Raman Intensities,” Journal of Chemical Physics 34, 1476–1484 (1961), doi:10.1063/1.1701032.
- P. F. Bernath, Spectra of Atoms and Molecules, 5th ed., Oxford University Press, 2025, doi:10.1093/oso/9780197754498.001.0001.
- P. A. M. Dirac, “The Quantum Theory of Dispersion,” Proceedings of the Royal Society A 114, 710–728 (1927), doi:10.1098/rspa.1927.0071.
- H. Goenaga Infante, J. Warren, J. Chalmers, G. Dent, J. L. Todoli, J. Collingwood, N. Telling, M. Resano, A. Limbeck, T. Schoenberger, D. B. Hibbert, A. Le Gresley, K. Adams, and D. Craston, “Glossary of Methods and Terms Used in Analytical Spectroscopy,” Pure and Applied Chemistry 93, 647–776 (2021), doi:10.1515/pac-2019-0203.
- International Union of Pure and Applied Chemistry, “depolarization ratio”, “Stokes–Raman scattering”, and “anti-Stokes Raman scattering”, Compendium of Chemical Terminology, 5th ed., online version 5.0.0, 2025.
- International Union of Pure and Applied Chemistry, “Raman scattering”, “Raman spectroscopy”, and “Raman wavenumber shift”, Compendium of Chemical Terminology, 5th ed., online version 5.0.0, 2025.
- D. A. Long, The Raman Effect: A Unified Treatment of the Theory of Raman Scattering by Molecules, Wiley, 2002, doi:10.1002/0470845767.
- R. L. McCreery, Raman Spectroscopy for Chemical Analysis, Wiley, 2000, doi:10.1002/0471721646.
- K. McNamara, D. V. Fursa, and I. Bray, “Efficient Calculation of Rayleigh and Raman Scattering,” Physical Review A 98, 043435 (2018), doi:10.1103/PhysRevA.98.043435.
- G. Placzek, Rayleigh-Streuung und Raman-Effekt, in E. Marx, ed., Handbuch der Radiologie, Vol. 6, Part 2, Akademische Verlagsgesellschaft, 1934.
- C. V. Raman and K. S. Krishnan, “A New Type of Secondary Radiation,” Nature 121, 501–502 (1928), doi:10.1038/121501c0.
- A. Urbas, K. Gierz, and D. D. Leber, Certification of Standard Reference Material 2246a: Relative Intensity Correction Standard for Raman Spectroscopy, 830 nm Excitation, NIST Special Publication 260-244, 2024, doi:10.6028/NIST.SP.260-244.
- E. B. Wilson Jr., J. C. Decius, and P. C. Cross, Molecular Vibrations: The Theory of Infrared and Raman Vibrational Spectra, McGraw–Hill, 1955; Dover reprint, 1980.