

Molecular Symmetry
Molecular symmetry turns a geometric observation into exact statements about a specified molecular model. Once the equilibrium geometry, isotopic composition, Hamiltonian, and coordinate convention are fixed, symmetry can:
- divide electronic, vibrational, and rotational problems into independent blocks;
- label states and orbitals without relying on a particular visualization;
- identify degeneracies protected by the model;
- prove that selected matrix elements vanish;
- organize normal modes and spectroscopic activity;
- expose when a numerical calculation has broken an intended symmetry.
The useful question is therefore not merely “what shape is the molecule?” It is:
Which transformations leave the relevant Hamiltonian unchanged, how do the states and operators transform, and what follows from those transformation laws?
A molecular point group is the practical fixed-geometry answer. It consists of rotations, reflections, inversion, and improper rotations that map the nuclear framework onto itself while leaving at least one point fixed. Point groups are powerful, but they are not the whole symmetry of a freely rotating, vibrating, permuting molecule.
Helpful background. Groups and representations explains invariant subspaces; selection rules proves the state–operator symmetry test. The character-table reference supplies the tables used below.
Molecular Point Groups
Section titled “Molecular Point Groups”A group attached to a geometry
Section titled “A group attached to a geometry”After removing overall translation, describe the nuclear framework by body-fixed positions
An orthogonal transformation is a point-group operation when it maps this framework onto itself, allowing exchange only among identical nuclear species. Schematically,
where is a permutation that preserves nuclear species. For an isotopically specified rovibrational problem, operations that exchange different isotopes need not remain symmetries of the nuclear kinetic energy.
The point-group label therefore belongs to a particular geometry and model. It is commonly assigned at an equilibrium structure, a transition structure, or another stationary geometry. A generic displaced geometry usually has less symmetry than the stationary point even when the displacement is a normal coordinate classified by the higher-symmetry reference group.
Common families
Section titled “Common families”Schoenflies notation is standard in molecular spectroscopy and quantum chemistry:
- contains only the identity.
- has one mirror plane, and has inversion.
- , , and have a principal -fold axis, with the indicated vertical or horizontal mirror planes.
- , , and also have twofold axes perpendicular to the principal axis.
- groups are generated by an improper rotation.
- , , and describe ideal tetrahedral, octahedral, and icosahedral frameworks.
- and are the linear-molecule groups without and with inversion, respectively.
Ideal examples include:
- bent water, ;
- pyramidal ammonia, ;
- trigonal-planar boron trifluoride, ;
- methane, ;
- benzene, ;
- hydrogen chloride, ;
- carbon dioxide, .
These labels assume idealized isolated structures. Substitution, isotopic labeling, an external field, a crystal environment, or a distorted geometry can reduce the symmetry.
A reliable identification sequence
Section titled “A reliable identification sequence”For a finite equilibrium geometry:
- Check whether the molecule is linear.
- Check for one of the high-symmetry polyhedral groups.
- Find the highest-order proper rotation axis and call it the principal axis.
- Look for twofold axes perpendicular to that principal axis.
- Test for a horizontal mirror plane, vertical or dihedral mirror planes, an inversion center, and improper axes.
- Verify closure by listing every operation, not only every visible symmetry element.
The last step matters. A symmetry element is a geometric object such as an axis or plane. A symmetry operation is a transformation such as or . One threefold axis supplies two nonidentity rotations.
Point groups are stabilizers, not molecular trajectories
Section titled “Point groups are stabilizers, not molecular trajectories”The point group of contains operations that leave the reference framework invariant as a set. It does not include arbitrary internal motions that carry the molecule through other geometries. Internal rotation, inversion tunneling, pseudorotation, and exchange of identical nuclei through feasible paths may require a molecular symmetry or permutation–inversion group.
That distinction is essential in high-resolution spectroscopy. A rigid equilibrium point group is often the correct first tool, but it is not always the exact symmetry group of the full rovibronic Hamiltonian.
Symmetry Operations
Section titled “Symmetry Operations”The basic operations
Section titled “The basic operations”The standard spatial operations are:
For a chosen principal axis:
- is perpendicular to the axis;
- contains the axis;
- contains the axis and bisects two perpendicular axes.
With the principal axis chosen as , one may write
Powers contain the reflection only for odd , since . The order of factors is harmless here because the rotation about commutes with reflection in the plane perpendicular to . For more general products, operator order must be kept explicit.
How an operation acts on a wavefunction
Section titled “How an operation acts on a wavefunction”At fixed nuclear geometry, a spatial operation acts on every electronic coordinate. For a spinless coordinate-space wavefunction,
with the nuclei simultaneously mapped onto symmetry-equivalent positions. If the clamped-nuclei electronic Hamiltonian respects the operation,
The inverse in the wavefunction argument makes the operators compose in the same order as the geometric transformations.
Water and its axis convention
Section titled “Water and its axis convention”Take water in the plane, with the axis along . The two vertical planes are then:
- , which exchanges the two hydrogens;
- , the molecular plane, which fixes all three nuclei.
The rotation also exchanges the hydrogens. All four operations
map the nuclear framework onto itself and form .
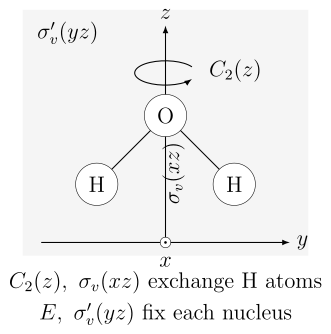
Axis convention for the worked example. The molecular plane is ; and exchange the hydrogens, while fixes each nucleus.
Axis conventions are not cosmetic. If the molecule is instead placed in the plane, labels called and in one convention can be exchanged in another. The displacement pattern and measurable polarization rule remain the same.
Spin and external fields
Section titled “Spin and external fields”Ordinary point groups act on spatial coordinates. If spin–orbit coupling is important, spinors transform under a double group, and a rotation cannot simply be represented as the identity on the spin state. Magnetic fields also require care because axial vectors and antiunitary time reversal enter the symmetry analysis.
An external electric field restricts the group to operations that preserve the field direction and polarity. A calculation performed in a field must therefore use the symmetry of “molecule plus field,” not the field-free molecular point group.
Irreducible Representations
Section titled “Irreducible Representations”States transform in invariant subspaces
Section titled “States transform in invariant subspaces”Let label an irreducible representation of . A basis of one copy of that irrep transforms as
Here:
- is the irrep dimension;
- labels components inside one irrep;
- distinguishes repeated copies of the same irrep.
If the Hamiltonian commutes with the group, its matrix is block diagonal in irreducible sectors:
States belonging to inequivalent irreps cannot mix under a symmetry-preserving Hamiltonian. Repeated copies of the same irrep can mix, so a symmetry label alone need not identify a unique state.
Symmetry-protected degeneracy
Section titled “Symmetry-protected degeneracy”On each isotypic sector, Schur’s lemma gives : the Hamiltonian can mix repeated copies but acts identically on their irrep components. Diagonalizing produces energy multiplets with degeneracy at least . Thus a multidimensional irrep enforces degeneracy:
The converse is false. Distinct one-dimensional irreps can cross, and unrelated levels can be accidentally degenerate. Conversely, numerical use of an Abelian subgroup can print the components of a parent multidimensional irrep under different labels even though the exact parent symmetry keeps them degenerate.
Mulliken labels
Section titled “Mulliken labels”For common molecular point groups:
- and usually denote one-dimensional irreps;
- denotes a two-dimensional irrep;
- denotes a three-dimensional irrep;
- subscripts distinguish irreps with different behavior under selected operations;
- and denote even and odd parity under inversion;
- primes and double primes denote even and odd behavior under a designated horizontal reflection.
The precise subscript convention depends on the group and chosen axes. Read the table header before interpreting a label. The symbol can denote either the identity operation or a two-dimensional irrep; context distinguishes them.
Reducible representations
Section titled “Reducible representations”A chosen set of coordinates, atomic orbitals, or displacement vectors usually transforms reducibly. If
then is the number of copies of irrep . This decomposition is the bridge from geometry to useful basis functions.
Character Tables
Section titled “Character Tables”Characters compress the representation
Section titled “Characters compress the representation”The character of a representation matrix is its trace:
Characters are invariant under a basis change and are constant on conjugacy classes. A character table therefore lists:
- conjugacy classes in columns;
- irreducible representations in rows;
- irreducible characters in the central block;
- common coordinate, rotation, and quadratic functions as interpretive aids.
The functions printed beside an irrep are examples of objects with that transformation law. They are not an exhaustive basis and do not mean that the irrep “is” a coordinate.
Orthogonality and reduction
Section titled “Orthogonality and reduction”For finite groups, irreducible characters obey
Taking the inner product of a reducible character with an irreducible character gives the multiplicity:
The sum is over conjugacy classes , and is the number of operations in the class. A valid reduction must give nonnegative integers and satisfy the dimension check
Failure of either check usually means that an atom-fixed count, vector trace, class size, or axis convention is wrong.
Projection operators
Section titled “Projection operators”The projector onto the full isotypic sector of irrep is
Applying it to trial orbitals or internal coordinates produces symmetry-adapted combinations. This character projector selects all copies of . Resolving individual rows and repeated copies requires the fuller matrix-element projectors or an additional orthogonalization and diagonalization within the projected space.
Direct products
Section titled “Direct products”Products of states or operators transform in tensor-product representations. Their characters multiply:
The resulting character can be reduced with the same formula. For one-dimensional real irreps, direct products are especially simple: multiply the signs operation by operation.
The compact and tables used below are collected at Character Tables.
Worked Example: Water Vibrations
Section titled “Worked Example: Water Vibrations”The Cartesian displacement representation
Section titled “The Cartesian displacement representation”Each of the three nuclei contributes three Cartesian displacement coordinates, so water begins with a nine-dimensional representation . Its character under operation is
Only nuclei left at their own positions contribute to the trace. A nucleus moved to another site contributes zero to the diagonal trace, even when the destination contains an identical nucleus.
For the axis convention in the figure:
The factors , , and are traces of the corresponding three-dimensional vector transformations. Reducing the character
against the table gives
The dimensions add to , as required.
Remove translations and rotations
Section titled “Remove translations and rotations”With along the axis and the molecule in the plane,
Therefore
The two modes are conventionally the symmetric stretch and bend; the mode is the asymmetric stretch in this axis convention. All irreps are one dimensional, so does not enforce a vibrational degeneracy.
The subtraction is subtraction of irrep multiplicities after decomposition. It is not subtraction of sets or individual displacement arrows.
Selection Rules
Section titled “Selection Rules”The totally symmetric test
Section titled “The totally symmetric test”For a transition operator , consider
The matrix element can be nonzero only if
where is the totally symmetric irrep. Equivalently, the multiplicity of the totally symmetric irrep is
If , symmetry forces the matrix element to vanish within the stated model. If , the transition is symmetry allowed, but its intensity can still be extremely small or zero for dynamical reasons.
Electric-dipole transitions
Section titled “Electric-dipole transitions”The electric-dipole components transform like , , and . In the water convention:
From an initial state, an -, -, or -polarized transition can reach , , or , respectively, provided the other quantum numbers and approximations also permit the transition.
For a fundamental vibration from a totally symmetric vibrational ground state, the normal coordinate must transform like at least one dipole component. Water’s fundamentals are therefore all infrared active in the electric-dipole and harmonic approximations.
Raman transitions
Section titled “Raman transitions”The leading nonresonant Raman operator is the symmetric polarizability tensor. Its components transform like
A normal mode is first-order Raman active when its irrep occurs among those quadratic functions. This is a different condition from infrared activity.
For a centrosymmetric molecule, dipole components are ungerade and polarizability components are gerade. A harmonic fundamental from a gerade ground state therefore cannot be both electric-dipole infrared active and first-order Raman active. This mutual-exclusion rule depends on inversion symmetry and on the stated leading operators.
Which forbidden lines can gain intensity?
Section titled “Which forbidden lines can gain intensity?”Mixing can relax an approximate label such as pure spin, electronic symmetry, or a harmonic mode assignment. A vibronic state, for example, transforms as the combined electronic and nuclear state rather than either factor separately. External fields or isotope substitution can also reduce the exact symmetry, and a magnetic-dipole or electric-quadrupole operator has a different transformation law from the electric dipole.
However, symmetry-preserving mixing cannot defeat an exact selection rule for the full states and the specified operator. If inversion remains exact, electric-dipole transitions still require opposite total parities, while electric-dipole–electric-dipole Raman transitions require equal parities. Anharmonicity or resonant intensity borrowing alone does not remove that distinction.
Normal Modes and Spectroscopy
Section titled “Normal Modes and Spectroscopy”Why the Hessian respects symmetry
Section titled “Why the Hessian respects symmetry”At a symmetry-preserving stationary geometry, the mass-weighted Hessian commutes with the displacement representation:
It can therefore be block diagonalized by irreducible representation. This gives three immediate consequences:
- modes from inequivalent irreps do not mix while the symmetry is preserved;
- multidimensional irreps enforce degenerate harmonic frequencies;
- transition-property derivatives can be screened by symmetry before an intensity calculation.
Modes of the same irrep can mix freely. If several modes are labeled , the label does not uniquely mean “the symmetric stretch.” Frequencies, displacement vectors, and internal-coordinate projections are needed for a physical assignment.
Degenerate modes
Section titled “Degenerate modes”For ammonia in , an vibrational mode has two components. Any orthonormal basis spanning that two-dimensional eigenspace is valid. Numerical eigenvectors can rotate within the subspace as the method, basis, or geometry changes while the observable subspace and common harmonic frequency remain unchanged.
A small computed splitting of an expected pair can signal:
- a geometry that is not exactly symmetry adapted;
- loose optimization or integration thresholds;
- unequal numerical treatment of the components in a lower-symmetry computational subgroup;
- a genuinely symmetry-breaking perturbation;
- an electronically unstable high-symmetry reference.
Using a subgroup alone does not physically split an exact parent-symmetry multiplet. An unstable high-symmetry geometry can have a degenerate pair of negative Hessian eigenvalues; instability does not by itself split that pair.
Correlation with lower symmetry
Section titled “Correlation with lower symmetry”When a molecule distorts from group to subgroup , an irrep of may remain irreducible or split when restricted to :
Correlation tables record this restriction. They are indispensable for tracking orbital and vibrational labels along reaction paths, symmetry-breaking coordinates, and Jahn–Teller distortions. Equal subgroup labels permit mixing; they do not guarantee that mixing is large.
The detailed normal-coordinate construction and intensity derivatives are developed in Normal Modes of Polyatomics. Raman Spectroscopy specializes the quadratic-function test to measured Raman tensors, depolarization ratios, and oriented samples.
Orbital Classification
Section titled “Orbital Classification”Symmetry-adapted linear combinations
Section titled “Symmetry-adapted linear combinations”Suppose the two hydrogen functions in water are and . Their two-dimensional permutation representation has character
because and exchange the functions, whereas and fix them. Character reduction gives
For normalized real functions with overlap , normalized symmetry-adapted combinations are
The coefficients apply only to an orthonormal site basis, or after neglecting overlap. Symmetry determines the relative signs, while the inner product determines normalization.
On oxygen, the and orbitals transform as , as , and as in this convention. A totally symmetric one-electron Hamiltonian couples only matching symmetry sectors:
Thus the hydrogen combination can mix with oxygen functions, and the combination can mix with oxygen . The oxygen function has no partner in this minimal hydrogen basis.
Symmetry proves the zeros. It does not determine the nonzero coupling strengths, orbital ordering, occupation, or degree of bonding.
Orbitals, configurations, and states
Section titled “Orbitals, configurations, and states”A molecular orbital has an irrep label, but a many-electron state is classified by the direct product of all occupied spin-orbital transformation laws, reduced with the required fermionic antisymmetry and spin coupling. The symmetry of a single frontier orbital is not automatically the symmetry of an excited state.
For open-shell systems or strong spin–orbit coupling, one may need:
- spin-adapted configuration state functions;
- double-group irreps;
- total angular-momentum projections for linear molecules;
- explicit treatment of near-degenerate electronic configurations.
Computational block structure
Section titled “Computational block structure”Quantum-chemistry programs often exploit only an Abelian subgroup even when the geometry has a non-Abelian point group. This keeps all working irreps one-dimensional and simplifies integral storage. Components of a parent or irrep then receive separate subgroup labels.
Compare the full point group and axis convention when matching labels from different programs: two different subgroup labels may be components of the same parent multiplet.
Beyond the Rigid Point Group
Section titled “Beyond the Rigid Point Group”Isotopic substitution
Section titled “Isotopic substitution”Electronic Coulomb interactions depend on nuclear charges, not nuclear masses. Replacing one isotope can therefore leave the clamped-nuclei electronic potential symmetric at the same geometry while reducing the symmetry of the nuclear kinetic energy and the full rovibrational problem.
This is why an isotopic substitution can activate weak vibrational lines or split rovibrational structure without substantially changing the electronic potential-energy surface.
Large-amplitude and nonrigid motion
Section titled “Large-amplitude and nonrigid motion”For tunneling between equivalent minima, internal rotation, umbrella inversion, or pseudorotation, the experimentally relevant states can extend over several geometries. A single equilibrium point group cannot represent every feasible permutation and inversion operation. Molecular symmetry groups and permutation–inversion groups provide the appropriate global classification.
Electronic degeneracy and distortion
Section titled “Electronic degeneracy and distortion”A nonlinear molecule in a degenerate electronic state can be unstable to a symmetry-lowering distortion through the Jahn–Teller effect. The high-symmetry point group still organizes the parent electronic and vibrational spaces, but the distorted minima belong to subgroups.
At intersections of electronic potential-energy surfaces, point-group symmetry can permit or forbid couplings on special subspaces, while generic conical intersection topology depends on the local branching plane. See Conical Intersections for the branching-plane construction.
Approximate symmetry
Section titled “Approximate symmetry”Near symmetry is often chemically informative, but “almost an irrep” is not an exact group-theory concept. If a perturbation is small, write
where has the higher symmetry. The labels of then remain useful zeroth-order labels, and controls mixing and splitting. Quantifying is more informative than assigning a higher point group by visual tolerance alone.
Exercises
Section titled “Exercises”Exercise 1: Identify idealized point groups
Section titled “Exercise 1: Identify idealized point groups”Assign the equilibrium point group of idealized , , , , and . State the geometric feature that distinguishes each answer.
Solution
- is linear and heteronuclear, so it is .
- is linear and centrosymmetric, so it is .
- Bent has one axis and two vertical mirror planes, so it is .
- Pyramidal has one axis and three vertical mirror planes, so it is .
- Ideal tetrahedral has tetrahedral symmetry, .
These are equilibrium-geometry labels. Isotopic substitution, distortion, or an external environment can reduce them.
Exercise 2: Replace one hydrogen by deuterium
Section titled “Exercise 2: Replace one hydrogen by deuterium”At the same fixed geometry, which point-group operations of water remain symmetries of the HDO nuclear kinetic energy? Does this change the clamped-nuclei Coulomb electronic potential?
Solution
Only and the molecular-plane reflection preserve the masses at every site, giving . The operations that exchange H and D no longer preserve the kinetic energy. The clamped-nuclei Coulomb electronic potential still has symmetry because H and D have the same nuclear charge. Thus the electronic potential and isotope-specific rovibrational Hamiltonian need not have the same point group.
Exercise 3: Construct hydrogen SALCs
Section titled “Exercise 3: Construct hydrogen SALCs”Apply symmetry to the two water hydrogen functions. Show that their representation is and construct normalized symmetry-adapted linear combinations for normalized functions with real overlap .
Solution
The identity and molecular-plane reflection fix both functions, while and exchange them. Thus
which reduces to . Their squared unnormalized norms are and , respectively, giving
Their relative signs describe transformation behavior. An overall sign change of either SALC has no physical effect.
Exercise 4: Polarization selection in water
Section titled “Exercise 4: Polarization selection in water”Assume an initial state in . Which final-state irreps can be reached by -, -, and -polarized electric-dipole transitions in the axis convention used on this page?
Solution
The coordinate components transform as
For one-dimensional real irreps, the product
contains precisely when . Therefore
Other quantum numbers and the actual transition moment can impose additional restrictions or make an allowed line weak.
Exercise 5: Prove vibrational mutual exclusion
Section titled “Exercise 5: Prove vibrational mutual exclusion”For a centrosymmetric molecule with a gerade vibrational ground state, show that a one-quantum harmonic fundamental cannot be both electric-dipole infrared active and first-order Raman active.
Solution
The electric dipole is ungerade. A nonzero infrared matrix element from a gerade ground state therefore requires an ungerade final vibrational state:
The polarizability tensor is gerade. A nonzero first-order Raman matrix element from the same ground state requires a gerade final state:
A normal mode of a centrosymmetric molecule is either gerade or ungerade, not both. Hence the same fundamental cannot satisfy both conditions within these approximations. With exact inversion, the same pair of parity eigenstates cannot acquire both E1 and E1–E1 intensity merely through anharmonic or vibronic mixing. Breaking inversion or using a different multipole process changes the symmetry test.
Exercise 6: Prove the symmetry block zero
Section titled “Exercise 6: Prove the symmetry block zero”Let and belong to inequivalent irreps of a finite point group, and let be totally symmetric. Why must
Solution
The matrix element transforms in
By irreducible-character orthogonality, the totally symmetric irrep occurs in only when the two irreps are equivalent. For inequivalent irreps, group averaging projects the matrix element to zero. This is the finite-group form of the block-diagonal statement behind Schur’s lemma.
Exercise 7: Change the water axes
Section titled “Exercise 7: Change the water axes”Move the water molecule from the plane to the plane while keeping as the axis. What happens to the and labels? Does the infrared polarization physics change?
Solution
Interchanging the roles of and interchanges the two vertical reflection planes. Under the standard convention, labels attached to and therefore exchange:
The asymmetric stretch receives the other printed label. Its displacement pattern and the statement that it couples to dipole polarization in the molecular plane perpendicular to the axis do not change. The relabeling is conventional; the observable relation is geometric.
Exercise 8: Diagnose a split degenerate mode
Section titled “Exercise 8: Diagnose a split degenerate mode”A calculation on idealized ammonia is run without imposed symmetry. Two modes expected to form an pair differ by a small frequency. Give a diagnostic sequence that distinguishes numerical symmetry breaking from a physical instability.
Solution
First inspect the optimized Cartesian geometry and quantify deviations from under a stated atom-matching tolerance. Then:
- tighten geometry, integration-grid, self-consistent-field, and Hessian thresholds;
- symmetrize the geometry and recompute the analytic or finite-difference Hessian;
- project the two eigenvectors onto the expected displacement subspace;
- test whether the splitting shrinks with numerical convergence;
- inspect the electronic and vibrational stability of the high-symmetry stationary point;
- repeat with symmetry explicitly imposed and with it disabled.
At an exactly geometry with an invariant Hamiltonian, the pair must remain degenerate even if its Hessian eigenvalue is negative. Persistent splitting there means that the Hamiltonian or numerical calculation breaks the assumed symmetry. A lower-energy distorted geometry can have physically split modes because its group is smaller; compare those modes with the unstable but degenerate parent pair.
Term symbols connects these point-group labels to linear-molecule angular projections and total-parity conventions.
References
Section titled “References”- P. W. Atkins and R. S. Friedman, Molecular Quantum Mechanics, 5th ed., Oxford University Press, 2011.
- D. M. Bishop, Group Theory and Chemistry, Dover, 1993.
- P. R. Bunker and P. Jensen, Molecular Symmetry and Spectroscopy, 2nd ed., NRC Research Press, 1998, ISBN 978-0-660-17519-5.
- P. R. Bunker and P. Jensen, Fundamentals of Molecular Symmetry, IOP Publishing, 2005, doi:10.1201/9781315273334.
- F. A. Cotton, Chemical Applications of Group Theory, 3rd ed., Wiley, 1990.
- G. Herzberg, Molecular Spectra and Molecular Structure II: Infrared and Raman Spectra of Polyatomic Molecules, Van Nostrand, 1945.
- International Union of Pure and Applied Chemistry, “Point Group,” Compendium of Chemical Terminology, 5th ed. (2025), doi:10.1351/goldbook.P04703.
- NIST Computational Chemistry Comparison and Benchmark Database, species organized by molecular point group, Standard Reference Database 101, Release 22 (May 2022).
- C. J. H. Schutte, J. E. Bertie, P. R. Bunker, J. T. Hougen, I. M. Mills, J. K. G. Watson, and B. P. Winnewisser, “Notations and Conventions in Molecular Spectroscopy: Part 2. Symmetry Notation,” Pure and Applied Chemistry 69, 1641–1650 (1997), doi:10.1351/pac199769081641.
- M. Tinkham, Group Theory and Quantum Mechanics, Dover, 2003.
- E. B. Wilson Jr., J. C. Decius, and P. C. Cross, Molecular Vibrations: The Theory of Infrared and Raman Vibrational Spectra, McGraw–Hill, 1955.