

Potential Energy Surfaces
A molecular potential energy surface assigns an energy to every admissible nuclear geometry for a specified electronic state and electronic Hamiltonian. Its minima organize stable structures, its curvature controls small-amplitude vibration, its saddle regions organize candidate reaction pathways, and its asymptotes identify dissociation channels.
The word surface is historical. A nonlinear molecule with nuclei has internal coordinates, so the object is usually a scalar field in a high-dimensional shape space rather than a literal two-dimensional surface. A contour plot is a projection or slice of that field.
Required background. The molecular Hamiltonian defines the fixed-geometry electronic operator; Born–Oppenheimer in molecules and scale separation explain when its eigenvalues can guide nuclear motion.
H₂⁺ gives an explicit one-coordinate example. Nonadiabatic coupling treats transfer between surfaces, and conical intersections develops their degeneracy geometry.
The Defining Eigenvalue Problem
Section titled “The Defining Eigenvalue Problem”Let denote a complete nuclear geometry after overall translation has been removed. For each fixed , solve
Throughout this page, includes the internuclear repulsion. The Born–Oppenheimer surface for electronic state is therefore
Some authors reserve for the electronic energy without internuclear repulsion and write
These conventions describe the same physics when translated consistently. A quoted minimum or barrier is ambiguous unless the electronic state, Hamiltonian, energy zero, and correction level are stated.
One geometry, several electronic states
Section titled “One geometry, several electronic states”The electronic Hamiltonian has a family of eigenvalues,
so a molecule generally has many potential energy surfaces. A label such as “the surface” usually means one specified electronic state over one specified region. Energy ordering is not always a reliable identity label: near avoided crossings, states can exchange character while their ordered eigenvalues remain smooth.
Spin, spatial symmetry, charge, and asymptotic channel labels help identify states. In computations, overlaps, transition properties, and reduced density matrices may be needed to track electronic character from one geometry to the next.
Born–Oppenheimer and adiabatic surfaces
Section titled “Born–Oppenheimer and adiabatic surfaces”Terminology is not completely uniform. A useful distinction is:
where is the diagonal Born–Oppenheimer correction, or DBOC. In a simple Cartesian representation,
The projected form is gauge invariant. It also shows that the corrected surface depends on nuclear masses, whereas the clamped-nuclei Born–Oppenheimer surface does not. Relativistic, radiative, finite-nuclear-size, external-field, and environmental corrections define still other effective surfaces.
Geometry Space, Not a Coordinate List
Section titled “Geometry Space, Not a Coordinate List”For distinct nuclei away from collisions, labeled Cartesian configurations lie in
Overall translations and rotations do not change an isolated molecule’s internal shape. Schematically, its shape space is
with further identifications under permutations of identical nuclei. Away from singular geometries, a nonlinear molecule has
local internal degrees of freedom. At a linear equilibrium the infinitesimal vibrational count is , because rotation about the molecular axis produces no displacement. The rotational quotient is singular there: this count is not the dimension of a regular global shape-space chart. Collinear geometries and identical-particle identifications require local care.
Coordinate representations
Section titled “Coordinate representations”Common coordinates include:
- Cartesian positions with translation and rotation projected out;
- bond lengths, bond angles, and dihedral angles;
- Jacobi coordinates for fragmentation and scattering;
- symmetry-adapted coordinates;
- mass-weighted normal coordinates near a stationary point;
- invariant distances or descriptors for fitted potentials.
The numerical function written as changes its appearance under a coordinate transformation, but the energy assigned to a physical geometry does not.
Gradients need a metric
Section titled “Gradients need a metric”The differential
is coordinate covariant. To convert it into a steepest-descent vector, one needs a metric . If the classical nuclear kinetic energy is
then
Thus “steepest descent” is incomplete without a coordinate metric. Mass-weighted Cartesian coordinates supply the metric used by the conventional intrinsic reaction coordinate.
Forces and electronic response
Section titled “Forces and electronic response”In Cartesian coordinates, the force on nucleus is
For an exact, normalized, nondegenerate electronic eigenstate in a fixed representation, the Hellmann–Feynman theorem gives
Approximate wavefunctions, incomplete optimization, and atom-centered bases can require response and Pulay terms. A smooth-looking energy fit can also have poor derivatives; forces must be validated as first-class observables of the representation.
What a Surface Is Not
Section titled “What a Surface Is Not”A PES is indispensable, but several neighboring objects must remain distinct.
| Object | Precise meaning |
|---|---|
| Born–Oppenheimer surface | A clamped-nuclei electronic eigenvalue , with a stated convention for |
| Corrected adiabatic surface | plus specified diagonal finite-mass or other corrections |
| Diabatic model | A smooth matrix whose eigenvalues reproduce coupled adiabatic surfaces |
| Force field | An analytic or learned approximation , often with a restricted chemical domain |
| Free-energy profile | A temperature- and ensemble-dependent reduction in which other coordinates have been averaged |
| Molecular spectrum | Eigenvalues or resonances obtained only after nuclear motion and required couplings are included |
A PES is not directly observed as a complete function. Experiments constrain it through rovibrational levels, scattering cross sections, reaction rates, equilibrium structures, diffraction data, and other observables after a nuclear model has been supplied. Inverse reconstruction is therefore model dependent and may be nonunique.
Nor is a surface an exact molecular spectrum with the nuclei permanently fixed. Zero-point motion, rotation, tunneling, exchange symmetry, nonadiabatic coupling, and external conditions still matter.
Stationary Points and Molecular Structure
Section titled “Stationary Points and Molecular Structure”A stationary geometry satisfies
For a nondegenerate stationary point, its local character is determined by the Hessian
After translations, rotations, constraints, and redundant coordinates have been handled, the index is the number of negative Hessian eigenvalues:
| Index | Local interpretation |
|---|---|
| minimum | |
| first-order saddle; candidate elementary transition structure | |
| higher-order saddle | |
| local maximum |
Additional internal zero eigenvalues require higher-order terms: and have identical zero Hessians but opposite stability. A positive-semidefinite Cartesian Hessian with six zero eigenvalues is not yet evidence of six floppy vibrations: for a nonlinear isolated molecule those zero modes represent three translations and three rotations. A linear molecule has five such external zero modes.
Why the classification is coordinate independent
Section titled “Why the classification is coordinate independent”Under a smooth nonsingular change , the Hessian transforms as
At a stationary point, the second line vanishes. The remaining congruence transformation preserves the numbers of positive, negative, and zero directions. Away from a stationary point, an ordinary coordinate Hessian is not a tensor; covariant derivatives or a specified coordinate convention are required.
Local, global, and symmetry-related minima
Section titled “Local, global, and symmetry-related minima”A local minimum is lower than nearby geometries. A global minimum is lowest over the stated domain and electronic state. Distinct local minima may be conformers, structural isomers, or symmetry-related copies of one physical arrangement.
The lowest electronic minimum need not be the most populated structure at finite temperature. Entropy, zero-point energy, nuclear spin statistics, and barriers between basins affect observed populations and interconversion.
Equilibrium versus averaged structure
Section titled “Equilibrium versus averaged structure”The minimum of an electronic surface is an equilibrium geometry in the clamped-nuclei model. A rotationally or vibrationally inferred structure samples a nuclear wavefunction and is generally different:
Isotope substitution leaves unchanged to leading order but changes the nuclear wavefunction, vibrational averaging, zero-point energy, and the DBOC. This distinction matters in precision structural work.
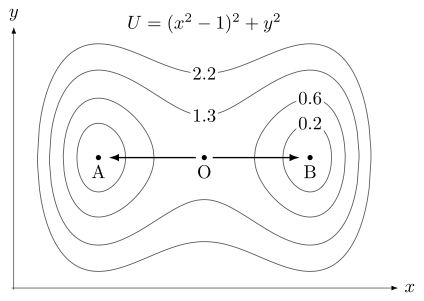
Contours of the model : A and B are minima at and O is the index-one saddle. The straight path joins them; arrows mark the two downhill branches from O, not a time-dependent trajectory. The contour labels give in model units.
Vibrational Motion Near a Minimum
Section titled “Vibrational Motion Near a Minimum”Let be a nondegenerate minimum and write . The local expansion is
In mass-weighted Cartesian coordinates, the projected Hessian
has internal eigenvectors satisfying
Positive eigenvalues give harmonic frequencies. The local nuclear Hamiltonian becomes a sum of oscillators,
with approximate energies
This is the molecular realization of the oscillator as a universal local model. The approximation is local. Anharmonicity, mode coupling, torsion, inversion, dissociation, and tunneling probe higher derivatives and distant regions of the surface.
For a single bond coordinate, Vibrations of Diatomics turns this curvature into isotope-dependent levels, tests anharmonic and Morse descriptions, and shows why local constants do not uniquely determine dissociation.
At an index-one saddle, one projected Hessian eigenvalue is negative. Electronic-structure programs often report the corresponding direction as an imaginary frequency. That label diagnoses local negative curvature; it is not an oscillatory normal mode and does not by itself prove that the saddle connects the intended reactant and product.
Reaction Coordinates
Section titled “Reaction Coordinates”A reaction coordinate is a function
chosen to summarize progress through a process. It might be a bond-length difference, an angle, a collective variable, a path parameter, or a learned function. Its level set
contains all geometries assigned the same progress value.
Reducing a high-dimensional molecule to discards information. A good coordinate for plotting energy need not be a good coordinate for kinetics, and a coordinate that separates reactants from products need not resolve hidden intermediates or competing channels.
Paths through geometry space
Section titled “Paths through geometry space”A path is a curve
Its energy profile is the composition
Different curves between the same basins generally give different profiles. The plotted profile is therefore not “the PES”; it is a one-dimensional restriction of the PES to a chosen curve.
Minimum-energy paths
Section titled “Minimum-energy paths”For a path with unit tangent in a specified metric, a minimum-energy path satisfies
Here ; in orthonormal coordinates the projector is . The gradient has no component normal to the path. Its tangential component need not vanish. Chain-of-states algorithms such as the nudged elastic band and string methods approximate this condition with a discrete sequence of geometries.
A minimum-energy path is useful for finding passes and organizing local vibrational coordinates. It is not necessarily unique, globally lowest, or dynamically dominant.
Intrinsic reaction coordinates
Section titled “Intrinsic reaction coordinates”The conventional intrinsic reaction coordinate, or IRC, follows mass-weighted steepest descent away from an index-one saddle. In mass-weighted coordinates ,
At the saddle the gradient is zero, so the integration is initialized by small displacements along the negative-curvature Hessian eigenvector in both directions. Both branches use the minus sign: their different initial displacements select the reactant and product descents. They identify the minima reached by this local construction.
The IRC depends on the mass metric and on the surface used. Nonuniform changes in isotopic masses can change the geometric path itself, even in ordinary Cartesian positions, because the descent direction is proportional to . Multiplying every mass by the same constant changes only the parameterization.
A path is not a trajectory
Section titled “A path is not a trajectory”A classical trajectory obeys a second-order equation,
and depends on initial positions and momenta. It can cross contour lines obliquely, oscillate across a valley, recross a dividing surface, or leave the neighborhood of a minimum-energy path.
A quantum nuclear wavepacket is more different still: it spreads, interferes, tunnels, and may occupy several electronic surfaces. A reaction path describes geometry; a trajectory or wavepacket additionally describes its evolution.
Barriers and Transition States
Section titled “Barriers and Transition States”For a reactant minimum and a connecting saddle , the electronic barrier on one surface is
Several quantities are often called “the barrier”:
They are not interchangeable. The first is a property of a specified PES; the others include nuclear and thermodynamic information.
Saddle point versus dynamical transition state
Section titled “Saddle point versus dynamical transition state”An index-one saddle is often called a transition structure. A transition state in rate theory is more fundamentally a dividing surface separating reactant and product regions in phase space. The best dividing surface need not pass through the lowest potential-energy saddle, especially for barrierless capture, entropic bottlenecks, roaming dynamics, strong recrossing, or reactions with several coupled coordinates.
For a unimolecular first-order rate, conventional transition-state theory gives
Here is per molecule and ; molar free energies require . Bimolecular rate constants additionally require a consistent concentration standard-state factor. A dynamical correction is often written
The interpretation of depends on convention: recrossing, tunneling, nonadiabatic transitions, and nonequilibrium preparation may be separated or bundled differently. A barrier height alone is never a rate constant.
Barrierless does not mean structureless
Section titled “Barrierless does not mean structureless”If energy decreases monotonically along a chosen entrance path, the reaction is barrierless on that path. Long-range capture, angular-momentum barriers, zero-point constraints, narrow dynamical bottlenecks, and free-energy barriers can still control the rate. Conversely, quantum tunneling can produce reaction below a classical saddle energy.
Potential Energy and Free Energy
Section titled “Potential Energy and Free Energy”At temperature , suppose all coordinates except a collective variable are averaged in a classical canonical ensemble. A potential of mean force can be written
The measure contains the coordinate metric, constraints, and any Jacobian factors. Consequently:
- depends on temperature and ensemble;
- it depends on the definition and normalization of ;
- it includes entropy from coordinates integrated out;
- it is not obtained by merely evaluating along one optimized path.
For quantum nuclei, the constrained object is defined through a density operator or path integral rather than the classical configurational integral above. Solvent, electronic excitations, and external fields can contribute further free-energy terms.
Entropic barriers
Section titled “Entropic barriers”Imagine a valley with the local form
Integrating over classically gives
A narrow region with large can therefore be a free-energy bottleneck even if has no potential barrier. Energy landscapes and free-energy landscapes answer different questions.
Asymptotic Channels
Section titled “Asymptotic Channels”A global reactive surface must describe more than stationary points. As fragments separate,
where labels a dissociation channel. The threshold must be consistent with the electronic method used in the interaction region.
Long-range interactions can contain charge–charge, charge–dipole, induction, dispersion, and anisotropic multipole terms. A flexible interpolant that fits a finite cloud of points need not extrapolate to the correct power law. Size consistency, separated-fragment degeneracies, and channel energy ordering are essential validation targets.
Local surfaces designed for spectroscopy near one minimum need not be trusted for bond breaking. Conversely, a global reactive surface may sacrifice the sub-wavenumber accuracy required for precision spectroscopy. Domain and intended observables belong in the model specification.
Crossings and Conical Intersections
Section titled “Crossings and Conical Intersections”One scalar surface is inadequate when several electronic states become close. A local real two-state Hamiltonian can be written
Its adiabatic eigenvalues are
A degeneracy requires
For a real electronic Hamiltonian, these are generically two independent conditions, with linearly independent gradients at an ordinary conical intersection. In an -dimensional internal space, conical intersections therefore form seams of dimension . The two directions that lift the degeneracy are the branching plane. With a genuinely complex Hermitian two-state Hamiltonian, a coefficient supplies a third condition, so the generic codimension changes.
The noncrossing rule in context
Section titled “The noncrossing rule in context”Along one tunable coordinate, two states of the same symmetry generically avoid crossing because one parameter cannot satisfy two independent degeneracy conditions. States protected from mixing by different exact symmetries may cross. In the full multidimensional nuclear space, same-symmetry conical-intersection seams are common because enough coordinates are available.
A one-dimensional scan can therefore be misleading:
- an avoided crossing in the scan may be a nearby conical intersection in the full space;
- a visible crossing may be symmetry protected;
- ordering states by energy can swap their electronic character.
Adiabatic and diabatic representations
Section titled “Adiabatic and diabatic representations”The diagonal adiabatic representation makes energies simple but derivative couplings can become large or singular near a degeneracy. A smooth multistate model instead uses a matrix
whose eigenvalues reproduce the adiabatic surfaces. Such a representation is usually called diabatic or quasi-diabatic. A strictly derivative-coupling-free diabatic basis need not exist: nonzero connection curvature in a truncated electronic subspace can obstruct it locally, and topology can obstruct a global construction.
Adiabatic energies alone are insufficient for nonadiabatic dynamics. One also needs derivative couplings, a diabatic matrix with off-diagonal interactions, or equivalent gauge-covariant information. Born–Oppenheimer Berry phase derives the sign change around a conical intersection.
A Solvable Two-Dimensional Landscape
Section titled “A Solvable Two-Dimensional Landscape”Consider
with , , and . The stationary equations are
There are minima at and a saddle at . The Hessian is
At either minimum,
whereas at the central point,
The central point has index one. The minimum-energy path is , and the electronic barrier is exactly
If the kinetic energy is
the minimum frequencies are
At the saddle, the unstable direction has
This model separates several ideas cleanly: stationary-point index is determined by curvature, the path is geometric, the barrier is an energy difference, and the dynamical time scales require masses.
Computational Representation
Section titled “Computational Representation”Except for very small systems, a useful surface is reconstructed from finite electronic-structure data. A typical data set is
Not every project contains every derivative or coupling. The representation should be designed around the intended observable: stationary structures need reliable gradients and Hessians, trajectories need smooth forces, spectroscopy needs delicate local curvature and anharmonicity, and scattering needs correct asymptotes.
Representation families
Section titled “Representation families”| Family | Strengths and cautions |
|---|---|
| Grids, splines, and local polynomials | Transparent for low dimension; scale poorly and can misbehave outside the grid |
| Physically motivated analytic forms | Encode dissociation and long range; may be too rigid in complex reactive regions |
| Many-body expansions | Organize fragment limits; truncation and switching functions require care |
| Permutationally invariant polynomials | Enforce exchange symmetry exactly; basis size can grow rapidly |
| Reproducing kernels and Gaussian processes | Smooth interpolation and, for Gaussian processes, uncertainty estimates; kernels and asymptotes must be chosen deliberately |
| Neural and equivariant potentials | Flexible and scalable; extrapolation, data coverage, and physical constraints remain central |
| Diabatic matrix fits | Represent coupled surfaces and crossings; gauge consistency and off-diagonal data add difficulty |
The phrase machine-learned potential identifies the regression strategy, not the underlying physical accuracy. A model trained on density-functional data reproduces that reference level plus fitting error; it does not silently become exact electronic structure.
Required invariances
Section titled “Required invariances”For an isolated molecule, the fitted energy should satisfy
for every rigid rotation and translation . Exchanging identical nuclei must also leave the energy unchanged:
These properties may be built through internal coordinates, invariant descriptors, symmetrized polynomial bases, equivariant architectures followed by scalar readout, or explicit data augmentation. Exact architectural invariance is usually more reliable than hoping a finite training set teaches it everywhere.
Energy–force consistency
Section titled “Energy–force consistency”When forces are used for dynamics, they should derive from one scalar model:
This guarantees conservative forces within the modeled isolated system. Fitting independent force components without integrability can produce path-dependent work and energy drift.
A common joint objective is
where regularizes the representation. The weights define which errors the fit prioritizes; they are part of the model, not mere numerical decoration.
Testing a fitted surface
Section titled “Testing a fitted surface”Near-equilibrium data rarely constrain repulsive walls, competing conformers, bond breaking, or a crossing. Test those regions with independent electronic calculations before using the fit there. Splitting adjacent frames from one trajectory randomly between training and testing exaggerates independence; separating trajectories, basins, or reaction channels is more informative.
Check the quantities needed for the application: gradients and Hessians for structure and vibration, asymptotic thresholds for scattering, and couplings for multistate dynamics. A small average energy error can hide a large error in a narrow reactive region. Conservative fitted forces support energy conservation, but an actual trajectory also needs a converged numerical integrator.
Separate errors of the physical Hamiltonian, electronic approximation, fit, and nuclear dynamics. Their effects can cancel for one observable. Posterior variance or agreement among related fits measures neither shared electronic bias nor unrepresented physics. The coordinate conventions, numerical implementation, reference data, and domain of the fit must be available to reproduce its predictions.
Exercises
Section titled “Exercises”1. When the Hessian cannot decide
Section titled “1. When the Hessian cannot decide”Classify the origin for and . What would an index-only classification miss?
Solution
Both have zero gradient and Hessian , with no negative eigenvalues. Yet has a strict minimum, whereas decreases along and increases along , making the origin a degenerate saddle. The zero internal curvature makes the nondegenerate Hessian classification inconclusive.
2. Coordinate changes and Hessian index
Section titled “2. Coordinate changes and Hessian index”Explain why a nonlinear coordinate change can alter the ordinary Hessian away from a stationary point but cannot change the index of a nondegenerate stationary point.
Solution
For ,
The second line is generally nonzero, so the ordinary Hessian is not a tensor away from stationarity. At a stationary point , leaving
For nonsingular , Sylvester’s law of inertia says that this congruence preserves the numbers of positive and negative eigenvalues. The index is therefore invariant.
3. Path versus trajectory
Section titled “3. Path versus trajectory”A minimum-energy path descends from a saddle to a reactant minimum. Must a classical trajectory launched near the saddle follow that curve? Give two reasons.
Solution
No. First, a minimum-energy path is defined by a first-order geometric condition on the gradient normal to the path, whereas a trajectory obeys the second-order Newton equation and carries momentum. A transverse initial velocity immediately moves the trajectory away from the path.
Second, the path is usually defined with a chosen mass or coordinate metric. Dynamics also depends on the full mass matrix and can exchange energy among transverse modes. Recrossing and valley oscillation are therefore possible even on a perfectly known surface.
4. An entropic bottleneck
Section titled “4. An entropic bottleneck”For
integrate out in a classical canonical ensemble and show how a narrow valley alters the free-energy profile.
Solution
At fixed ,
Hence
Large means a narrow transverse distribution and less configurational entropy. It raises even if is flat or decreasing.
5. Isotopes and descent directions
Section titled “5. Isotopes and descent directions”At one geometry let in two Cartesian coordinates. Compare the directions of a mass-weighted descent for and .
Solution
Up to a positive normalization, . The first direction is , with slope ; the second is , with slope . The surface is unchanged, but the paths have different tangent directions. A common rescaling of both masses would leave that direction unchanged.
References
Section titled “References”- A. P. Bartók, M. C. Payne, R. Kondor, and G. Csányi, “Gaussian approximation potentials: the accuracy of quantum mechanics, without the electrons,” Physical Review Letters 104, 136403 (2010), doi:10.1103/PhysRevLett.104.136403.
- J. Behler and M. Parrinello, “Generalized neural-network representation of high-dimensional potential-energy surfaces,” Physical Review Letters 98, 146401 (2007), doi:10.1103/PhysRevLett.98.146401.
- M. Born and K. Huang, Dynamical Theory of Crystal Lattices, Oxford University Press (1954), Chapters IV and V.
- M. Born and R. Oppenheimer, “Zur Quantentheorie der Molekeln,” Annalen der Physik 389, 457–484 (1927), doi:10.1002/andp.19273892002.
- J. M. Bowman, B. J. Braams, S. Carter, C. Chen, G. Czakó, B. Fu, X. Huang, E. Kamarchik, A. R. Sharma, B. C. Shepler, Y. Wang, and Z. Xie, “Ab initio-based potential energy surfaces for complex molecules and molecular complexes,” Journal of Physical Chemistry Letters 1, 1866–1874 (2010), doi:10.1021/jz100626h.
- S. Chmiela, A. Tkatchenko, H. E. Sauceda, I. Poltavsky, K. T. Schütt, and K.-R. Müller, “Machine learning of accurate energy-conserving molecular force fields,” Science Advances 3, e1603015 (2017), doi:10.1126/sciadv.1603015.
- H. Eyring, “The activated complex in chemical reactions,” Journal of Chemical Physics 3, 107–115 (1935), doi:10.1063/1.1749604.
- K. Fukui, “Formulation of the reaction coordinate,” Journal of Physical Chemistry 74, 4161–4163 (1970), doi:10.1021/j100717a029.
- K. Fukui, “The path of chemical reactions: the IRC approach,” Accounts of Chemical Research 14, 363–368 (1981), doi:10.1021/ar00072a001.
- G. Henkelman, B. P. Uberuaga, and H. Jónsson, “A climbing image nudged elastic band method for finding saddle points and minimum energy paths,” Journal of Chemical Physics 113, 9901–9904 (2000), doi:10.1063/1.1329672.
- T.-S. Ho, T. Hollebeek, H. Rabitz, L. B. Harding, and G. C. Schatz, “A global HO potential energy surface for the reaction O() + H → OH + H,” Journal of Chemical Physics 105, 10472–10486 (1996), doi:10.1063/1.472977.
- T.-S. Ho and H. Rabitz, “Reproducing kernel Hilbert space interpolation methods as a paradigm of high dimensional model representations,” Journal of Chemical Physics 119, 6433–6442 (2003), doi:10.1063/1.1603219.
- H. C. Longuet-Higgins, U. Öpik, M. H. L. Pryce, and R. A. Sack, “Studies of the Jahn–Teller effect. II. The dynamical problem,” Proceedings of the Royal Society A 244, 1–16 (1958), doi:10.1098/rspa.1958.0022.
- C. A. Mead and D. G. Truhlar, “On the determination of Born–Oppenheimer nuclear motion wave functions including complications due to conical intersections and identical nuclei,” Journal of Chemical Physics 70, 2284–2296 (1979), doi:10.1063/1.437734.
- W. H. Miller, N. C. Handy, and J. E. Adams, “Reaction path Hamiltonian for polyatomic molecules,” Journal of Chemical Physics 72, 99–112 (1980), doi:10.1063/1.438959.
- J. N. Murrell and K. J. Laidler, “Symmetries of activated complexes,” Transactions of the Faraday Society 64, 371–377 (1968), doi:10.1039/TF9686400371.
- J. von Neumann and E. Wigner, “Über das Verhalten von Eigenwerten bei adiabatischen Prozessen,” Physikalische Zeitschrift 30, 467–470 (1929).
- H. Partridge and D. W. Schwenke, “The determination of an accurate isotope dependent potential energy surface for water from extensive ab initio calculations and experimental data,” Journal of Chemical Physics 106, 4618–4639 (1997), doi:10.1063/1.473987.
- E. Teller, “The crossing of potential surfaces,” Journal of Physical Chemistry 41, 109–116 (1937), doi:10.1021/j150379a010.
- D. G. Truhlar and B. C. Garrett, “Variational transition-state theory,” Accounts of Chemical Research 13, 440–448 (1980), doi:10.1021/ar50156a002.
- Z. Xie and J. M. Bowman, “Permutationally invariant polynomial basis for molecular energy surface fitting via monomial symmetrization,” Journal of Chemical Theory and Computation 6, 26–34 (2010), doi:10.1021/ct9004917.
- D. R. Yarkony, “Diabolical conical intersections,” Reviews of Modern Physics 68, 985–1013 (1996), doi:10.1103/RevModPhys.68.985.