

Molecular Orbitals
A molecular orbital is a one-electron function used to represent electronic structure at a specified molecular geometry. It may be an exact eigenfunction for a genuine one-electron molecule, an eigenfunction of a mean-field operator such as the Hartree–Fock or Kohn–Sham operator, or a member of some other deliberately chosen one-particle basis. The word orbital therefore identifies a mathematical role only after the Hamiltonian, approximation, basis, geometry, and spin convention have been stated.
Molecular-orbital language is powerful because it turns a complicated electronic problem into a sequence of one-particle ideas: atom-centered functions mix, symmetry filters the allowed mixing, levels split, electrons occupy spin-orbitals, and the resulting configurations organize bonding and spectroscopy. The price is conceptual discipline. An orbital diagram is not the exact many-electron wavefunction, an orbital coefficient is not an observable, and a HOMO–LUMO separation is not automatically an excitation energy.
Required background. Molecular Hamiltonian and Born–Oppenheimer separation define the fixed-geometry problem; atomic orbitals distinguishes one-particle functions from many-electron states; change of basis supplies matrix representations.
The Fixed-Geometry One-Particle Problem
Section titled “The Fixed-Geometry One-Particle Problem”Use atomic units for electronic energies and Coulomb operators below. Choose a nuclear geometry and solve the clamped-nuclei electronic problem at that geometry. A molecular orbital is a function of one electron coordinate,
or, when spin is included explicitly, a spin-orbital
The geometry label matters. Changing a bond length or angle changes the electronic operator and therefore changes the orbitals, their ordering, and often their qualitative character.
Exact only for a one-electron electronic problem
Section titled “Exact only for a one-electron electronic problem”For a one-electron molecular ion, the clamped-nuclei electronic equation has the form
Apart from the stated nonrelativistic and fixed-nucleus approximations, can then be an exact electronic eigenstate.
For several electrons, the Coulomb interaction depends on pairs of coordinates,
so the exact state is a function
not a list of independent one-electron wavefunctions. Hartree–Fock, Kohn–Sham density-functional theory, configuration-interaction expansions, Green-function methods, and reduced-density-matrix analyses introduce orbitals in different ways. The same plotted shape can therefore have different physical status in different calculations.
Three objects that should not be conflated
Section titled “Three objects that should not be conflated”| Object | Typical notation | What it means |
|---|---|---|
| atom-centered basis function | a chosen function used to span a finite one-particle space | |
| molecular orbital | a linear combination or eigenfunction in that space | |
| many-electron state | the antisymmetric electronic state from which observables are calculated |
An “atomic orbital” in an LCAO calculation is commonly an atom-centered Gaussian or numerical basis function. It need not be an exact eigenstate of an isolated atom. A contracted basis may contain several functions of the same angular character on one center, polarization functions absent from the isolated occupied shell, and diffuse functions designed to represent molecular tails.
Linear Combination of Atomic Orbitals
Section titled “Linear Combination of Atomic Orbitals”Let be linearly independent atom-centered basis functions. A molecular orbital is expanded as
This is the linear combination of atomic orbitals, or LCAO, representation. The coefficients say how the chosen basis functions combine. They are representation-dependent numbers; changing the basis changes the coefficients even when the represented function changes little.
The overlap matrix is the metric
Section titled “The overlap matrix is the metric”Atom-centered functions on different nuclei are generally not orthogonal. Define
For a normalized orbital with coefficient vector ,
Mutual orthonormality of a set of molecular orbitals is
The matrix , not the Euclidean dot product of coefficient columns, supplies the inner product in a nonorthogonal basis. Thus is generally false.
Generalized secular equation
Section titled “Generalized secular equation”Let be the one-particle operator being diagonalized. It might be an exact one-electron Hamiltonian, a converged Fock operator, or a Kohn–Sham operator. Define
Stationarity of the Rayleigh quotient
gives
Collecting the eigenvectors gives the generalized matrix equation
This is an ordinary Hermitian eigenproblem only when . If is positive definite, a symmetric orthogonalization uses
and transforms the problem to
The back-transformed coefficients are .
Basis covariance and convergence
Section titled “Basis covariance and convergence”An invertible change among basis functions changes , , and together while preserving the represented subspace and generalized eigenvalues. A coefficient attached to one primitive function is therefore not invariant. Even atom-by-atom populations require a specified partitioning prescription.
Near-linear dependence is a numerical warning. If has a very small eigenvalue, some linear combination of basis functions has a very small norm. Orthogonalization then amplifies roundoff and integral error through . Practical calculations monitor overlap eigenvalues, remove redundant combinations when justified, and test whether energies and observables are stable under basis changes.
Two Equivalent Centers
Section titled “Two Equivalent Centers”The smallest molecular-orbital calculation uses two normalized real functions and related by symmetry. Let
The normalized in-phase and out-of-phase combinations are
and
Their orthogonality follows directly from the symmetry-equivalent diagonal overlaps:
Energies in the two-function model
Section titled “Energies in the two-function model”For a one-particle operator , define
and, after a real phase convention,
The generalized eigenvalues are
Their splitting is
The in-phase combination is lower than the isolated diagonal level precisely when
Under the same condition, the out-of-phase combination lies above . This criterion is more informative than saying “overlap lowers the bonding orbital.” Overlap enters both the normalization and the Hamiltonian matrix element. The energetic result comes from their combination.
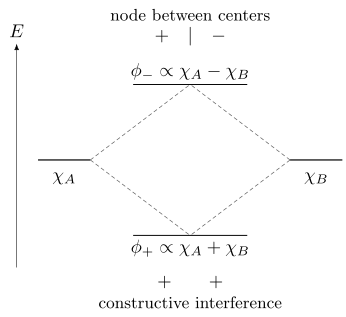
Two symmetry-equivalent atom-centered functions generate orthogonal in-phase and out-of-phase combinations. The lower combination has constructive amplitude between the centers in the usual bonding case; the upper combination has an internuclear node. The signs indicate relative wavefunction phase, not electric charge.
Interference in the density
Section titled “Interference in the density”For real basis functions,
The cross term is the interference term. In a region where and have the same sign, it increases the in-phase density and decreases the out-of-phase density. A nodal surface appears wherever the amplitudes cancel.
Phase itself is conventional: replacing by interchanges which algebraic combination is written with a plus sign. What is invariant is the spatial symmetry, nodal structure, density, and energy of each eigenfunction.
Why a node often raises the energy
Section titled “Why a node often raises the energy”A node forces stronger spatial variation. Through the kinetic operator,
that variation often increases kinetic energy. Redistribution of density also changes electron–nuclear attraction, electron–electron repulsion in a many-electron model, and nuclear-screening effects. A molecular bond cannot generally be assigned to a single term without a specified energy-decomposition method.
The two-state algebra is the same as the Tight-Binding Dimer. The molecular application differs because the atom-centered basis is usually nonorthogonal and because the one-electron operator may be self-consistent.
H₂⁺ Ion applies this two-center algebra to an actual Coulomb Hamiltonian. It evaluates the overlap and Hamiltonian integrals in closed form, adds the proton–proton term to obtain potential curves, and compares the minimal basis with an accurate one-electron benchmark.
Hydrogen Molecule shows what changes when two electrons occupy the same orbital basis: a restricted determinant acquires ionic contamination at dissociation, while mixing and restores the neutral spin-singlet limit.
Unequal Centers and Polarized Orbitals
Section titled “Unequal Centers and Polarized Orbitals”For a heteronuclear two-function illustration, first work in an orthonormal basis and choose phases so the coupling is with :
Let
The eigenvalues are
A convenient parametrization of the lower eigenvector is
where
The upper eigenvector is the orthogonal combination
Two tests control the interpretation:
- Symmetry compatibility: only basis functions belonging to compatible symmetry sectors can mix.
- Energy matching: mixing is strong when is comparable to the diagonal mismatch and weak when .
When , the lower orbital is concentrated on the lower-energy center and the upper orbital on the higher-energy center. When , both approach equal-weight combinations. This is the one-particle origin of orbital polarization in a heteronuclear bond, but partial charge and dipole moment require the occupied many-electron density, not one coefficient squared.
In a nonorthogonal basis, the same physics is obtained from
Ignoring changes both the eigenvalues and normalization and can produce a qualitatively wrong mixing analysis.
Bonding, Antibonding, and Nonbonding
Section titled “Bonding, Antibonding, and Nonbonding”The three labels describe how occupation of an orbital affects a stated molecular interaction within a stated model.
| Label | Typical signature | Interpretation and qualification |
|---|---|---|
| bonding | constructive amplitude in an internuclear region | occupation tends to stabilize shorter separation, but this must be tested against a dissociation reference or energy derivative |
| antibonding | destructive amplitude with an additional node | occupation tends to oppose the corresponding bond; the asterisk is not a universal symmetry quantum number |
| nonbonding | weak coupling to the partner fragment | occupation has a relatively small first-order effect on that bond but may still polarize, hybridize, or affect other bonds |
A label is relative to a coordinate
Section titled “A label is relative to a coordinate”An orbital can be bonding with respect to one internuclear distance and antibonding with respect to another. In a polyatomic molecule, a delocalized orbital may stabilize one set of contacts while destabilizing another. A robust local diagnostic asks how an orbital contribution or occupied state changes as the relevant nuclear coordinate is varied.
For an orbital eigenvalue model, one sometimes examines
Even that derivative is method-dependent and is not by itself the derivative of the total molecular energy. Nuclear repulsion, all occupied orbitals, relaxation, and correlation contribute to the physical force.
The simple MO bond-order index
Section titled “The simple MO bond-order index”For a diagram with clearly paired bonding and antibonding levels, the elementary index is
Two electrons in a bonding level and none in its antibonding partner give ; equal populations give zero. This is a useful bookkeeping rule in minimal models. It is not a unique observable or a universal definition of chemical bond order. Basis choice, delocalization, correlation, orbital relaxation, and the choice of which levels form a pair all matter.
Population analyses, density-based indices, valence-bond weights, and energy decompositions answer related but different questions. Numerical bond orders should therefore be reported with their definition and method.
Sigma, Pi, and Delta Orbitals
Section titled “Sigma, Pi, and Delta Orbitals”For a linear molecule, choose the internuclear axis as . In the ideal cylindrically symmetric clamped-nuclei problem, the projection of one-electron orbital angular momentum on the axis can label an orbital. If
then the magnitude
is denoted
for one-electron orbitals. Uppercase usually label the projection of the total electronic orbital angular momentum of an electronic term. An orbital label and a many-electron term label are not interchangeable.
Sigma orbitals
Section titled “Sigma orbitals”A orbital has and is invariant under rotation about the molecular axis, apart from its trivial phase. Atom-centered functions and functions can contribute to a block when the axis is . The label says nothing by itself about bonding:
may denote bonding and antibonding members of a qualitative pair, but both have axial character.
Pi and higher axial character
Section titled “Pi and higher axial character”A orbital has . For a cylindrically symmetric, spin-independent operator with no magnetic field, reflection or complex conjugation relates the degenerate and functions. Axial rotations alone do not require this degeneracy: an axial magnetic field can preserve those rotations while splitting the two projections. Real and orbitals are unitary combinations of those complex eigenfunctions and span the same two-dimensional subspace.
Similarly, denotes . Bending a linear molecule or placing it in an anisotropic environment lowers the symmetry and can split or mix components according to the remaining point group.
Inversion and reflection labels are independent
Section titled “Inversion and reflection labels are independent”For a centrosymmetric molecule, inversion through the center gives an additional parity label:
The label is not the same as bonding/antibonding. For example, whether an in-phase combination is or depends on the intrinsic parity and orientation of the contributing atom-centered functions.
For a many-electron term, a superscript or can specify reflection symmetry in a plane containing the molecular axis. That sign also does not mean bonding or antibonding.
Polyatomic molecules
Section titled “Polyatomic molecules”In a nonlinear molecule, point-group irreducible representations replace the continuous axial label. Matrix elements between basis functions of incompatible irreducible representations vanish when the operator respects the symmetry. The secular problem therefore block-diagonalizes:
Only functions within compatible symmetry blocks can mix. Low symmetry permits more mixing; it does not guarantee strong mixing, because energetic mismatch can still suppress it. Molecular Symmetry develops character reduction, projection operators, and symmetry-adapted orbital combinations; Symmetry Applications to Molecular Physics supplies the wider application map.
How to Build and Read an MO Diagram
Section titled “How to Build and Read an MO Diagram”A qualitative diagram compresses a matrix eigenproblem. Start with fragment functions at a fixed geometry, sort them into symmetry sectors, and mix functions within each sector according to their couplings and energy mismatch. Fill the resulting spin-orbitals, then determine the spin and spatial symmetry of the many-electron state.
The vertical scale may represent an exact one-electron eigenvalue, a mean-field eigenvalue, or only an ordering. The two-function H₂⁺ calculation makes the overlap matrix and energy convention explicit.
Occupations and spin
Section titled “Occupations and spin”For orthonormal spin-orbitals, the Pauli principle permits occupation number zero or one. In a restricted closed-shell spatial-orbital model, two electrons of opposite spin occupy each spatial orbital. The density is then
In an unrestricted or open-shell model, separate spin densities are needed:
The total density and spin density are
Degenerate or nearly degenerate levels require care. Filling them to maximize spin may be a useful first model, but the actual state ordering depends on electron repulsion, symmetry, spin–orbit interaction, vibronic coupling, and correlation.
A configuration is not yet an electronic state
Section titled “A configuration is not yet an electronic state”An occupation string such as
specifies a configuration in a chosen orbital basis. It does not by itself specify all spin couplings, spatial symmetry, or configuration mixing. Several symmetry-adapted states can arise from the same open-shell occupation pattern.
Slater Determinants explains the antisymmetric construction. A determinant built from occupied spin-orbitals is
That determinant is a many-electron wavefunction. The orbital diagram is only a compact specification of the one-particle ingredients used to build it.
Canonical and Localized Molecular Orbitals
Section titled “Canonical and Localized Molecular Orbitals”Canonical Hartree–Fock orbitals diagonalize the converged Fock operator,
They are often delocalized and symmetry-adapted. Let the occupied projector be
For any unitary matrix acting within the occupied subspace,
and
For occupied spin-orbitals the determinant changes only by the overall phase . If these are occupied spatial orbitals rotated identically in both spin sectors of a closed shell, the phase is . The density and Hartree–Fock energy do not change in either case.
Localized bond and lone-pair orbitals exploit this freedom by optimizing a localization criterion. Boys, Edmiston–Ruedenberg, and Pipek–Mezey orbitals generally differ. None is the uniquely correct set of bonds hidden beneath the canonical solution. Each is a chosen basis for a subspace, useful when its criterion matches the question.
Rotations that mix occupied and virtual orbitals are different: they generally change the determinant and its energy. Likewise, rotating nondegenerate canonical orbitals produces functions that no longer diagonalize the Fock operator, even though they may span the same selected space.
Orbital Energies
Section titled “Orbital Energies”An orbital energy has meaning only relative to the operator that generated it.
| Orbital construction | Meaning of | What it is not automatically |
|---|---|---|
| exact one-electron molecule | eigenvalue of the clamped-nuclei one-electron Hamiltonian | a full rovibronic transition energy |
| canonical Hartree–Fock | Lagrange-multiplier eigenvalue of the converged Fock operator | an exact removal, addition, or neutral excitation energy |
| Kohn–Sham | eigenvalue of an auxiliary density-reproducing system | a generic quasiparticle spectrum |
| localized or natural orbital | no unique energy unless an extra operator or convention is supplied | an eigenstate energy merely because the orbital is plotted |
Hartree–Fock eigenvalues are not additive
Section titled “Hartree–Fock eigenvalues are not additive”In an orthonormal spin-orbital notation,
The electronic Hartree–Fock energy is
Therefore
A naive sum of occupied orbital energies double-counts the mean electron–electron interaction. Internuclear repulsion must also be added for the fixed-geometry molecular energy when it is not included in the electronic Hamiltonian convention.
Koopmans’ frozen-orbital statement
Section titled “Koopmans’ frozen-orbital statement”For canonical restricted closed-shell or unrestricted Hartree–Fock spin-orbitals, removing an electron from occupied orbital without allowing any other orbital to relax gives the exact difference between those two frozen determinants:
Approximating the physical ionization energy by this frozen difference is Koopmans’ approximation. Restricted open-shell orbital energies require a separate convention and do not all obey this simple interpretation. Orbital relaxation, electron correlation, different ionic-state mixing, relativistic effects, and nuclear motion modify the measured ionization energy. Virtual Hartree–Fock eigenvalues have no equally direct general interpretation as electron affinities.
HOMO, LUMO, and Frontier Language
Section titled “HOMO, LUMO, and Frontier Language”For a specified one-particle spectrum:
- HOMO means the highest occupied molecular orbital;
- LUMO means the lowest unoccupied molecular orbital;
- SOMO means a singly occupied molecular orbital in an open-shell description;
- frontier orbitals usually means orbitals near the occupation boundary that dominate a chosen low-energy interaction or reactivity model.
These labels depend on electron number, geometry, spin treatment, charge state, external field, method, and basis. In an unrestricted calculation there can be separate and frontier levels. Degeneracy may produce a frontier subspace rather than one unique orbital.
Three gaps that must be separated
Section titled “Three gaps that must be separated”The orbital gap is
At the same nuclear geometry, the exact fundamental charge gap is
A neutral excitation energy is
An optical absorption onset additionally depends on transition matrix elements, selection rules, temperature, line broadening, and nuclear motion. In general,
are different quantities.
For the multiplicative-potential form of exact ground-state Kohn–Sham theory, with consistent integer-electron ground states and the particle-number limits of ensemble DFT, the fundamental gap can be written
where is the exchange-correlation derivative discontinuity. Approximate functionals and generalized Kohn–Sham schemes require their own interpretation. A small computed HOMO–LUMO gap is therefore a warning about low-energy orbital rearrangement, not a universal measured excitation energy.
Frontier-orbital reasoning
Section titled “Frontier-orbital reasoning”Frontier-orbital theory asks whether an occupied orbital of one reactant and an unoccupied orbital of another:
- have compatible symmetry;
- overlap in the reacting region;
- have a favorable energy separation;
- retain the assumed character along the reaction coordinate.
This can organize trends and selection rules. It does not replace a potential-energy surface, transition-state calculation, solvent model, spin analysis, or nonadiabatic treatment. A reaction is governed by the total many-electron energy and dynamics, not by the visual proximity of two orbital lines.
Molecular Orbitals Versus Exact Many-Electron States
Section titled “Molecular Orbitals Versus Exact Many-Electron States”A correlated electronic eigenstate can be expanded in determinants,
The orbitals and configuration coefficients together define the representation. A unitary change of the complete one-particle basis changes the determinant expansion but not the exact vector represented by a full configuration-interaction expansion.
The one-body reduced density matrix
Section titled “The one-body reduced density matrix”For a normalized -electron state, the spin-orbital one-body density matrix is
Its diagonal gives the one-electron density,
Natural spin-orbitals diagonalize :
with
A single determinant has occupation numbers equal to one and all others zero. For a pure electronic state, fractional natural occupations establish that the state is not a single determinant. A statistical mixture can also have fractional occupations without each component being correlated. Reduced Density Matrices develops the formalism.
State-specific removal amplitudes
Section titled “State-specific removal amplitudes”Photoelectron observables are often better connected to a Dyson orbital, which overlaps an -electron state with a specified -electron ionic state:
This is a transition amplitude between two many-electron states, not proof that a particular electron permanently occupied a visible orbital. Different ionic channels have different Dyson orbitals and removal energies.
Photoelectron Spectroscopy shows how those channel-specific overlaps enter measured electron energies, intensities, satellites, and angular distributions.
Worked Example: Heteronuclear Two-Level Mixing
Section titled “Worked Example: Heteronuclear Two-Level Mixing”Consider the orthonormal matrix, in electronvolts,
Here
The eigenvalues are
so
The mixing angle satisfies
which gives
The lower-orbital weights in this orthonormal two-function model are
The lower orbital is therefore strongly polarized toward the lower-energy fragment function. Coupling stabilizes it by relative to and destabilizes the upper level by the same amount relative to .
This calculation does not imply a charge on fragment . Charge requires the occupations of all relevant orbitals, the overlap metric if the basis is nonorthogonal, and an explicitly chosen real-space or population partition.
Exercises
Section titled “Exercises”1. Change the phase of a fragment orbital
Section titled “1. Change the phase of a fragment orbital”Replace by in the equivalent-center problem. What happens to , , and ? Does a bonding state become antibonding?
Solution
Both and change sign, so the formulas called and exchange names. The represented physical functions, energies, and densities are unchanged: the previously in-phase state is now written as a difference. Bonding character is not determined by the written plus sign.
2. Explore heteronuclear mixing
Section titled “2. Explore heteronuclear mixing”For
show that the lower-state weight on is
Evaluate the limits and .
Solution
With
one has
Because the lower state is
its weight is
which gives the stated result. If , then : the state localizes on the lower-energy center. If , then : equivalent centers give equal mixing.
3. Separate axial, inversion, and bonding labels
Section titled “3. Separate axial, inversion, and bonding labels”For a centrosymmetric linear molecule, explain why each of the following statements is false:
- every orbital is bonding;
- every antibonding orbital is ungerade;
- the sign on a electronic term means constructive orbital interference.
Solution
states have ; this is an axial angular-momentum label. Both bonding and antibonding orbitals can occur.
Gerade and ungerade describe inversion parity. Bonding character describes the energetic effect of occupation along a coordinate. Their relation depends on the intrinsic parity and orientation of the fragment functions, so antibonding does not universally imply .
For a electronic term, the superscript means even reflection parity in a plane containing the molecular axis. It is a many-electron spatial-symmetry label, not a sign between two basis functions.
4. Prove occupied-subspace invariance
Section titled “4. Prove occupied-subspace invariance”Let be orthonormal occupied orbitals and define
where is unitary. Show that the occupied projector and closed-shell density are unchanged.
Solution
The transformed projector is
The closed-shell density is twice the position-space diagonal:
It is therefore unchanged. Individual orbital shapes can change even though the determinant ray, projector, and density do not.
5. Distinguish three energy gaps
Section titled “5. Distinguish three energy gaps”Suppose a calculation reports
while total-energy differences give and . The first neutral excited state lies above the ground state but is dipole forbidden. Compute the orbital and fundamental gaps, and state what additional information is needed for the optical onset.
Solution
The orbital gap is
The fundamental gap is
The lowest neutral excitation is , already distinct from both. Because that state is dipole forbidden, the absorption onset requires transition moments to higher states, vibronic intensity borrowing, temperature and nuclear-motion effects, and an experimental detection threshold and line-shape convention.
6. Diagnose a correlated state
Section titled “6. Diagnose a correlated state”A pure four-electron state has spin-orbital natural occupations
with all remaining occupations negligible. Verify the trace and explain why no single four-orbital determinant exactly represents the state.
Solution
The occupations sum to
as required for four electrons. A single determinant would have exactly four spin-orbital occupations equal to one and all others equal to zero. Here occupation has leaked into additional natural orbitals, while the leading four are depleted. The state is therefore not exactly a single determinant in any spin-orbital basis.
References
Section titled “References”- P. W. Atkins and R. S. Friedman, Molecular Quantum Mechanics, 5th ed. (Oxford University Press, 2011), Chapters 8–10.
- S. F. Boys, “Construction of Some Molecular Orbitals to Be Approximately Invariant for Changes from One Molecule to Another,” Reviews of Modern Physics 32, 296–299 (1960), doi:10.1103/RevModPhys.32.296.
- F. A. Cotton, Chemical Applications of Group Theory, 3rd ed. (Wiley, 1990), Chapters 3–9.
- C. Edmiston and K. Ruedenberg, “Localized Atomic and Molecular Orbitals,” Reviews of Modern Physics 35, 457–465 (1963), doi:10.1103/RevModPhys.35.457.
- K. Fukui, T. Yonezawa, and H. Shingu, “A Molecular Orbital Theory of Reactivity in Aromatic Hydrocarbons,” Journal of Chemical Physics 20, 722–725 (1952), doi:10.1063/1.1700523.
- T. Helgaker, P. Jørgensen, and J. Olsen, Molecular Electronic-Structure Theory (Wiley, 2000), Chapters 2, 10, and 11, doi:10.1002/9781119019572.
- G. Herzberg, Molecular Spectra and Molecular Structure. I. Spectra of Diatomic Molecules, 2nd ed. (Van Nostrand, 1950), Chapters I–III.
- J. F. Janak, “Proof That in Density-Functional Theory,” Physical Review B 18, 7165–7168 (1978), doi:10.1103/PhysRevB.18.7165.
- W. Kohn and L. J. Sham, “Self-Consistent Equations Including Exchange and Correlation Effects,” Physical Review 140, A1133–A1138 (1965), doi:10.1103/PhysRev.140.A1133.
- T. Koopmans, “Über die Zuordnung von Wellenfunktionen und Eigenwerten zu den einzelnen Elektronen eines Atoms,” Physica 1, 104–113 (1934), doi:10.1016/S0031-8914(34)90011-2.
- A. I. Krylov, “From Orbitals to Observables and Back,” Journal of Chemical Physics 153, 080901 (2020), doi:10.1063/5.0018597.
- J. E. Lennard-Jones, “The Electronic Structure of Some Diatomic Molecules,” Transactions of the Faraday Society 25, 668–686 (1929), doi:10.1039/TF9292500668.
- I. N. Levine, Quantum Chemistry, 7th ed. (Pearson, 2014), Chapters 8–15.
- P.-O. Löwdin, “Quantum Theory of Many-Particle Systems. I. Physical Interpretations by Means of Density Matrices, Natural Spin-Orbitals, and Convergence Problems in the Method of Configurational Interaction,” Physical Review 97, 1474–1489 (1955), doi:10.1103/PhysRev.97.1474.
- R. McWeeny, Methods of Molecular Quantum Mechanics, 2nd ed. (Academic Press, 1992), Chapters 3–6.
- R. S. Mulliken, “Electronic Structures of Polyatomic Molecules and Valence,” Physical Review 40, 55–62 (1932), doi:10.1103/PhysRev.40.55.
- R. S. Mulliken, “Electronic Structures of Polyatomic Molecules and Valence. II. General Considerations,” Physical Review 41, 49–71 (1932), doi:10.1103/PhysRev.41.49.
- J. P. Perdew and M. Levy, “Physical Content of the Exact Kohn–Sham Orbital Energies: Band Gaps and Derivative Discontinuities,” Physical Review Letters 51, 1884–1887 (1983), doi:10.1103/PhysRevLett.51.1884.
- J. Pipek and P. G. Mezey, “A Fast Intrinsic Localization Procedure Applicable for ab initio and Semiempirical Linear Combination of Atomic Orbital Wave Functions,” Journal of Chemical Physics 90, 4916–4926 (1989), doi:10.1063/1.456588.
- C. C. J. Roothaan, “New Developments in Molecular Orbital Theory,” Reviews of Modern Physics 23, 69–89 (1951), doi:10.1103/RevModPhys.23.69.
- L. J. Sham and M. Schlüter, “Density-Functional Theory of the Energy Gap,” Physical Review Letters 51, 1888–1891 (1983), doi:10.1103/PhysRevLett.51.1888.
- A. Szabo and N. S. Ostlund, Modern Quantum Chemistry: Introduction to Advanced Electronic Structure Theory (Dover, 1996), Chapters 2–3.