

Rotations of Molecules
Molecular rotation is the quantized reorientation of a molecule’s mass distribution in space. In a semirigid molecule, rotational spacings are usually much smaller than vibrational and electronic spacings, so a rotor Hamiltonian is often the first effective theory of nuclear motion on a potential-energy surface.
Rotational constants measure inverse moments of inertia. Turning those constants into bond lengths requires a model of vibrational averaging and, for polyatomic molecules, enough isotopic or structural information to constrain the geometry.
Required background. Molecular Hamiltonian and Born–Oppenheimer separation define the nuclear problem. Rigid rotor and rotational spectra derive the angular eigenstates and ideal line ladder used here.
From Molecular Motion to a Rotor
Section titled “From Molecular Motion to a Rotor”Separate translation first
Section titled “Separate translation first”For nuclei with laboratory positions and masses , define the nuclear center of mass
In the leading Born–Oppenheimer nuclear model, coordinates relative to this nuclear center of mass separate a free translational term from internal motion. Exact separation of the whole molecule uses the center of mass of nuclei and electrons and retains finite-electron-mass recoil corrections; see Molecular Hamiltonian.
On one Born–Oppenheimer surface, an internal nuclear Hamiltonian has the schematic organization
where denotes internal shape coordinates and contains rotation–vibration coupling. The decomposition requires a molecule-fixed frame; an Eckart frame is the standard local choice near an equilibrium geometry.
Rovibrational coupling develops the joint term values, P/Q/R branches, and Coriolis effects obtained when these couplings are retained.
Freeze shape, not quantum mechanics
Section titled “Freeze shape, not quantum mechanics”For a nonlinear molecule in a nondegenerate electronic state with no electronic angular momentum, let be the nuclear inertia tensor at a reference geometry. Neglecting spin, vibration, and rotation–vibration coupling gives
In principal axes ,
For a linear molecule has a zero eigenvalue: this inverse is taken only on the two perpendicular rotational directions, not on the molecular axis.
The molecule is not literally a perfectly rigid classical object. “Rigid rotor” means that shape coordinates have been projected onto a reference vibrational state and their residual effects are initially neglected. Rotational constants measured in a vibrational state already contain vibrational averaging.
The approximation is controlled by scale separation. A useful diagnostic is
Small supports a rotor expansion, but high rotational excitation, soft vibrations, internal rotation, near-degenerate vibronic states, or dissociation can invalidate a low-order treatment.
Diatomic Rigid Rotor
Section titled “Diatomic Rigid Rotor”Internal radial Hamiltonian
Section titled “Internal radial Hamiltonian”For two nuclei, use the relative vector
For a nondegenerate electronic state, neglecting nuclear spin and electronic recoil corrections, the leading internal nuclear Hamiltonian is
The first term changes bond length, the second reorients the internuclear axis, and is the chosen adiabatic potential. Freezing at a reference value gives
The corresponding spin-free rotational states are
with energies
In zero external field, the degeneracy is . This degeneracy concerns orientation relative to a laboratory axis. Nuclear-spin statistical weights and fine or hyperfine structure are additional factors, not part of the scalar rigid-rotor result.
Two rotational degrees of freedom
Section titled “Two rotational degrees of freedom”For point nuclei on a line, rotation about the internuclear axis does not change the nuclear configuration. The linear rotor therefore has two orientational degrees of freedom. Its axial principal moment is zero in the ideal nuclear model, while the two perpendicular moments are equal:
Writing an “axial rotational constant” proportional to is not a physical extra rotational ladder. Electronic orbital angular momentum, spin, bending angular momentum, and finite-size corrections can add structure in real molecules, but they require an enlarged Hamiltonian.
When the rotational label is N
Section titled “When the rotational label is N”Here describes rotation when electronic spin and electronic angular momentum vanish. The standard more general convention is
is nuclear end-over-end rotation; excludes electron spin but includes electronic orbital angular momentum. Adding again to would double count it. Different coupling limits determine which intermediate quantum numbers remain useful.
Identical nuclei
Section titled “Identical nuclei”For a homonuclear diatomic, exchanging the nuclei reverses the directed molecular axis. The total molecular wavefunction must have the exchange symmetry required by nuclear statistics. Rotational states of alternating parity can therefore pair with different nuclear-spin species or be absent. This is why ortho and para forms cannot be handled by multiplying every level by one universal spin degeneracy.
Rotational Constants
Section titled “Rotational Constants”Three common unit conventions
Section titled “Three common unit conventions”The same inertia is encoded in several constants:
| Convention | Definition |
|---|---|
| Energy | |
| Frequency | |
| Wavenumber |
Accordingly,
Spectroscopic papers often write all three constants simply as . The units determine the meaning. In this page, , , and are kept distinct whenever ambiguity matters.
The wavenumber is a frequency divided by :
whereas the angular wavevector magnitude in vacuum is . If is reported in , must be expressed in when converting to hertz.
Equilibrium and vibrational-state constants
Section titled “Equilibrium and vibrational-state constants”For a diatomic equilibrium distance ,
The experimentally fitted ground-vibrational constant is not generally . Rotation samples a vibrational probability distribution, so approximately
For a diatomic, the standard low-order expansion is
For a polyatomic molecule with mode degeneracies ,
Thus a distance obtained directly from ,
is an effective ground-state distance, not automatically the minimum of the potential. Precision work must state whether a reported structure is , , a substitution structure , or another defined effective structure.
Moment of Inertia
Section titled “Moment of Inertia”Inertia tensor and principal axes
Section titled “Inertia tensor and principal axes”At a fixed geometry, with nuclear coordinates
the nuclear inertia tensor is
Diagonalizing this real symmetric tensor defines principal axes and principal moments. The conventional ordering is
so the frequency rotational constants obey
Translation of every nucleus by the same vector does not change the center-of-mass inertia tensor. Rotation of the coordinate axes changes its matrix entries but not its three eigenvalues.
For a planar rigid geometry, choosing perpendicular to the plane gives the perpendicular-axis relation
The ground-state inertial defect
need not vanish, even for a planar equilibrium structure. Zero-point out-of-plane motion, electronic contributions, and nonrigid effects contribute to .
Isotopic substitution
Section titled “Isotopic substitution”Changing an isotope changes masses while leaving the electronic potential nearly unchanged at the Born–Oppenheimer level. The resulting shift
provides new constraints on atomic coordinates. For a diatomic with an isotope-independent reference distance,
For a polyatomic molecule, substitution also shifts the center of mass and can rotate the principal axes. Kraitchman substitution coordinates use changes in principal moments to estimate the absolute coordinates of the substituted atom. They are powerful, but they do not recover coordinate signs by themselves and become unstable for coordinates close to a principal plane or axis.
Three cautions are essential:
- three rotational constants provide only three independent inertia constraints for one isotopologue;
- ground-state moments contain vibrational averaging;
- a highly precise fit does not by itself make the inverse structural problem unique.
Multiple isotopologues, chemical constraints, ab initio vibrational corrections, and uncertainty propagation are normally combined for a defensible equilibrium structure.
Rotational Selection Rules
Section titled “Rotational Selection Rules”The transition operator matters
Section titled “The transition operator matters”The leading interaction with a microwave electric field is
A transition requires a nonzero matrix element
Energy-level spacings alone do not determine whether a line appears. The permanent body-fixed dipole, rotational wavefunctions, polarization, parity, exchange symmetry, and any coupled spins all enter the line strength.
For a polar linear rotor in a nondegenerate closed-shell electronic and vibrational state, electric-dipole pure rotational transitions obey
Absorption normally follows . The ideal frequency is
A homonuclear diatomic has no permanent electric dipole in a field-free isolated state, so its ordinary electric-dipole pure rotational spectrum is absent. The rotational levels still exist. Rotational Raman scattering, weak multipole or magnetic mechanisms, collision-induced absorption, and rovibrational transitions probe different operators and have different rules.
Symmetric-top and asymmetric-top rules
Section titled “Symmetric-top and asymmetric-top rules”For a symmetric top, is the projection of rotational angular momentum on the symmetry axis. A dipole component parallel to that axis gives
while a perpendicular component gives
The angular rule is , with . For a parallel permanent dipole, connects degenerate rigid-top levels and gives no nonzero-frequency absorption line; the ordinary ladder has . If a perpendicular component is allowed, changing can instead give a nonzero-frequency transition. Molecular symmetry determines which dipole components exist.
An asymmetric top has no exact , but its levels are conventionally labelled by correlation with prolate and oblate symmetric-top limits. If a dipole component lies along principal axis , , or , the corresponding parity rules are:
| Transition type | Changes of limiting labels |
|---|---|
| type | even, odd |
| type | odd, odd |
| type | odd, even |
Here “even” includes zero and “odd” includes positive or negative odd integers. The general angular condition is
Mixing can lend intensity to rules based on approximate quantum numbers; it cannot remove a zero protected by an exact symmetry that the Hamiltonian and transition operator still obey. Nuclear-spin symmetry can remove or reweight levels, and spin coupling can replace by a chain of labels. Selection rules must therefore be attached to a stated Hamiltonian, operator, and coupling scheme.
Centrifugal Distortion
Section titled “Centrifugal Distortion”Why the rigid ladder bends
Section titled “Why the rigid ladder bends”At larger , the rotational kinetic energy favors a larger average bond length and therefore a larger moment of inertia. In a simple diatomic model, write
For fixed
the effective radial energy is approximately
Minimization to leading order gives
Substitution back into the energy produces
where
If , then
In spectroscopic wavenumber units this becomes the familiar scale estimate
This derivation explains the sign and scale, but fitted distortion constants are effective Hamiltonian parameters. In polyatomic molecules they summarize couplings to many vibrational coordinates, and their numerical values depend on the Hamiltonian reduction and convention.
Effective term values
Section titled “Effective term values”For a linear molecule in a fixed nondegenerate vibrational state,
where all constants and use the same frequency or wavenumber units.
Writing , the adjacent absorption line is
The positive term lowers high- lines relative to the rigid ladder. Because the correction grows as , a fit that works for low can extrapolate poorly far beyond the measured range.
Evaluated HCN example
Section titled “Evaluated HCN example”The NIST evaluated triatomic tables report the following ground-vibrational constants:
| Constant | HCN | DCN |
|---|---|---|
| () | ||
| () |
For HCN, retaining and gives:
| () | Rigid excess () | |
|---|---|---|
The increasing residual is the centrifugal-distortion signature. The nitrogen nucleus also has an electric quadrupole moment, so an observed feature resolves into several hyperfine components under sufficient resolution. A rotational centroid, a hyperfine component, and the rigid-rotor prediction are three different frequencies.
The isotope shift from HCN to DCN is much larger than the distortion correction because deuteration changes the entire center-of-mass inertia. It is not legitimate to interpret that shift as a change in bond strength without a separate nuclear-motion analysis.
Symmetric and Asymmetric Tops
Section titled “Symmetric and Asymmetric Tops”General rigid-top Hamiltonian
Section titled “General rigid-top Hamiltonian”In frequency units, the principal-axis Hamiltonian is
The relations among classify the ideal rotor.
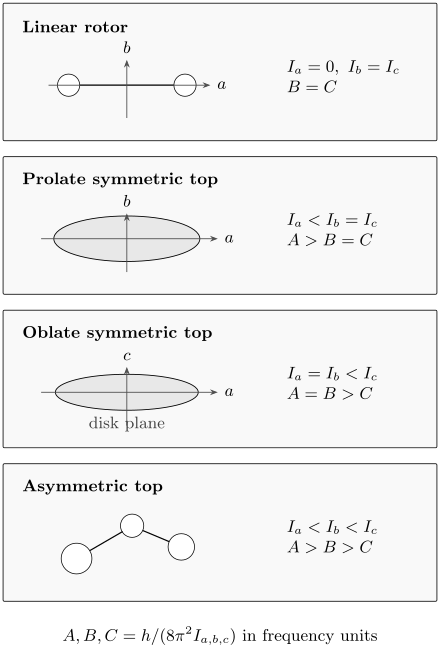
Rigid-rotor classes are statements about principal moments of inertia. Because each rotational constant is inversely proportional to its moment, the inequalities reverse between and . Molecular examples can carry inversion, internal-rotation, spin, or vibronic structure beyond the shapes shown.
Linear and spherical tops
Section titled “Linear and spherical tops”A linear rotor has
and only the two perpendicular rotations produce the scalar ladder .
A spherical top has
Its rotational Hamiltonian is also proportional to , but the configuration space is the full orientation of a three-dimensional body rather than only a molecular axis. Methane and sulfur hexafluoride are standard high-symmetry examples, although their ordinary pure rotational electric-dipole spectra vanish in the rigid equilibrium limit because they have no permanent dipole.
Symmetric tops
Section titled “Symmetric tops”A prolate symmetric top has
while an oblate symmetric top has
Let be the constant for the symmetry axis and the common perpendicular constant. Then
with
For a prolate top, ; for an oblate top, . In a field-free rigid symmetric top the energy depends on , so and are degenerate before additional interactions are included.
Asymmetric tops
Section titled “Asymmetric tops”Most polyatomic molecules satisfy
Because no body-axis projection commutes with the full asymmetric-top Hamiltonian, there is no exact quantum number and no universal closed-form energy formula. For each , one constructs and diagonalizes a finite matrix in a symmetric-top basis.
For , Ray’s asymmetry parameter is
The spherical top makes this ratio undefined. The prolate symmetric-top limit is and the oblate limit is . The conventional labels
record correlation with those two limits. They are useful labels, not simultaneous exact projections of on two body axes.
For semirigid asymmetric tops, precision work usually adds quartic and then sextic centrifugal-distortion operators. Watson’s reduced Hamiltonians remove parameter indeterminacies, but different reductions and axis representations produce different numerical parameter sets. Constants should therefore be compared only after matching conventions.
Microwave Spectroscopy Connection
Section titled “Microwave Spectroscopy Connection”Microwave line frequencies constrain differences of effective rotational energies. Rotational spectroscopy explains line assignment, fits, and evaluated catalogs. The HCN example above illustrates why the rigid frequency, a distortion-corrected rotational centroid, and a hyperfine component must be distinguished.
Positions, strengths, and populations
Section titled “Positions, strengths, and populations”Line positions come from energy differences:
Integrated strengths also depend on the transition moment and lower-state population. For an ideal linear rotor in thermal equilibrium,
where is the nuclear-spin statistical weight. Ignoring that factor and treating as continuous, the most populated region is near
This continuous estimate is useful when several levels are populated. A negative value is replaced by the boundary ; at low temperature compare the discrete populations directly. The estimate concerns level population, not the intensity of an individual line. Dipole components, Hönl–London factors, stimulated emission, partition functions, instrumental response, optical depth, collisions, and nonequilibrium populations can all matter.
What different patterns constrain
Section titled “What different patterns constrain”| Spectroscopic evidence | Primary constraint |
|---|---|
| Line spacings and assignments | and distortion constants |
| Isotopologue shifts | mass distribution and structural coordinates |
| Stark shifts | body-fixed dipole components |
| Hyperfine splittings | nuclear quadrupole, spin-rotation, and spin-spin couplings |
| Relative intensities | populations, dipole components, and line-strength factors |
| Residual trends | missing distortion, coupling, perturbation, or misassignment |
A rotational spectrum is often a distinctive molecular fingerprint because different isomers and isotopologues have different inertia tensors. Enantiomers in an isolated achiral environment, however, have the same ordinary rotational constants. Additional chiral discrimination requires a chiral interaction or phase-sensitive multi-axis protocol.
A low-order distortion expansion is useful only over a limited range of rotation. Localized level perturbations, tunneling splittings, and large-amplitude motion require the relevant coupling or internal coordinate, not merely more powers of . Constants from different Watson reductions or axis conventions should be transformed before comparison.
Exercises
Section titled “Exercises”1. Inertia and an effective bond length
Section titled “1. Inertia and an effective bond length”A closed-shell diatomic has and reduced mass . Estimate and . Use .
Solution
In frequency units,
With ,
The reduced mass is
Therefore
Because the input is , this is an effective ground-state distance , not automatically .
2. Diatomic isotope scaling
Section titled “2. Diatomic isotope scaling”Approximate the masses of , , and by their mass numbers. If the bond length is unchanged, estimate .
Solution
The reduced masses are
and
Since at fixed distance,
The heavier isotopologue has the smaller rotational constant and more closely spaced lines.
3. Distorted HCN line
Section titled “3. Distorted HCN line”Using and , calculate the HCN centroid frequency through quartic distortion.
Solution
For ,
At ,
This is a rotational centroid within the truncated model. Nitrogen quadrupole coupling splits an actual high-resolution feature into hyperfine components.
4. Rotation and electron spin
Section titled “4. Rotation and electron spin”In a state, take and . Which values arise from and ? Neglect spin–rotation coupling and verify the number of states in each manifold.
Solution
For , only occurs, with two magnetic substates. For , angular-momentum addition gives , with substates. These agree with . In this limit is the nuclear rotor quantum number and the common energy is ; a spin–rotation interaction splits the two components.
5. Classify a rotor
Section titled “5. Classify a rotor”A molecule has principal moments in the ratio
Classify the rotor and find the ratio .
Solution
Because
the molecule is a prolate symmetric top. Rotational constants are inversely proportional to moments:
6. Identify an asymmetric-top transition type
Section titled “6. Identify an asymmetric-top transition type”Classify each transition by its -, -, or -type parity rule:
Solution
For
both changes are odd:
It is therefore type.
For
the changes are
Even and odd identify an -type transition.
7. Thermal rotational population
Section titled “7. Thermal rotational population”For a linear rotor with at , estimate the most populated , ignoring nuclear-spin weights. Use .
Solution
The continuous estimate is
Since
we find
The nearest integer is . Exact discrete populations should be compared at and , and observed line intensity still requires a transition-strength factor.
8. Precision versus structural meaning
Section titled “8. Precision versus structural meaning”An asymmetric-top fit determines to relative precision . Explain why this does not establish every equilibrium bond length and angle to relative precision .
Solution
The fitted constants determine three ground-state principal moments with high precision. A polyatomic geometry has more than three internal coordinates, so one isotopologue does not make the inverse problem unique. Moreover, include zero-point vibrational averaging, while an equilibrium structure refers to the potential minimum.
Recovering requires additional information such as multiple isotopologues, structural constraints, and calculated rotation–vibration corrections. Frequency precision, effective-Hamiltonian precision, and structural accuracy are distinct quantities.
References
Section titled “References”- P. F. Bernath, Spectra of Atoms and Molecules, 4th ed., Oxford University Press, 2020.
- J. M. Brown and A. Carrington, Rotational Spectroscopy of Diatomic Molecules, Cambridge University Press, 2003.
- P. R. Bunker and P. Jensen, Molecular Symmetry and Spectroscopy, 2nd ed., NRC Research Press, 1998.
- C. C. Costain, “Determination of Molecular Structures from Ground State Rotational Constants,” Journal of Chemical Physics 29, 864–874 (1958), doi:10.1063/1.1744602.
- C. P. Endres, S. Schlemmer, P. Schilke, J. Stutzki, and H. S. P. Müller, “The Cologne Database for Molecular Spectroscopy, CDMS, in the Virtual Atomic and Molecular Data Centre, VAMDC,” Journal of Molecular Spectroscopy 327, 95–104 (2016), doi:10.1016/j.jms.2016.03.005.
- W. Gordy and R. L. Cook, Microwave Molecular Spectra, 3rd ed., Wiley, 1984.
- J. Kraitchman, “Determination of Molecular Structure from Microwave Spectroscopic Data,” American Journal of Physics 21, 17–24 (1953), doi:10.1119/1.1933338.
- NIST Physical Measurement Laboratory, Triatomic Spectral Database: molecular parameters and energy-level formulation and evaluated HCN isotopologue constants.
- H. M. Pickett, R. L. Poynter, E. A. Cohen, M. L. Delitsky, J. C. Pearson, and H. S. P. Müller, “Submillimeter, Millimeter, and Microwave Spectral Line Catalog,” Journal of Quantitative Spectroscopy and Radiative Transfer 60, 883–890 (1998), doi:10.1016/S0022-4073(98)00091-0.
- C. H. Townes and A. L. Schawlow, Microwave Spectroscopy, McGraw–Hill, 1955; Dover reprint, 1975.
- J. K. G. Watson, “Determination of Centrifugal Distortion Coefficients of Asymmetric-Top Molecules,” Journal of Chemical Physics 46, 1935–1949 (1967), doi:10.1063/1.1840957.
- J. K. G. Watson, “Simplification of the Molecular Vibration-Rotation Hamiltonian,” Molecular Physics 15, 479–490 (1968), doi:10.1080/00268976800101381.
- E. B. Wilson Jr., J. C. Decius, and P. C. Cross, Molecular Vibrations, McGraw–Hill, 1955; Dover reprint, 1980.
- R. N. Zare, Angular Momentum: Understanding Spatial Aspects in Chemistry and Physics, Wiley, 1988.