

Born–Oppenheimer Approximation as Scale Separation
The Born–Oppenheimer approximation uses the large disparity between nuclear and electronic masses to organize molecular quantum mechanics into fast electronic and slow nuclear problems. At each nuclear configuration, one first diagonalizes an electronic Hamiltonian. Its eigenvalues become potential-energy surfaces for the nuclear motion; derivatives of its eigenvectors couple those surfaces.
The slogan “electrons adjust instantaneously to fixed nuclei” captures the leading picture, but it hides the approximation’s actual content. The nuclei are not made classical, “fixed nuclei” describes only an auxiliary electronic eigenproblem, and a small mass ratio does not by itself guarantee single-surface dynamics. The decisive questions are whether the relevant electronic subspace remains spectrally isolated and whether nuclear motion can resolve the changing electronic eigenvectors.
Required background. Scale separation and error estimates explain the expansion; the adiabatic theorem introduces isolated spectral subspaces. The molecular Hamiltonian supplies the Coulomb model. Applications to spectra and structures are developed in Born–Oppenheimer molecular physics; nonadiabatic coupling treats motion on several surfaces.
Electronic Channels and Nuclear Motion
Section titled “Electronic Channels and Nuclear Motion”The construction is a sequence, not a single product ansatz:
The channel expansion on the last line is exact when the electronic basis is complete, with continuum integrals as needed in addition to the discrete sum. The approximation begins when one truncates the electronic channels or neglects terms generated when the nuclear kinetic energy differentiates the -dependent electronic states.
For an isolated channel , the leading one-surface equation is
This simple equation is powerful because a many-electron problem has been compressed into a scalar surface . Its reliability must be established from the omitted derivative couplings.
Heavy and Light Degrees of Freedom
Section titled “Heavy and Light Degrees of Freedom”Let denote electronic coordinates and nuclear coordinates. Before removing the overall center of mass, the nonrelativistic Coulomb Hamiltonian is
After separating center-of-mass motion, write the internal Hamiltonian schematically as
Here acts on electronic variables while the nuclear positions label the operator. Throughout this page, includes the internuclear repulsion, so its eigenvalues are molecular potential-energy surfaces. Another common convention leaves that scalar term outside ; either convention is valid if used consistently. Exact internal coordinates can also generate reduced-mass and mass-polarization terms. The displayed channel derivation below uses a diagonal nuclear Laplacian, as in the laboratory-coordinate model above or an appropriate slow–fast model in Jacobi coordinates. Exact internal kinetic cross terms must be included if present; they cannot be discarded while calling the internal-coordinate equations exact.
Which small parameter?
Section titled “Which small parameter?”For a representative nuclear mass , the fundamental dimensionless ratio is
There is no contradiction among the different powers of used in the literature. They organize different scalings:
- In atomic units, the nuclear kinetic operator carries a coefficient of order .
- A semiclassical slow-coordinate Hamiltonian is often written with , so that is proportional to .
- Near a smooth nondegenerate molecular minimum, the original Born–Oppenheimer ordering uses . Nuclear displacements are then of order , vibrational energies of order , and rotational energies typically of order .
Thus , , and are bookkeeping parameters adapted to different questions. A statement such as “the Born–Oppenheimer error is of order ” is incomplete unless it specifies the states, energy window, time scale, spectral-gap assumptions, and norm in which the error is measured. The general discipline is the same as in Small Parameters and Error Estimates: identify both the nominal small parameter and every denominator that can defeat it.
Why heavy particles are slow
Section titled “Why heavy particles are slow”For comparable kinetic energies, momentum scales as but velocity scales as . Heavy nuclei therefore move more slowly through configuration space than electrons. The electronic state can often track this motion, provided the electronic gap remains large enough.
Mass separation is only the beginning of the argument. Near an electronic degeneracy, eigenvectors may rotate sharply with , and even slow nuclei can generate substantial channel mixing. Conversely, a symmetry can force a coupling matrix element to vanish even at an energy crossing. The relevant comparison always combines nuclear motion, electronic gaps, and matrix elements.
Fixed-Configuration Electronic Problem
Section titled “Fixed-Configuration Electronic Problem”For each admissible nuclear geometry , solve
with electronic inner products
The subscript emphasizes that electronic variables are integrated while is held as a parameter. Antisymmetry, spin labels, and molecular symmetry should be imposed in this electronic Hilbert space. For a nondegenerate isolated eigenvalue, can be chosen smoothly in a local region. Near degeneracies, it is generally safer to retain a multiplet and work with its spectral projector rather than insist on one eigenvector.
The phrase clamped nuclei refers only to this parameter-dependent eigenproblem. Once and have been found, the nuclear coordinates return as quantum variables in the coupled molecular equation.
Exact Electronic-Channel Equations
Section titled “Exact Electronic-Channel Equations”Expand the molecular wavefunction in the fixed-configuration electronic basis:
Define the first- and second-derivative couplings
Applying the nuclear Laplacian to one channel gives
Projecting the full Schrödinger equation onto yields
where
These are the Born–Huang channel equations. No single-surface approximation has yet been made. They show exactly where electronic transitions enter: nuclear differentiation of an -dependent electronic basis produces off-diagonal operators.
For a complete smooth electronic basis,
The matrix behaves as a connection under an -dependent change of electronic basis. Individual derivative couplings are therefore basis dependent, whereas the full coupled-channel operator and its predictions are basis covariant. Truncating channels without transforming all terms consistently can spoil Hermiticity or create gauge-dependent results.
Potential-Energy Surfaces
Section titled “Potential-Energy Surfaces”Each electronic eigenvalue defines a potential-energy surface over nuclear configuration space. On one surface:
- minima identify candidate equilibrium geometries;
- gradients determine forces;
- Hessians near a minimum determine harmonic restoring forces;
- barriers and saddles organize tunneling and reaction pathways;
- asymptotic regions organize dissociation channels.
For an exact normalized nondegenerate electronic eigenstate, the Hellmann–Feynman relation gives
Approximate electronic wavefunctions or geometry-dependent finite bases can add response terms, often called Pulay terms. The formal identity should not be used to discard those numerical corrections.
A potential-energy surface is an effective object, not the exact molecular energy spectrum with the nuclei literally frozen forever. Nuclear zero-point motion, tunneling, rotation, geometric terms, and coupling to other electronic surfaces still have to be included in the nuclear problem. Molecular Quantum Mechanics connects these steps to bonding, normal modes, and spectroscopy.
From Exact Channels to One Surface
Section titled “From Exact Channels to One Surface”Suppose channel is separated from the other relevant electronic channels over the region explored by the nuclear state. The leading Born–Oppenheimer approximation keeps and drops the off-diagonal couplings to . Several related approximations should be distinguished:
| Level | Retained information | What is omitted |
|---|---|---|
| Clamped-configuration electronic problem | and at each | Nuclear dynamics |
| Leading one-surface Born–Oppenheimer equation | All derivative couplings | |
| Diagonal adiabatic correction | Diagonal scalar and geometric terms | Transitions to other surfaces |
| Multisurface Born–Huang treatment | A selected matrix of surfaces and derivative couplings | Electronic channels outside the retained set |
| Full molecular problem | All electronic and nuclear degrees of freedom | Nothing at the nonrelativistic model level |
In a local real gauge for a nondegenerate state, one may set . The diagonal second-derivative term then gives the positive diagonal Born–Oppenheimer correction
when the displayed electronic basis is complete. This correction is small when the mass-weighted eigenvector gradients are small on the energy scale of interest; a gap alone does not bound it. It often grows near a degeneracy. With , projection onto one channel gives the gauge-covariant operator
This diagonal projection is not automatically the full second-order dynamical approximation: virtual transitions to other channels also generate momentum-dependent corrections when states explore kinetic energies of order one in the semiclassical scaling (Panati, Spohn, and Teufel, 2007).
The leading approximation is therefore not merely the factorization
It is the claim that the omitted action of on has a controlled effect on the states and observables of interest.
Nonadiabatic Couplings
Section titled “Nonadiabatic Couplings”Differentiate the electronic eigenvalue equation with respect to and project onto a distinct state . One obtains
This off-diagonal Hellmann–Feynman identity makes the danger visible: derivative couplings are enhanced by small electronic gaps. It also shows why a gap alone is not the whole story; the numerator can vanish because of symmetry or selection rules.
For a semiclassical nuclear component with momentum and velocity , the first-derivative channel coupling has the energy scale
A useful local diagnostic is therefore
Using the derivative-coupling identity shows the familiar gap-squared structure:
One should also compare the second-derivative term with the gap, for example through
These are diagnostics in a specified smooth electronic gauge, not universal error bounds. In particular changes under coordinate-dependent phases; its magnitude alone is not gauge invariant. The complete combination of derivative terms acting on the transformed nuclear amplitudes is the physical object. A quantum nuclear wavepacket samples a region of phase space, so the relevant estimate must cover its support rather than one classical trajectory. Long evolution can accumulate small local errors, and threshold scattering or resonances can amplify corrections to particular observables. Rigorous versions use almost-invariant electronic subspaces, dressed projectors, and norms adapted to a specified energy and time window.
Avoided Crossing Example
Section titled “Avoided Crossing Example”Consider one nuclear coordinate and a two-state electronic Hamiltonian
The adiabatic surfaces are
with minimum gap at . In a smooth real gauge, the magnitude of the derivative coupling is
It is largest exactly where the surfaces approach most closely. Combining it with the energy gap gives
so at the center . Slow motion helps, but narrowing the avoided crossing defeats the one-surface approximation quadratically in .
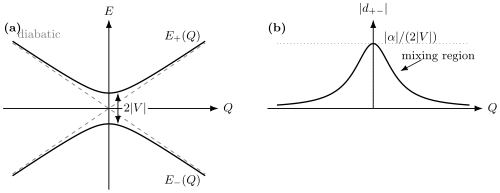
The avoided crossing has a minimum gap between the adiabatic energies at . The derivative coupling peaks in the same narrow region, so a nuclear packet can change electronic character even though its coordinate motion is slow.
This model isolates the local mechanism behind Landau–Zener transitions, but the two descriptions answer different questions. The Born–Oppenheimer channel equations describe how nuclear kinetic energy couples coordinate-dependent electronic states. A Landau–Zener formula additionally assumes a prescribed passage, usually with a locally linear time dependence, and estimates a transition probability. It should not replace a check of the molecular wavepacket, the dimensionality of the crossing region, or the validity of the two-state truncation.
Where the Approximation Fails
Section titled “Where the Approximation Fails”Single-surface dynamics becomes suspect in several recurring situations:
- Avoided crossings: a small gap produces sharply varying electronic eigenvectors and enhanced derivative couplings.
- Conical intersections: two surfaces become degenerate in a multidimensional nuclear space, so one isolated electronic line bundle no longer describes the crossing region.
- Fast or energetic nuclei: collisions, photodissociation, and highly excited vibrations can increase the momentum-dependent coupling.
- Light nuclei: hydrogen transfer and proton-coupled electron transfer can make nuclear delocalization and nonadiabatic effects especially important.
- Broad wavepackets: a packet centered in a benign region may still overlap a small-gap region through its tails.
- Ionization or dense electronic spectra: separation from electronic continua or nearby Rydberg manifolds can fail.
- Strong time-dependent driving: the relevant quasienergy or dressed-state gaps can differ from field-free electronic gaps.
An exact degeneracy does not automatically imply a transition between every pair of states. Symmetry may block the matrix element, and a degenerate multiplet can sometimes be retained as a whole while remaining isolated from all other states. In that case the correct effective theory is matrix valued rather than a scalar one-surface theory.
Berry Phase Preview
Section titled “Berry Phase Preview”For an isolated electronic state, the diagonal derivative coupling defines a local Berry connection
If the nuclei traverse a closed loop , the electronic state can acquire the holonomy
On a simply connected patch one can set only when its Berry curvature vanishes, as for a smooth real nondegenerate eigenstate. For a general complex Hamiltonian with nonzero curvature, even a local patch need not admit that gauge. Globally, a loop around a conical intersection may obstruct one single-valued choice and produce a nontrivial phase. That phase changes nuclear boundary conditions and interference even when the trajectory remains on one adiabatic surface.
The canonical derivation of the covariant nuclear momentum, gauge transformations, Born–Huang scalar term, and conical-intersection holonomy is on Born–Oppenheimer Berry Phase. The present page needs only the methodological warning: “neglect transitions” does not imply “neglect geometry.”
A Prescribed Nuclear Trajectory
Section titled “A Prescribed Nuclear Trajectory”Treating a nuclear trajectory as prescribed converts electronic following into a time-dependent adiabatic problem, with
That is useful for intuition and mixed quantum–classical schemes. The full Born–Oppenheimer problem is richer because is itself an operator and a quantum coordinate. Nuclear branching, tunneling, interference, and geometric phase cannot in general be represented by one prescribed path.
Exercises
Section titled “Exercises”1. Why the fourth root appears
Section titled “1. Why the fourth root appears”In atomic units, consider one nuclear normal coordinate near a minimum:
where is of order unity and . Determine the scaling of the oscillator frequency, ground-state width, and zero-point energy with . Express the result using .
Solution
The effective mass is , so
The ground-state width scales as
up to factors involving . The zero-point energy scales as
With , the displacement is of order and the vibrational energy is of order . This is why the original molecular expansion naturally uses a fourth-root parameter even though the nuclear kinetic prefactor is .
2. Gauge change in one electronic channel
Section titled “2. Gauge change in one electronic channel”Let
How must transform so that the molecular product is unchanged? Find the transformation of .
Solution
The product is invariant if
The transformed connection is
Consequently, only gauge-covariant combinations in the nuclear Hamiltonian and closed-loop holonomies are physical. A local gauge with need not extend globally around a degeneracy.
3. Does zero nuclear velocity remove all corrections?
Section titled “3. Does zero nuclear velocity remove all corrections?”At the center of the avoided crossing, let a nuclear wavefunction vary so slowly that its first derivative can be neglected. Find the diagonal scalar correction for either electronic state. Does the vanishing first-derivative contribution justify using only ?
Solution
At , , so
The scalar correction remains and grows as the gap closes. Comparing it with gives . Thus a velocity-based diagnostic alone can miss an important correction; near the single-channel projection itself becomes unsuitable.
References
Section titled “References”- M. Born and R. Oppenheimer, “Zur Quantentheorie der Molekeln,” Annalen der Physik 389, 457–484 (1927), doi:10.1002/andp.19273892002.
- M. Born and K. Huang, Dynamical Theory of Crystal Lattices, Clarendon Press (1954).
- G. A. Hagedorn, “A time dependent Born–Oppenheimer approximation,” Communications in Mathematical Physics 77, 1–19 (1980).
- T. Jecko, “On the mathematical treatment of the Born–Oppenheimer approximation,” Journal of Mathematical Physics 55, 053504 (2014), doi:10.1063/1.4870855.
- C. A. Mead and D. G. Truhlar, “On the determination of Born–Oppenheimer nuclear motion wave functions including complications due to conical intersections and identical nuclei,” The Journal of Chemical Physics 70, 2284–2296 (1979), doi:10.1063/1.437734.
- G. Panati, H. Spohn, and S. Teufel, “The time-dependent Born–Oppenheimer approximation,” ESAIM: Mathematical Modelling and Numerical Analysis 41, 297–314 (2007), doi:10.1051/m2an:2007023.
- H. Spohn and S. Teufel, “Adiabatic decoupling and time-dependent Born–Oppenheimer theory,” Communications in Mathematical Physics 224, 113–132 (2001).
- B. T. Sutcliffe and R. G. Woolley, “On the quantum theory of molecules,” The Journal of Chemical Physics 137, 22A544 (2012), doi:10.1063/1.4755287.
- D. R. Yarkony, “Diabolical conical intersections,” Reviews of Modern Physics 68, 985–1013 (1996), doi:10.1103/RevModPhys.68.985.