

Exchange Interactions
A magnetic exchange interaction is the dependence of a system’s low-energy electronic energy on the relative quantum state of magnetic moments. It is not an additional fundamental force placed beside electromagnetism. Rather, fermionic antisymmetry changes the allowed many-electron states, while Coulomb matrix elements, orbital overlap, hopping, charge excitation energies, itinerant response, and spin–orbit coupling determine whether those states have different energies.
The result is often summarized by a spin Hamiltonian,
but the number is the end of a reduction, not the beginning of an explanation. Different microscopic pathways can produce the same apparent bilinear coupling. Several pathways can coexist on one bond, and some mechanisms do not reduce accurately to a field-independent pairwise at all.
The central question is which electronic process couples the moments. Direct exchange, superexchange, double exchange, and RKKY coupling arise in different limits, with different signs and energy scales. Exchange and Correlation derives atomic Coulomb integrals; the Heisenberg Model develops the spin model that often results.
Required background. Spin and Spatial Wavefunctions explains the required fermionic symmetry, while the Hubbard Model supplies hopping and charge-excitation scales used in virtual exchange.
Helpful background. Singlet and Triplet States fixes coupled-spin basis conventions, while the Schrieffer–Wolff Transformation supplies a controlled low-energy projection.
Exchange Symmetry Is Not an Extra Force
Section titled “Exchange Symmetry Is Not an Extra Force”For two identical electrons, the total state must be antisymmetric under particle exchange. A spin singlet is antisymmetric in spin and must therefore be paired with a symmetric spatial state. A spin triplet is symmetric in spin and must be paired with an antisymmetric spatial state.
Let and be orthonormal spatial orbitals. The normalized exchange eigenstates are
For a spin-independent two-body interaction , define the direct and exchange matrix elements
Their interaction energies are
For electrons, accompanies the spin singlet and the triplet. The interaction contribution to the singlet–triplet splitting is therefore
For the positive Coulomb kernel and fixed orbitals, , so this contribution favors the triplet or leaves it degenerate. That conclusion is narrower than the slogan “Pauli exclusion causes ferromagnetism.” The full energy also contains kinetic, one-body, orthogonalization, hybridization, and correlation contributions. Those can reverse the net splitting.
Most importantly, antisymmetry by itself is kinematic. If the relevant orbitals have disjoint support and no process connects them, and the hopping matrix element vanish. The spin states can then remain degenerate even though the total wavefunction is still exactly antisymmetric. An exchange splitting requires state-dependent Hamiltonian matrix elements.
Exchange signs, spin units, and bond counting
Section titled “Exchange signs, spin units, and bond counting”Throughout this page, is a dimensionless spin operator:
The isotropic pair Hamiltonian is
With this convention:
- favors antiparallel correlations and is called antiferromagnetic;
- favors parallel correlations and is called ferromagnetic;
- has units of energy.
For two spin- moments,
The singlet and triplet energies are
so
This dimer identity is the simplest operational definition of the sign.
The permutation operator on two spin- states is
It has eigenvalue on the singlet and on the triplet. This identity explains why a two-state exchange splitting can be rewritten as a Heisenberg coupling, but it does not identify the microscopic mechanism that generated the splitting.
Why published signs disagree
Section titled “Why published signs disagree”Other common conventions include
and Hamiltonians written using unit vectors rather than spin operators. Some sums count every bond twice. Consequently, a quoted positive “exchange constant” may mean ferromagnetic coupling in one paper and antiferromagnetic coupling in another.
For example, changing to , with unit vectors , gives in a classical fixed-length description. An ordered-pair sum instead of one unordered-bond sum adds another factor of two. Comparing the Hamiltonians resolves the sign and normalization before any numerical values are compared.
Four Microscopic Pathways
Section titled “Four Microscopic Pathways”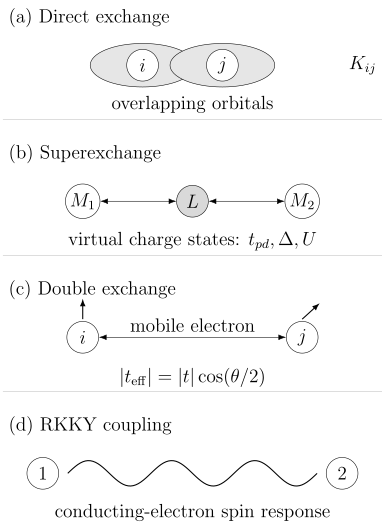
Four routes by which electronic motion and interaction couple magnetic moments. The superexchange arrows indicate virtual charge transfer; their direction depends on the occupied orbitals. The double-exchange carrier remains mobile. In RKKY coupling the intervening electron gas responds to a local spin. The diagram identifies processes, not universal signs for every material.
These mechanisms are not mutually exclusive labels for materials. A transition-metal oxide can have antiferromagnetic superexchange, ferromagnetic Hund coupling, weaker direct overlap, and longer-range itinerant exchange simultaneously. The effective is the low-energy sum after the active orbitals and charge fluctuations have been specified.
Direct Exchange
Section titled “Direct Exchange”Direct exchange couples magnetic orbitals that overlap without requiring a distinct ligand or conduction sea as mediator. At the two-orbital level, its characteristic Coulomb contribution is the exchange integral
It depends on the transition density , so it decreases rapidly as localized orbitals separate. Orbital orientation, radial extent, screening, covalency, and orthogonalization all matter.
For fixed orthonormal orbitals, the bare Coulomb exchange term lowers the triplet relative to the singlet and maps to a ferromagnetic contribution,
This equation should not be promoted into a universal sign theorem for an interatomic bond. Once orbital relaxation and hopping are allowed, virtual charge fluctuations often add an antiferromagnetic term. What a fitted model calls “nearest-neighbor exchange” may be the sum of both.
Intra-atomic Hund exchange
Section titled “Intra-atomic Hund exchange”Different orbitals on the same ion also have direct Coulomb exchange. A common local convention writes
This favors a high-spin atomic multiplet. Hund exchange is crucial in both superexchange and double exchange, but it is not identical to an intersite Heisenberg coupling. Crystal-field splitting, pair hopping, and the full multiorbital Coulomb tensor are needed for quantitative work.
Superexchange
Section titled “Superexchange”Superexchange is an indirect coupling generated by virtual hopping through high-energy charge configurations. It is especially important in magnetic insulators, where real dc charge transport is suppressed but virtual charge transfer remains allowed.
Two-site Hubbard derivation
Section titled “Two-site Hubbard derivation”The cleanest example is the half-filled, one-orbital two-site Hubbard model. Choose the phase of the orbitals so that its single hopping amplitude is real:
At half filling and , the low-energy subspace has one electron on each site. A hop temporarily creates one empty site and one doubly occupied site, costing energy . A second hop returns to the low-energy sector.
To second order,
within the singly occupied subspace. The constant is chosen so that the triplet energy shift is zero. The singlet is lowered by , and therefore
This antiferromagnetic sign follows from Pauli blocking and virtual kinetic energy: the singlet can access the relevant doubly occupied intermediate state, whereas the triplet cannot in a one-orbital model.
The result is controlled only when charge excitations remain well separated:
Near a metal–insulator transition, higher orders, longer-range hopping, ring exchange, and residual charge fluctuations can become important.
Ligand-mediated superexchange
Section titled “Ligand-mediated superexchange”In a transition-metal–ligand–transition-metal path, virtual charge transfer between metal and ligand states connects the two magnetic sites. The allowed direction depends on occupancy: for a filled ligand shell, a first electron hop can leave a ligand hole, rather than add an electron to an already occupied spin orbital. If is a metal–ligand hopping and a charge-transfer energy, a schematic downfolding gives
and an antiferromagnetic scale of order
This scaling identifies the small parameters, not a complete material formula. Real denominators include metal and ligand Coulomb repulsions, crystal-field levels, Hund couplings, and multiple orbital paths.
Goodenough–Kanamori–Anderson guidance
Section titled “Goodenough–Kanamori–Anderson guidance”Orbital symmetry provides useful qualitative rules:
- a nearly path between half-filled orbitals that hybridize with the same ligand orbital commonly gives strong antiferromagnetic exchange;
- a near- geometry can suppress that shared-orbital path, allowing ligand or metal Hund coupling to favor ferromagnetic exchange;
- half-filled–empty and half-filled–full combinations can have signs different from the half-filled–half-filled case;
- distortions can change both hopping magnitude and the active orbital channel.
These are chemistry-aware limiting rules, not a lookup table based on bond angle alone. The valence, orbital occupancy, local axes, ligand states, and competing paths must all be specified.
Double Exchange
Section titled “Double Exchange”Double exchange couples mixed-valence local moments through the kinetic energy of a mobile carrier. A minimal form contains hopping and a strong ferromagnetic Hund coupling between the carrier spin and a core moment:
Here is a dimensionless core spin of length , and is the dimensionless carrier spin. Each unordered bond is counted once. For the following semiclassical result, treat the cores as fixed classical directions and take : the carrier spin then follows the local core. Anderson and Hasegawa also analyze quantum cores; their finite-spin problem is not exactly the same as two fixed directions. If neighboring core moments subtend an angle , the overlap of their aligned spinors has magnitude
The projected amplitude is . Its magnitude is therefore
The complex phase also matters around a loop; replacing every amplitude by its magnitude would discard the associated geometric phase. For a single bond that phase can be removed by a change of orbital phases.
Parallel moments maximize each projected hopping magnitude and can lower the occupied-band kinetic energy. Antiparallel moments suppress hopping in the ideal limit. This produces a ferromagnetic tendency tied to carrier motion.
The comparison highlights the different physical processes:
| Property | Superexchange | Double exchange |
|---|---|---|
| Charge process | Virtual excursion and return | Real carrier delocalization |
| Typical regime | Integer filling, charge localized | Mixed valence or partial filling |
| Energy scale | Often proportional to | Often proportional to occupied-band kinetic energy |
| Angular form | Bilinear at leading order | Can scale as and be non-Heisenberg |
The ferromagnetic tendency is not unconditional. At an empty or completely filled active band there may be no kinetic-energy gain, and antiferromagnetic superexchange between core moments can compete strongly. Lattice distortions, orbital order, polarons, and phase separation can also reshape the simple picture.
RKKY interaction through a conducting host
Section titled “RKKY interaction through a conducting host”Localized moments can interact even when their orbitals scarcely overlap. Each polarizes the conducting electrons, whose response is felt by the other. Use a lattice-normalized coupling
where both spins are dimensionless and has units of energy. Define the static host response to by . For a spin-isotropic reciprocal host, its quadratic free-energy change is . Substitution of gives, for distinct sites,
This minus sign follows from the stated physical response convention. It is not an independent choice once and the plus-sign pair Hamiltonian are fixed. Continuum contact couplings have different units because the electron spin is then a density; RKKY Interaction gives that normalization and the full range-function derivation.
For a clean three-dimensional isotropic free-electron host, the large-distance coupling oscillates as , so different separations can favor parallel or antiparallel moments. In a crystal, the full Fermi surface determines periods and directions. Disorder, temperature, dimensionality, and spin–orbit coupling change the range or tensor structure.
This is a weak-coupling result. Kondo Effect describes the competing screening of a local moment; Kondo Lattices develops the competition in a dense array of moments.
Exchange Beyond an Isotropic Scalar
Section titled “Exchange Beyond an Isotropic Scalar”Spin–orbit coupling and low bond symmetry allow a general bilinear tensor,
Decomposing into scalar, antisymmetric, and symmetric-traceless parts gives
Here:
- is isotropic exchange;
- is the Dzyaloshinskii–Moriya vector;
- is symmetric and traceless anisotropic exchange.
Reversing the bond orientation gives ; this compensates the sign change of .
An inversion center at the midpoint of a bond forbids for that bond. Broken inversion is necessary but not sufficient; the remaining point-group operations constrain its direction. Strong spin–orbit-coupled materials can also generate bond-dependent Ising, compass, or Kitaev-like terms. Spin–Orbit Coupling in Solids explains how local orbital physics and crystal symmetry feed these effective interactions.
Single-ion anisotropy, magnetic dipole interactions, and Zeeman coupling are not exchange, even though they coexist in the same spin Hamiltonian. Magnetic Anisotropy explains their orientation-dependent material consequences and their competition with anisotropic exchange.
What Sets Sign and Strength
Section titled “What Sets Sign and Strength”An exchange parameter is controlled by several linked scales:
Distance and orbital orientation
Section titled “Distance and orbital orientation”Direct overlap and hopping usually decrease rapidly with distance. Because superexchange is proportional to several hopping amplitudes, modest bond-length or bond-angle changes can strongly alter . Pressure, epitaxial strain, and structural transitions can therefore tune exchange.
Occupancy and charge gaps
Section titled “Occupancy and charge gaps”Virtual exchange denominators depend on onsite repulsion, charge-transfer energy, crystal-field splitting, and Hund coupling. Changing valence or gating a narrow band can move the system between superexchange-dominated and carrier-mediated regimes.
Fermi-surface geometry
Section titled “Fermi-surface geometry”RKKY and itinerant exchange inherit wavevectors and anisotropy from the susceptibility . Nesting, multiple pockets, and spin–orbit splitting can select noncollinear or long-period order.
Competing paths
Section titled “Competing paths”Two bonds of equal length need not have equal exchange if their ligand chemistry differs. Several paths can cancel, making a small net highly sensitive to structural details. A sign change is often evidence for competition, not for one mechanism changing its intrinsic nature.
Choice of low-energy model
Section titled “Choice of low-energy model”Downfolding into different orbital sets redistributes physics among hopping, onsite interactions, and exchange terms. A static pairwise extracted near one reference state can differ from a coupling fitted over large spin rotations or at another temperature. Exchange parameters are model coordinates, not basis-independent observables.
How Exchange Is Determined
Section titled “How Exchange Is Determined”No single measurement reads out every directly. Reliable inference combines complementary information.
Singlet–triplet spectroscopy
Section titled “Singlet–triplet spectroscopy”For an isolated spin- dimer in this convention,
Inelastic neutron scattering, electron-spin resonance, optical spectroscopy, or tunneling spectroscopy can resolve this gap. Extra anisotropy splits the triplet and must be included rather than absorbed blindly into one .
Magnon dispersion
Section titled “Magnon dispersion”In an ordered magnet, exchange controls the momentum dependence of spin-wave energies. Inelastic neutron scattering measures the dynamic structure factor , allowing multiple neighbor couplings to be fitted across the Brillouin zone. Magnons develops the quantization and spectral-weight framework.
A zone-center stiffness alone usually constrains only a weighted moment of the . Several exchange networks can share the same long-wavelength slope but differ at the zone boundary.
Thermodynamics
Section titled “Thermodynamics”Susceptibility, specific heat, and ordering temperature constrain exchange scales, but they are inverse problems. Frustration, dimensionality, anisotropy, disorder, and quantum fluctuations can suppress the ordering temperature far below the largest bond energy. The estimate is dimensional guidance, not an extraction formula.
Electronic-structure mapping
Section titled “Electronic-structure mapping”One can calculate total energies for several magnetic configurations and fit them to a spin Hamiltonian. For one isolated classical pair of equal spin length ,
In a lattice, bond multiplicities and all simultaneously changed pairs must be counted. The fitted result depends on the spin normalization, reference state, electronic-structure approximation, orbital projection, and chosen interaction terms.
Infinitesimal-rotation or magnetic-force-theorem methods extract the curvature of electronic energy around a reference state. Correlated methods can add frequency-dependent self-energies and vertex information. Agreement with measured magnon dispersions is a stronger test than agreement with one collinear energy difference.
Competing exchange paths and the limits of a spin model
Section titled “Competing exchange paths and the limits of a spin model”Consider a bond with and . Its one-orbital superexchange estimate is
A direct Coulomb exchange integral adds the leading contribution , giving . This additive estimate assumes small charge admixture and neglects mixed higher-order corrections. The small net value reflects cancellation between larger paths. Modest structural changes can therefore reverse it even when neither microscopic process changes its own sign.
A fixed-length Heisenberg model works when charge and orbital excitations are well separated and the angular energy is approximately bilinear. Compare several spin orientations or observables: one collinear energy difference cannot reveal a biquadratic interaction or changing moment length. The review by Szilva and collaborators explains how exchange extraction depends on the reference electronic state and the retained variables.
If moment amplitudes change during rotation, orbital states remain active, or charge fluctuations are strong, include those degrees of freedom. Multispin, anisotropic, retarded, or state-dependent interactions can then matter. A useful effective model specifies the energy and temperature range in which its parameters predict the observations.
Exercises
Section titled “Exercises”1. Spinor overlap in double exchange
Section titled “1. Spinor overlap in double exchange”Let
represent a spin aligned with a unit vector. Show that the magnitude of the overlap of two aligned spinors separated by angle is .
Solution
For pure spin- states with Bloch vectors and ,
Since ,
For , the magnitude is . Parallel core spins maximize hopping; antiparallel spins suppress it in the ideal strong-Hund limit.
2. The angular energy of one mobile electron
Section titled “2. The angular energy of one mobile electron”For two classical core directions separated by and one strongly Hund-aligned electron, the two-site hopping amplitude has magnitude , with . Find the electronic ground-state energy and its small-angle Heisenberg approximation . Can this approximation reproduce the whole angular curve?
Solution
The two eigenvalues of the projected hopping matrix are for . The one-electron energy is
Since has quadratic term , matching the curvature gives . Adjusting the constant to match then predicts , whereas the exact projected value is zero. A good small-angle fit does not establish a bilinear interaction over large rotations.
3. Why midpoint inversion forbids Dzyaloshinskii–Moriya exchange
Section titled “3. Why midpoint inversion forbids Dzyaloshinskii–Moriya exchange”Consider
Show why an inversion center at the bond midpoint requires .
Solution
Inversion about the midpoint exchanges sites and . Spin is an axial vector, so each spin direction is unchanged by spatial inversion, but the site labels swap:
The bond is mapped onto itself. Invariance would therefore require the same coefficient multiplying both a term and its negative. The only possibility is .
4. Energy-mapping normalization
Section titled “4. Energy-mapping normalization”A calculation uses two classical spins of length and the convention . It finds for an isolated pair. Determine and its magnetic sign.
Solution
For parallel and antiparallel classical vectors,
Thus
With ,
The negative sign is ferromagnetic in this convention. In a lattice calculation, every changed bond and any moment-length change would also need to be counted.
The next step depends on the question. Ferromagnetism and Antiferromagnetism connect exchange to order; Spin Waves and Magnons in Materials connects an exchange network to measured excitation spectra. For two controlled spins, Silicon Spin Qubits uses exchange to generate gates and discusses charge-noise and leakage limits.
References
Section titled “References”-
P. W. Anderson, “New Approach to the Theory of Superexchange Interactions,” Physical Review 115, 2–13 (1959), doi:10.1103/PhysRev.115.2.
-
P. W. Anderson and H. Hasegawa, “Considerations on Double Exchange,” Physical Review 100, 675–681 (1955), doi:10.1103/PhysRev.100.675.
-
J. B. Goodenough, “Theory of the Role of Covalence in the Perovskite-Type Manganites,” Physical Review 100, 564–573 (1955), doi:10.1103/PhysRev.100.564.
-
J. Kanamori, “Superexchange interaction and symmetry properties of electron orbitals,” Journal of Physics and Chemistry of Solids 10, 87–98 (1959), doi:10.1016/0022-3697(59)90061-7.
-
D. I. Khomskii, Transition Metal Compounds (Cambridge University Press, 2014), doi:10.1017/CBO9781139096782.
-
T. Moriya, “Anisotropic Superexchange Interaction and Weak Ferromagnetism,” Physical Review 120, 91–98 (1960), doi:10.1103/PhysRev.120.91.
-
M. A. Ruderman and C. Kittel, “Indirect Exchange Coupling of Nuclear Magnetic Moments by Conduction Electrons,” Physical Review 96, 99–102 (1954), doi:10.1103/PhysRev.96.99.
-
A. Szilva, Y. Kvashnin, E. A. Stepanov, L. Nordström, O. Eriksson, A. I. Lichtenstein, and M. I. Katsnelson, “Quantitative theory of magnetic interactions in solids,” Reviews of Modern Physics 95, 035004 (2023), doi:10.1103/RevModPhys.95.035004.