

Tight-Binding Models
A tight-binding model represents crystalline quantum states in a retained set of localized orbitals and replaces continuum motion by coherent Hamiltonian matrix elements between them. Translation symmetry then converts the real-space hopping matrices into a finite-dimensional Bloch Hamiltonian for each crystal momentum.
The method is not synonymous with nearest-neighbor hopping, one orbital per site, or nonoverlapping atomic orbitals. A tight-binding representation can contain many orbitals, long-range and complex hopping, spin–orbit coupling, nonorthogonal basis functions, and parameters downfolded from first-principles bands. Its accuracy is controlled by the chosen subspace and the terms retained after projection.
We construct the crystalline Bloch matrix from its localized orbitals, then compare simple chains and two-sublattice models. The many-body tight-binding introduction explains occupation and finite boundaries; flat bands examines the special case of a momentum-independent energy.
Required background. Bloch’s Theorem supplies the translation-sector decomposition, while Crystals and Lattices supplies primitive cells, motifs, orbital positions, and finite geometry.
Localized Orbital Basis
Section titled “Localized Orbital Basis”Choose localized one-particle states
where labels a Bravais-lattice cell and labels an orbital, sublattice, layer, or other retained mode within the cell. Translation covariance means
For an orthonormal basis,
The corresponding real-space wavefunctions are
They may be idealized atomic orbitals, symmetry-adapted combinations, molecular orbitals, numerically constructed Wannier functions, or phenomenological modes. The word “orbital” describes a basis state, not necessarily an isolated-atom eigenfunction.
Projection from a continuum Hamiltonian
Section titled “Projection from a continuum Hamiltonian”Given a one-particle operator , its projected matrix elements are
Translation symmetry makes the matrix element depend only on the cell displacement:
If the retained orbitals span an invariant subspace and every matrix element is kept, this is an exact change of representation within that subspace. Approximation enters when remote states are excluded, energy dependence from downfolding is neglected, or long-range matrix elements are truncated.
Basis nonuniqueness
Section titled “Basis nonuniqueness”Localized orbitals are not observables. A unitary transformation among retained orbitals changes individual onsite energies, hopping amplitudes, and spatial spreads while preserving predictions when all operators are transformed consistently. Statements such as “the nearest-neighbor hopping is ” are meaningful only after the basis, gauge, orbital embedding, and fitting procedure are specified.
Real-Space Hamiltonian
Section titled “Real-Space Hamiltonian”Using the hopping orientation fixed in Conventions for Quantum Matter, write
multiplies the transfer from orbital to . Hermiticity requires
The block contains onsite energies and intracell hybridization. In a basis that diagonalizes that block,
but diagonal onsite form is a convenience rather than a requirement.
Hopping is coherent
Section titled “Hopping is coherent”A hopping amplitude is an off-diagonal Hamiltonian matrix element. It produces unitary superposition and interference; it is not a classical rate or a stochastic jump probability. The probability of transfer depends on the full time evolution, all available paths, detuning, and state preparation.
The conventional minus sign
Section titled “The conventional minus sign”For one orbital on a chain, it is common to write
with . In the general convention above,
Some sources instead call the signed matrix element itself . Always infer the sign from the written Hamiltonian before comparing dispersions.
Bloch Hamiltonian
Section titled “Bloch Hamiltonian”Use the cell-convention transform
Substitution gives
with
Hermiticity in real space implies
The cell convention also makes the matrix periodic:
At each , solve
With orthonormal spinless orbitals per primitive cell, there are bands and one-particle states in a finite periodic crystal. Physical spin doubles this count before spin splitting or additional constraints are considered.
Band operators
Section titled “Band operators”Choose normalized eigenvectors,
and define
Then
The diagonalization is exact for the specified quadratic model. Whether that model accurately represents a material is a separate question.
Orbital Embedding and Bloch Gauge
Section titled “Orbital Embedding and Bloch Gauge”The cell transform excludes the basis position . An orbital-position convention instead uses
Define
Then
and
The eigenvalues are identical, but the embedded matrix is generally sewn rather than strictly periodic:
Orbital embedding matters for position, polarization, optical matrix elements, and Berry geometry. Two Hamiltonian matrices that look different can represent the same model in different -dependent bases.
Nonorthogonal Orbitals
Section titled “Nonorthogonal Orbitals”Atomic-orbital bases often have a nontrivial overlap,
Its Bloch transform is
The band problem is then generalized:
For a linearly independent Bloch basis at each , is positive definite and eigenvectors obey . The real-space second-quantized formulas above assumed canonical operators in an orthonormal basis. Creation operators for the actual overlapping orbitals do not satisfy those canonical relations; orthogonalize before using that form of the many-body Hamiltonian. Replacing this by an ordinary eigenproblem without first orthogonalizing the basis changes the spectrum. Orthogonalization itself can lengthen the range of the effective hopping, so “nearest neighbor” is not basis invariant.
Hopping Range and Locality
Section titled “Hopping Range and Locality”For well-localized orbitals and a local or short-ranged one-particle operator, matrix elements often decay with the separation
This motivates truncation to onsite, nearest-neighbor, or a few neighbor shells. The decay can be anisotropic and orbital dependent, and downfolding can generate longer-range terms even when the original microscopic Hamiltonian was local.
A hopping cutoff should be justified by a convergence test. Compare full and truncated models over the entire target Brillouin zone, not only along a few high-symmetry lines.
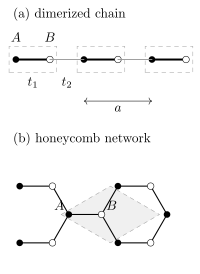
Two examples with two orbitals per primitive cell and therefore two bands before spin. In the dimerized chain, the dashed boxes choose cells and the bond strengths distinguish from . The honeycomb network has triangular translations and two sublattices; the shaded parallelogram is one primitive cell. The hopping graph and orbital positions supply different information and both enter physical matrix elements.
Worked Example: One-Dimensional Chain
Section titled “Worked Example: One-Dimensional Chain”For the Hamiltonian written above,
so
For the continuous bulk band and , the minimum is at , the maximum is at , and the bandwidth is
A finite periodic chain samples only its allowed momenta. For odd , is absent and the sampled bandwidth is , approaching as . The dispersion is broad when neighboring orbitals hybridize strongly and collapses to the atomic level as . The Tight-Binding Chain develops the finite spectra and boundary conditions.
Worked Example: SSH Chain
Section titled “Worked Example: SSH Chain”The Su–Schrieffer–Heeger chain has two orbitals per cell, intracell hopping , and intercell hopping . With real hoppings,
In the cell convention,
The bands are
For positive , the direct gap at is
The gap closes at , where the chosen two-site cell merely folds the uniform chain. Boundary zero modes in the minimal hopping model rely on chiral symmetry and the termination; their interpretation also requires a consistent unit cell. See the SSH model for the winding number and boundary calculation.
Worked Example: Square Lattice
Section titled “Worked Example: Square Lattice”For one orbital per rectangular cell and nearest-neighbor hoppings ,
If , the bandwidth is
For the isotropic square lattice,
the band runs from at to at the zone corner. The saddle points at and later produce a two-dimensional van Hove singularity in the density of states.
Longer-range diagonal hopping adds
changing particle–hole symmetry and the Fermi-surface shape without changing the one-orbital band count.
Worked Example: Honeycomb Lattice
Section titled “Worked Example: Honeycomb Lattice”The honeycomb structure is not a Bravais lattice. It has a triangular Bravais lattice and two sublattices . For nearest-neighbor hopping in a common cell convention,
where one convenient choice is
The bands are
They touch where
at inequivalent Brillouin-zone corners. Expanding near such a point gives a two-component Dirac Hamiltonian whose Pauli matrices act on sublattice pseudospin, not physical spin.
An onsite sublattice imbalance adds
and opens a gap in this minimal model. Graphene Dirac Model gives the compact low-energy Hamiltonian. Graphene and Dirac Materials develops the material interpretation, pseudospin and Berry diagnostics, magnetic spectrum, realistic corrections, and moiré bridge.
Multi-Orbital Models
Section titled “Multi-Orbital Models”With several orbitals per cell, is an Hermitian matrix. Its entries can include:
- crystal-field-split onsite energies;
- intracell hybridization between orbitals of compatible symmetry;
- direction-dependent intercell hopping;
- longer-range hopping;
- layer and sublattice couplings;
- spin-dependent onsite and hopping terms.
Orbital symmetry strongly constrains the angular dependence. Slater–Koster parameterizations organize two-center matrix elements such as , , , and by bond direction. Their tabulated form is a modeling convention, not a substitute for checking the retained basis and environment.
Symmetry covariance
Section titled “Symmetry covariance”If a unitary crystal symmetry maps to , a correctly constructed Bloch Hamiltonian obeys
with possible reciprocal sewing when is returned to the chosen zone. Symmetry can force degeneracies, forbid hybridization, or relate hopping parameters, but it does not determine their numerical values.
For an antiunitary symmetry, such as time reversal, the left side instead contains between the two unitary matrices. Symmetry of Bloch States explains how these relations constrain degeneracies and hybridization.
Spin–Orbit Coupling
Section titled “Spin–Orbit Coupling”With physical spin, each orbital carries a spinor and acts in orbital spin space. Atomic spin–orbit coupling takes the onsite form
Here and are dimensionless, so has units of energy. A nontrivial onsite orbital angular momentum requires a suitable orbital multiplet. Projection to a single nondegenerate real orbital can remove the first-order onsite term while virtual transitions generate spin-dependent hopping; this is not obtained by assigning an arbitrary orbital Pauli matrix to physical spin.
For a single orbital channel, any spin block can be expanded in the identity and Pauli matrices. In a time-reversal-compatible real orbital basis, a useful form with real and is
Hermiticity then requires
The odd spin-dependent term becomes momentum odd. These real-coefficient conditions are not a general parameterization for complex multi-orbital bases, where time reversal can also mix orbital components. The full spinful Bloch matrix must satisfy the appropriate time-reversal, point-group, and Hermiticity constraints. Spin–Orbit Coupling develops the angular-momentum calculation. Spin–Orbit Coupling in Solids develops crystal-field projection, Bloch-band symmetry constraints, and Rashba and Dresselhaus invariants; a material model must still specify its orbital and spatial symmetry representation.
Complex Hopping and Magnetic Phases
Section titled “Complex Hopping and Magnetic Phases”Local orbital rephasings,
move phases among bonds without changing gauge-invariant loop products. A magnetic vector potential can be incorporated approximately through the Peierls phase,
Only the accumulated phase around a closed loop is directly tied to magnetic flux. The path choice, orbital extent, and additional magnetic couplings must be controlled in precision work; see Peierls Phase Preview.
From Bands to Wannier Functions
Section titled “From Bands to Wannier Functions”Wannier functions turn an appropriate Bloch subspace into translated localized orbitals. Wannierization workflows explain how to choose and converge that subspace numerically. An isolated group with nonzero Chern number has no exponentially localized orthonormal Wannier basis related by lattice translations. A larger group with canceling total Chern number can remove that obstruction; demanding additional symmetries can impose further restrictions.
For a complete localized basis, the band-derived hopping matrix is the inverse Fourier transform of the Wannier-gauge Bloch matrix:
Within a fixed selected subspace, a unitary Bloch-frame change and complete Fourier transform are exact basis changes: they preserve the projector and the represented spectrum. Subspace selection or disentanglement, hopping truncation, parameter fitting, and errors in the reference electronic structure are distinct sources of approximation error and should be tested separately.
Relation to Hubbard Models
Section titled “Relation to Hubbard Models”The base tight-binding Hamiltonian is quadratic. A Hubbard model adds interactions in the same localized basis, for example
More complete projections generate interorbital repulsion, Hund exchange, pair hopping, and nonlocal density interactions. Their values depend on orbital localization and on what screening has already been included.
Adding does not merely “correct the band energies.” It changes the many-body problem and can invalidate an independent-particle description. Hubbard Model develops the interacting limits. Hubbard Physics in Materials explains screened interaction tensors, electron counts and the danger of counting the same interaction effect twice.
How much hopping can be neglected?
Section titled “How much hopping can be neglected?”Choose the energy range and physical observable first. An accurate dispersion near the Fermi energy may need fewer orbitals than an optical calculation involving higher bands. Fit or compute matrix elements in a declared orbital basis, and test Hermiticity, translation symmetry and any assumed point or time-reversal symmetry before interpreting the bands.
Suppose an orthonormal model omits hopping blocks . The operator norm gives a useful uniform bound,
Every energy-ordered eigenvalue then moves by at most . For example, removing a second-neighbor chain hopping changes the band by , whose maximum magnitude is exactly . Small individual matrix elements are insufficient if many omitted terms accumulate.
This energy bound does not by itself control eigenvectors near a degeneracy. Optical transitions, Berry curvature, polarization and interaction matrix elements require converged eigenvectors and consistently projected operators as well as energies. A gap larger than cannot close under this bounded hopping change, but a much smaller gap may disappear. Compare successive hopping ranges over the full zone, including regions away from the usual plotted path.
Exercises
Section titled “Exercises”Exercise 1: prove Hermiticity in momentum space
Section titled “Exercise 1: prove Hermiticity in momentum space”Use
to prove .
Solution
Take the conjugate transpose:
Relabel :
Exercise 2: square-lattice checks
Section titled “Exercise 2: square-lattice checks”For isotropic nearest-neighbor hopping on a square lattice,
find the energies at , , and . Identify the saddle point.
Solution
Direct substitution gives
Near , write
To quadratic order,
The curvatures have opposite signs, so is a saddle point.
Exercise 3: generalized eigenproblem
Section titled “Exercise 3: generalized eigenproblem”Suppose a nonorthogonal two-orbital basis has positive-definite overlap matrix . Show how to convert
into an ordinary Hermitian eigenproblem.
Solution
Positive definiteness gives a Hermitian square root and inverse . Set
Multiplying the generalized equation by gives
The transformed Hamiltonian is Hermitian because and are Hermitian:
This symmetric orthogonalization preserves the generalized eigenvalues but changes the real-space shape and range of the effective orbitals and hopping. For a concrete example, take
The symmetric and antisymmetric vectors have energies and . With and , these are and , rather than . Their coefficient vectors must be normalized with , not with the Euclidean norm.
Exercise 4: a time-reversal-invariant spin-dependent hop
Section titled “Exercise 4: a time-reversal-invariant spin-dependent hop”One spinful orbital per cell has hopping , with real and the Hermitian reverse hop. Find , check time reversal , and identify the momenta at which the spin splitting vanishes.
Solution
The two directed hops give
Time reversal changes both spin and momentum: . The spin-resolved energies are , paired between and . Their splitting vanishes at and , the time-reversal-invariant momenta of the continuous band. A finite odd ring does not sample the latter point. Time reversal alone does not force spin degeneracy at a generic momentum.
References
Section titled “References”- N. W. Ashcroft and N. D. Mermin, Solid State Physics (Holt, Rinehart and Winston, 1976), Chapters 10–12.
- A. H. Castro Neto, F. Guinea, N. M. R. Peres, K. S. Novoselov, and A. K. Geim, “The electronic properties of graphene,” Reviews of Modern Physics 81, 109–162 (2009), doi:10.1103/RevModPhys.81.109.
- N. Marzari, A. A. Mostofi, J. R. Yates, I. Souza, and D. Vanderbilt, “Maximally localized Wannier functions: Theory and applications,” Reviews of Modern Physics 84, 1419–1475 (2012), doi:10.1103/RevModPhys.84.1419.
- J. C. Slater and G. F. Koster, “Simplified LCAO Method for the Periodic Potential Problem,” Physical Review 94, 1498–1524 (1954), doi:10.1103/PhysRev.94.1498.
- W. P. Su, J. R. Schrieffer, and A. J. Heeger, “Solitons in Polyacetylene,” Physical Review Letters 42, 1698–1701 (1979), doi:10.1103/PhysRevLett.42.1698.