

Crystals and Lattices
A Bravais lattice is a discrete group of spatial translations. A crystal structure is the physical motif repeated by those translations. Keeping those statements separate prevents several persistent mistakes: lattice points are not automatically atoms, a honeycomb array is not a Bravais lattice, and a conventional crystallographic cell need not be primitive.
The distinction matters whenever we count electronic states, construct vibrational modes or interpret diffraction. We first describe translations and cells, then use square, triangular and honeycomb examples to separate the repeated geometry from its atoms and orbitals.
Helpful background. Conventions for Quantum Matter fixes the lattice and Fourier signs used later; Momentum Operator as Generator and Translation-Invariant Hamiltonians supply the abstract operator language. This page introduces the direct-space crystal concepts itself.
Discrete translations in a crystal
Section titled “Discrete translations in a crystal”A macroscopic crystal contains an enormous number of microscopic degrees of freedom. Exact repetition converts part of that complexity into symmetry. Instead of treating every spatial region independently, one identifies translations that leave the idealized system unchanged.
For a single-particle Hamiltonian
the relevant idealization is
The set is not merely a collection of visually repeated points. It is the translation group of the periodic structure. This group, rather than a preferred drawing of a unit cell, is the invariant object from which reciprocal space and Bloch theory follow.
There are several physically distinct ways such periodicity can arise:
- equilibrium ionic positions in a crystalline solid;
- a periodic mean field or effective potential for electrons;
- an externally imposed optical lattice for neutral atoms;
- a periodic network used directly as an effective lattice model;
- a superstructure generated by charge, spin, orbital, or structural order.
The same geometry can therefore organize very different microscopic systems. Conversely, two materials with the same Bravais lattice can have different motifs, orbitals, interactions, and phases.
Lattice as a Discrete Translation Group
Section titled “Lattice as a Discrete Translation Group”Let be linearly independent vectors in a -dimensional physical subspace. The lattice they generate is
Vector addition gives
so is an abelian group. Once a primitive basis is chosen, its abstract group structure is
The embedding into Euclidean space still matters. It supplies lengths, angles, orientation, and cell volume, none of which is contained in the abstract group alone.
Discrete means isolated
Section titled “Discrete means isolated”The word discrete adds an important condition: lattice points do not accumulate at finite separation. Equivalently, there is a neighborhood of the origin containing no lattice vector except . Linear independence of a finite primitive set guarantees this property for the integer span above.
A dense subgroup such as
is generated by integer combinations, but it is not a one-dimensional crystal lattice because its points can approach one another arbitrarily closely.
Translation operators
Section titled “Translation operators”With the active translation convention,
The lattice translations obey
An ideal lattice-periodic Hamiltonian satisfies
This is weaker than continuous translation invariance. Ordinary momentum need not be conserved in a periodic potential, even though all commute with . Momentum Operator as Generator develops the continuous group; Translation-Invariant Hamiltonians explains the operator statement in general.
Primitive Vectors Are Not Unique
Section titled “Primitive Vectors Are Not Unique”A set is a primitive basis of when every lattice vector has a unique expansion with integer coefficients. Put the primitive vectors into the columns of a matrix
Another set generates the same lattice precisely when
where is an integer matrix with
Such a matrix is unimodular. Its inverse also has integer entries, so each old primitive vector is an integer combination of the new ones and conversely.
This freedom is not cosmetic. For the square lattice, both
and
are primitive bases. They define differently shaped parallelograms but the same translation group.
By contrast,
do not form a primitive basis of the square lattice. The transformation has determinant , so the integer span of reaches only an index-two sublattice. A cell drawn from these vectors has twice the primitive area.
Cell measure
Section titled “Cell measure”The primitive cell measure is
In three dimensions this is the scalar triple product
Under ,
Every primitive basis therefore gives the same . An integer change with describes a cell containing primitive cells.
Cells and Fundamental Domains
Section titled “Cells and Fundamental Domains”For a chosen primitive basis, a convenient half-open primitive cell is
Its translates tile space without overlap except at boundaries:
where the half-open convention assigns each boundary point to one cell. A primitive cell is thus a fundamental domain of . It need not have a unique shape. Parallelepipeds, Wigner–Seitz cells, and many less symmetric regions can represent the same quotient.
The Wigner–Seitz cell about a chosen lattice point consists of all points at least as close to that lattice point as to every other lattice point. To construct it, draw the perpendicular-bisector hyperplane of each vector from the chosen point to a neighboring lattice point and keep the intersection of the half-spaces containing the chosen point. The resulting cell is primitive, displays the point symmetry of the lattice, and is independent of a chosen primitive-vector basis up to boundary conventions.
A conventional cell is chosen to display symmetry or match crystallographic standards. It may be primitive, but it need not be. Face-centered and body-centered conventional cubic cells are familiar examples whose centering makes the symmetry transparent while increasing the cell volume.
This produces a terminology warning:
In crystallographic tables, an unqualified “unit cell” commonly means the conventional cell specified by the reported cell parameters. Do not infer that it contains one primitive cell.
If a conventional cell has volume
then its multiplicity is . Counting corner, face, edge, and interior lattice points with fractional weights reproduces for familiar polyhedral cells, but the volume ratio is the coordinate-independent statement.
Bravais Lattice Versus Crystal Structure
Section titled “Bravais Lattice Versus Crystal Structure”A Bravais lattice says where translations take one cell into another. A crystal structure says what is repeated. Let label objects in a motif, with positions inside a chosen cell. Their ideal positions are
The motif may include:
- several atomic species;
- inequivalent sites or sublattices;
- molecular orientations;
- local magnetic moments;
- structural orbitals used in an effective model;
- occupancies or displacement parameters in a crystallographic refinement.
A continuous density assembled from localized motif functions can be written
Shifting by any and relabeling the sum gives
A lattice point marks a translation-equivalent reference position in each cell. An atom need not sit at a lattice point, and several atoms can belong to one lattice point’s motif.
Three meanings of basis
Section titled “Three meanings of basis”The word basis is overloaded:
- a primitive lattice basis is the set of vectors ;
- a structural basis or motif is the list repeated in every cell;
- a Hilbert-space basis is a set of orbitals, spin states, bands, or many-body states used to represent quantum operators.
These counts need not agree. One atom can support several relevant orbitals, and several crystallographic sites can be combined into fewer low-energy bands. Conventions for Quantum Matter fixes the notation used when these notions appear together.
Bravais Types and Crystal Systems
Section titled “Bravais Types and Crystal Systems”A Bravais type classifies lattices with the same characteristic metric symmetry. The exact vectors and lattice constants still vary continuously within a type.
In two dimensions there are five Bravais types:
| Bravais type | Conventional metric |
|---|---|
| oblique | arbitrary unequal lengths and generic angle |
| rectangular | perpendicular vectors with unequal lengths |
| centered rectangular | rectangular conventional cell with one centering translation |
| square | perpendicular vectors of equal length |
| hexagonal | equal lengths with an angle of or |
There are four two-dimensional lattice systems because primitive and centered rectangular lattices belong to the same rectangular system. This is why “five 2D Bravais types” and “four 2D lattice systems” are both correct.
In three dimensions there are fourteen Bravais types distributed among seven lattice systems: triclinic, monoclinic, orthorhombic, tetragonal, rhombohedral, hexagonal, and cubic. These types classify translation lattices, not all crystal structures. Adding a motif and the operations that act on it leads to point groups and space groups.
The Crystalline Symmetry Preview introduces that larger symmetry structure. International crystallographic notation is more precise than the informal labels often used in condensed-matter calculations, so material data should be read with its stated conventional cell and space group.
Five Worked Geometries
Section titled “Five Worked Geometries”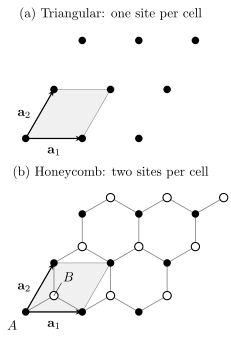
The same triangular translations generate two different structures. The shaded primitive cell has one site in panel (a), and an motif in panel (b), with boundary sites counted once across neighboring cells. Translating between the black and open honeycomb sites does not preserve the whole structure. Gray bonds guide the eye; translations define the lattice.
One-dimensional chain
Section titled “One-dimensional chain”Choose
Then
A monatomic chain has one motif site, . Each site has two nearest neighbors in the infinite chain, so its coordination number is . If alternating bonds have different strengths, the sites may remain spaced by , but the full Hamiltonian repeats only after . Its primitive translation cell then has two sites. The geometric spacing of sites alone does not determine the translation symmetry of the Hamiltonian.
Square lattice
Section titled “Square lattice”Take
The primitive area and monatomic nearest-neighbor coordination are
The square Bravais lattice has fourfold rotations about a lattice point. A motif can reduce that point symmetry even while preserving every translation.
Triangular lattice
Section titled “Triangular lattice”A standard primitive basis is
Its primitive area is
For one site per cell, the six vectors
form the nearest-neighbor shell, so . In two-dimensional crystallography this Bravais type is called hexagonal; “triangular lattice” describes the visible point net and avoids confusion with the non-Bravais honeycomb structure.
Honeycomb structure
Section titled “Honeycomb structure”Use the same triangular Bravais lattice and attach two sites:
Here is the length of a primitive Bravais translation. The nearest-neighbor bond length is
The full site set is
Translations in preserve the and sublattices separately. A displacement from to is not a symmetry translation: translating twice by produces the coset , which is neither nor . Thus the honeycomb structure is a triangular Bravais lattice with a two-site motif, not a sixth two-dimensional Bravais type.
Graphene realizes this geometry with carbon atoms, while hexagonal boron nitride places different species on the two motif sites. The same positions therefore do not imply the same sublattice symmetry or electronic Hamiltonian.
Simple-cubic lattice
Section titled “Simple-cubic lattice”Choose
Then
for a monatomic nearest-neighbor geometry. Simple cubic, body-centered cubic, and face-centered cubic are three different Bravais types. The latter two are often drawn using cubic conventional cells that contain two and four primitive cells, respectively.
Physical Crystal Examples
Section titled “Physical Crystal Examples”The same lattice-plus-motif language handles familiar structures:
| Structure | Translation lattice | Motif statement |
|---|---|---|
| monatomic face-centered cubic metal | face-centered cubic | one atom associated with each primitive cell |
| body-centered cubic elemental phase | body-centered cubic | one atom per primitive cell |
| graphene | two-dimensional triangular | two carbon sites, and |
| hexagonal boron nitride layer | two-dimensional triangular | boron and nitrogen occupy different motif sites |
| diamond structure | face-centered cubic | two identical atoms displaced along a body diagonal |
| rock-salt structure | face-centered cubic | two interpenetrating ionic sublattices |
Actual materials can change structure with temperature, pressure, composition, strain, or substrate. A structure label should therefore be understood as a statement about a specified phase and set of conditions.
Periodicity is not the same as crystallinity
Section titled “Periodicity is not the same as crystallinity”An optical lattice can impose
without the atoms spontaneously forming a rigid solid. Optical Lattices explains this controlled realization.
Conversely, a self-organized crystal is a phase of matter. In an ideal infinite system, crystalline order breaks continuous translations down to a discrete subgroup. The order is diagnosed through correlation functions and Bragg scaling, not merely by drawing particles near regular sites. See Long-Range Order for the many-body criterion.
Neighbor Shells, Bonds, and Models
Section titled “Neighbor Shells, Bonds, and Models”The lattice determines displacement vectors between translation-equivalent cells. It does not by itself determine which sites are chemically bonded, which orbitals overlap appreciably, or which couplings belong in an effective Hamiltonian.
For motif sites and , a relative displacement has the form
Neighbor shells group these vectors by length, possibly refined by point-group symmetry. A tight-binding representation can then assign amplitudes
Translation symmetry makes the amplitude depend on the cell difference rather than on two absolute cell labels. It does not require all geometrically equal distances to carry the same amplitude if the orbitals, species, magnetic order, or environment differ.
Coordination number is therefore model-dependent unless the neighbor criterion is stated. The values quoted for the chain, square, triangular, honeycomb, and simple-cubic examples refer to the nearest sites of the ideal monatomic or two-sublattice geometry.
Finite Samples and Boundary Conditions
Section titled “Finite Samples and Boundary Conditions”An infinite lattice is a mathematical limit. A finite supercell with primitive translations along direction contains
primitive cells. With structural sites and retained orbitals per site, a simple localized one-particle basis has
states before physical internal multiplicities such as spin are included. Nambu doubling or a gauge-redundant representation does not add physical one-particle states.
Periodic boundary conditions identify
The finite translation group becomes
This restores exact translation symmetry on a finite computational domain and leads to a discrete momentum mesh. It removes physical surfaces, however. Open boundaries retain edges and surface coordination changes but generally break translations near the boundary.
Boundary Conditions on Lattices develops finite many-body geometries, while Periodic Boundary Conditions gives the one-dimensional wave-mechanics foundation.
Thermodynamic-limit caution
Section titled “Thermodynamic-limit caution”For short-range bulk observables away from criticality, boundary contributions often scale like surface area while bulk contributions scale like volume. That heuristic does not justify discarding boundaries universally. Gapless modes, long-range interactions, topological boundary states, electrostatics, and transport contacts can keep boundary physics observable even in large samples.
Fractional Coordinates and Computational Geometry
Section titled “Fractional Coordinates and Computational Geometry”The lattice matrix converts fractional coordinates to Cartesian coordinates:
A motif position is commonly stored as
The direct-space metric in fractional coordinates is
so
This formula matters for nonorthogonal cells. Componentwise wrapping of fractional differences into does not always find the nearest periodic image in a strongly skewed cell. Robust neighbor searches compare an adequate set of translated images or use a lattice-reduction algorithm.
For example, take , and a fractional separation . Its Cartesian length is about , even though both fractional components already lie in the usual wrapped interval. Subtracting gives , whose length is only . Fractional proximity and physical proximity differ in a skew cell.
When reading coordinates from a file, first identify the vector order, units, origin and whether the coordinates are fractional or Cartesian. These choices determine the physical structure represented by the numbers.
Changing the primitive basis by also changes fractional coordinates:
Cartesian positions and physical observables remain unchanged when the motif and all tensor components are transformed consistently.
From Ionic Positions to a Periodic Hamiltonian
Section titled “From Ionic Positions to a Periodic Hamiltonian”In an electronic-structure problem, nuclei are not literally fixed classical points. The common starting point is a scale-separated description in which equilibrium nuclear positions define a reference structure and electronic motion is solved conditionally on nuclear coordinates. Born–Oppenheimer Scale Separation owns that approximation.
Write an equilibrium nuclear position as
The actual operator or instantaneous coordinate is more accurately represented by
where is a displacement. Setting every displacement to zero defines the ideal static lattice. Keeping small collective displacements leads to phonons; large, anharmonic, or topological distortions require more.
The periodic ionic reference can generate an electronic potential
but an effective one-electron Hamiltonian may also contain self-consistent fields, spin-orbit coupling, magnetic order, or nonlocal terms. The correct symmetry is the symmetry of the full Hamiltonian, not of one term considered in isolation.
The band-structure workflow uses this crystal and magnetic structure as the input to an electronic calculation.
What Diffraction Measures
Section titled “What Diffraction Measures”A periodic density admits a Fourier series over a discrete set of reciprocal vectors:
The next article derives which are allowed from the condition
For a motif, the amplitude contains a basis-dependent factor of the form
Peak positions primarily reveal the translation lattice and its metric. Peak intensities, systematic absences, and probe dependence contain information about the motif, species, form factors, displacements, and correlations. A lattice cannot generally be reconstructed from a photograph of a few peaks without indexing choices and structural modeling.
An infinite perfect lattice gives ideal reciprocal-space delta peaks. Finite size, thermal motion, defects, domains, strain, and instrumental resolution broaden or attenuate them. Structure Factors develops the correlation-function observable, and Electron Diffraction gives an experimental route to crystalline periodicity.
What the Ideal Lattice Approximation Ignores
Section titled “What the Ideal Lattice Approximation Ignores”The ideal lattice is a reference configuration, not a complete material model.
| Omitted feature | What changes physically |
|---|---|
| zero-point and thermal motion | atomic positions fluctuate; Bragg intensity is reduced and phonons appear |
| anharmonicity | normal modes interact; thermal expansion and finite lifetimes arise |
| vacancies and interstitials | translation symmetry is locally broken and carrier or phonon scattering changes |
| substitutional disorder | site energies, masses, and local environments vary |
| dislocations and strain | lattice vectors become local fields; topology and elasticity enter |
| surfaces and interfaces | coordination, electrostatics, and allowed states differ from the bulk |
| domains, twins, and grain boundaries | several orientations or order-parameter choices coexist |
| incommensurate order | no finite common primitive cell may exist |
| quasicrystalline order | sharp diffraction can occur without ordinary translation periodicity |
| finite correlation length | periodic order is approximate rather than asymptotically long-ranged |
Whether an omission is controlled depends on the observable and scale. A weak concentration of defects may barely change a band dispersion yet dominate low-temperature resistivity. Small atomic displacements can be negligible for static geometry but essential for heat capacity and superconducting pairing. A surface can be irrelevant to a bulk equation of state and decisive for photoemission or a topological phase.
Exercises
Section titled “Exercises”Exercise 1: primitive or doubled?
Section titled “Exercise 1: primitive or doubled?”For a square lattice with primitive matrix , consider
Do form a primitive basis? Repeat for
Solution
For the first pair,
and the integer matrix has determinant . It is unimodular, so are primitive and span the same lattice.
For the second pair,
whose determinant is . The cell area is , and the generated points satisfy a parity restriction on their original integer coordinates. The pair spans an index-two sublattice, not the full square lattice.
Exercise 2: why honeycomb is non-Bravais
Section titled “Exercise 2: why honeycomb is non-Bravais”Let
Show that no pure translation taking an site to a site preserves .
Solution
Any displacement taking an site to a site is congruent to modulo . It is therefore enough to test translation by :
The first coset is part of . Equality would require the second coset to equal . That would mean
which is false because its primitive coordinates are not integers. It also cannot equal , because their difference is not in . Thus a pure translation cannot interchange the two sublattices. The primitive translations are those of the triangular , and the motif has two sites.
Exercise 3: periodicity of a motif density
Section titled “Exercise 3: periodicity of a motif density”Starting from
prove for every . Identify the step that fails for a single vacancy.
Solution
Translate the argument:
Because is a group, the map
is a bijection of . Relabeling the dummy lattice vector therefore recovers .
A vacancy removes one selected term, so the translated sum no longer has the same set of terms. The relabeling argument fails at the missing cell even though the underlying reference lattice remains useful.
Exercise 4: interpreting an optical tail
Section titled “Exercise 4: interpreting an optical tail”A calculation with nuclei fixed at their ideal positions predicts a band gap, but an experiment measures optical absorption below that prediction. Give four distinct departures from the ideal lattice that could contribute, and state what additional evidence would help distinguish them. Can zero-point motion supply a phonon to be absorbed?
Solution
Possible mechanisms include:
- substitutional disorder or vacancies creating localized in-gap states;
- strain inhomogeneity shifting the local band edges;
- surfaces or interfaces supporting states absent from the bulk model;
- phonon-assisted transitions involving thermally occupied modes, or a shift of band edges caused by lattice motion;
- domains or secondary structural phases with different gaps;
- an incorrect motif, magnetic order, or interaction approximation in the nominally perfect model.
The optical and electronic thresholds must be distinguished. For an independent electron–hole pair costing energy and one phonon of energy , energy conservation gives
The oscillator matrix elements give occupation factors and , respectively. At zero temperature, the first vanishes: zero-point motion supplies no phonon to absorb. Emission remains possible but costs extra photon energy. It can enable absorption below a direct gap when the indirect pair energy is smaller; it does not by itself create absorption below the actual fundamental pair threshold. See Dresselhaus, Sec. 5.6. Zero-point and thermal motion can also renormalize that threshold relative to the fixed-nuclei calculation, a separate effect discussed by Zacharias, Patrick, and Giustino.
Useful discriminants include temperature dependence, thickness or surface sensitivity, spatially resolved spectroscopy, diffraction and pair-distribution measurements, controlled annealing or irradiation, polarization selection rules, and comparison across samples with quantified composition. Excitonic binding and errors in the electronic approximation are further possibilities beyond departures from the ideal lattice. The observation alone does not identify “disorder” as a unique cause.
The next step is the reciprocal lattice: the wavevectors whose phases are unchanged by every lattice translation.
References
Section titled “References”- M. I. Aroyo, ed., International Tables for Crystallography A: Space-Group Symmetry, 6th ed. (Wiley, 2016), especially the tables of two- and three-dimensional Bravais types.
- N. W. Ashcroft and N. D. Mermin, Solid State Physics (Holt, Rinehart and Winston, 1976), Chapters 4–7.
- P. M. Chaikin and T. C. Lubensky, Principles of Condensed Matter Physics (Cambridge University Press, 1995), Chapters 4–5.
- M. S. Dresselhaus, Solid State Physics, Part II: Optical Properties of Solids, MIT lecture notes, Sec. 5.6, Eqs. (5.49)–(5.65), open text.
- International Union of Crystallography, “Unit cell,” Online Dictionary of Crystallography, accessed 2026-10-14.
- G. L. Squires, Introduction to the Theory of Thermal Neutron Scattering, 3rd ed. (Cambridge University Press, 2012), Chapters 1–3.
- M. Zacharias, C. E. Patrick, and F. Giustino, “Stochastic Approach to Phonon-Assisted Optical Absorption,” Physical Review Letters 115, 177401 (2015), doi:10.1103/PhysRevLett.115.177401, open text. Distinguishes phonon-assisted optical transitions from electron–phonon renormalization of the band edges.