

Dipole Approximation
The electric-dipole approximation replaces the variation of an electromagnetic mode across a localized internal charge distribution by its value at one reference point. For a neutral atom or molecule in a prescribed classical field, the leading internal electric coupling is then
This compact Hamiltonian underlies much of spectroscopy, laser control, quantum optics, and atomic-clock physics. Its familiarity can hide several independent assumptions. In particular:
- it is a spatial approximation, not a weak-field approximation;
- it can be valid for internal motion while the center-of-mass phase remains essential;
- it does not imply that electric-dipole coupling is the leading nonzero transition amplitude;
- it does not by itself justify a two-level truncation, perturbation theory, a rotating-wave approximation, or a classical treatment of light.
The reliable question is therefore not merely “is the wavelength larger than the atom?” It is “which field gradients are resolved by the charge or current matrix element relevant to this observable?”
Required background. Minimal Coupling supplies the charge–field Hamiltonian; Gauge Choices explains how its length and velocity representations are related.
Internal Coordinates and the Electric Dipole
Section titled “Internal Coordinates and the Electric Dipole”Let the constituents have charges and positions
Here is an expansion point, often the center of mass, a nucleus, or a fixed molecular origin, and is an internal coordinate relative to it. Define
Most optical applications on this page concern a localized neutral system, . Charged systems require more care because uniform electric fields also accelerate the center of mass and because a permanent dipole depends on the choice of origin.
Repeated Cartesian indices are summed when an index form is used. The electric field is a real field unless a complex amplitude is displayed explicitly. The sign convention
then agrees with the electrostatic potential-energy expansion and extends to a time-dependent prescribed field through the length transformation.
Long-Wavelength Approximation
Section titled “Long-Wavelength Approximation”Taylor expansion of a smooth field
Section titled “Taylor expansion of a smooth field”Suppose the field is smooth over the support of the internal wavefunction. For each constituent,
Keeping only the first line makes the field uniform across the internal coordinates. It need not make the field uniform over the apparatus or over a center-of-mass trajectory.
For a plane-wave component,
The internal spatial factor has the expansion
If is a characteristic internal extent, the conventional long-wavelength parameter is
The leading internal replacement is controlled when . The factor of matters: the comparison is with the reduced wavelength , not merely with .
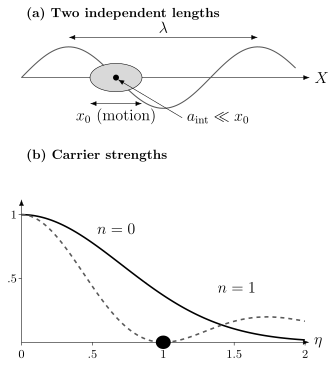
Top: a field can be nearly uniform over the internal coordinates while its phase changes over the center-of-mass wavepacket. Bottom: exact harmonic carrier strengths for depend on , not the internal atomic radius. The zero is motional interference. The derivation is given in Exercise 2.
What size should enter ka?
Section titled “What size should enter ka?”A geometric radius is only a first estimate. The relevant object is the charge or current transition distribution, not necessarily the full diameter of the system. Define the charge-density operator
and its transition density
At a fixed expansion point , an optional diagnostic length is
This absolute-density definition is a scale estimate when its denominator is nonzero and its second moment exists. A purely spin-changing transition can have zero transition charge density but nonzero magnetic coupling; then a charge-based size is undefined. A more decisive test is to calculate the first omitted interaction matrix element and compare it with the retained one.
This distinction matters in X-ray spectroscopy. A molecule may be several ångströms across while a particular core transition density is concentrated near one nucleus. Conversely, a Rydberg or continuum state can sample a much larger region than the ground-state radius suggests.
General modes and near fields
Section titled “General modes and near fields”For a cavity mode, focused beam, evanescent wave, or near field, may not be the shortest spatial scale. A local gradient length can be defined schematically by
where is a nonzero field scale appropriate to the experiment. The corresponding estimate is
At a node, using the local as the denominator is meaningless because it vanishes. One must compare the gradient-driven and field-driven matrix elements directly. Structured light can therefore make beyond-dipole coupling leading even when is very small.
Choosing the Expansion Point
Section titled “Choosing the Expansion Point”Neutral and charged systems
Section titled “Neutral and charged systems”Shift the reference point by a fixed vector :
The dipole operator transforms as
For a neutral system, as an operator. For a charged system, the permanent dipole is origin dependent. However, between distinct orthonormal states in the same charge sector,
Thus an off-diagonal transition dipole is origin independent under a fixed shift, even for a charged system. Diagonal moments and center-of-mass couplings still require the charge and origin to be stated.
Higher multipoles mix under shifts
Section titled “Higher multipoles mix under shifts”Beyond E1, shifting the expansion point redistributes contributions among multipole orders and changes the plane-wave phase. An exact, untruncated interaction is independent of that bookkeeping. A truncated expression is origin independent only when all terms required at the claimed order are kept consistently.
This is especially important for oscillator strengths beyond the dipole approximation. Keeping the square of an E2 or M1 amplitude while dropping interference terms of the same order can create spurious origin dependence.
Electric Dipole Hamiltonian
Section titled “Electric Dipole Hamiltonian”Multipolar form
Section titled “Multipolar form”For a localized neutral system in a smooth external field, the leading internal term in the multipolar expansion is
This is the length-form interaction for a prescribed field and fixed expansion point. Starting in a velocity representation, one can replace the vector potential by its value over the internal coordinates and perform the corresponding time-dependent unitary transformation. If the center of mass is dynamical, transforming its kinetic energy also generates motion-dependent terms, including the Röntgen coupling of a moving dipole to the magnetic field. Neglecting these requires a separate slow-motion approximation. “Dipole approximation” and “length gauge” are therefore related but not logically identical statements.
For a charged composite system, the complete long-wavelength Hamiltonian also contains center-of-mass coupling to the monopole charge. Writing only silently discards that dynamics.
Classical field convention
Section titled “Classical field convention”For a monochromatic classical field, write
Then
The factor belongs to this complex-amplitude convention. If is instead defined as a positive-frequency field without the usual real-field decomposition, the factor changes. Many apparent factor-of-two disagreements in Rabi frequencies are convention disagreements.
Quantized field
Section titled “Quantized field”In the quantized multipolar representation, distinguish the canonical radiative field from the physical transverse field . Here is the transverse matter polarization. For vacuum transverse modes normalized by , the canonical field has the expansion
The dipole interaction is then
Mode normalization may place a quantization volume or effective mode volume inside . A single-mode coupling is often defined by
up to a phase convention. The same long-wavelength test applies to the mode function: it must vary little over the internal transition distribution.
The interaction accompanies the polarization self-energy : together they come from expanding . Replacing by the physical field while leaving that self-energy unchanged gives a different Hamiltonian. The mode cutoff and finite spatial polarization must also be specified before using an ideal point dipole. Gauge Choices develops these representation and truncation issues.
Fixed nuclei and molecular coordinates
Section titled “Fixed nuclei and molecular coordinates”For electrons at and nuclei at ,
At fixed nuclear geometry, the nuclear term is proportional to the identity in the electronic Hilbert space and therefore drops from an off-diagonal electronic transition dipole. It does not disappear from permanent dipoles, vibrational transitions, vibronic coupling, or nuclear dynamics.
In the Born–Oppenheimer description, a geometry-dependent electronic dipole surface can couple vibrational states. Infrared intensity is controlled by how the molecular dipole changes along a normal coordinate, not merely by whether the equilibrium molecule has a permanent dipole.
Transition Dipole Moments
Section titled “Transition Dipole Moments”Definition
Section titled “Definition”For matter eigenstates and , define
The E1 matrix element is
For a classical complex amplitude, a common Rabi-frequency convention is
With the real-field convention above, the resonant rotating-wave Hamiltonian contains . A transition rate requires additional assumptions about time dependence, density of final states, field statistics, and line broadening.
Transition density interpretation
Section titled “Transition density interpretation”For a fixed expansion point (or internal matrix elements at fixed center of mass), the same matrix element can be written
For distinct states in one charge sector,
The transition density therefore carries no net transition charge. The transition dipole is its first spatial moment. Its generally complex contributions can cancel; positive and negative regions are the special case of a real transition density. A large extent therefore does not guarantee a large dipole matrix element.
Permanent and transition dipoles are different
Section titled “Permanent and transition dipoles are different”The diagonal quantity
is a permanent-state dipole. The off-diagonal quantity is a transition dipole. A state can have zero permanent dipole and still have strong E1 transitions. Conversely, a polar molecule can have a nonzero body-fixed permanent dipole while a particular transition dipole vanishes. A freely rotating, definite-parity molecular eigenstate nevertheless has zero laboratory-frame permanent electric dipole; orientation or parity mixing is required to obtain a nonzero laboratory expectation.
Polarization projection
Section titled “Polarization projection”For a mode with polarization , only
enters the E1 amplitude. The vector can be complex when the states or polarization basis are complex. Observable strengths involve absolute squares and, where appropriate, sums or averages over magnetic sublevels and polarizations.
The rotational and parity conditions under which this projection vanishes belong to Dipole Transitions. “Allowed” means not forced to zero by the stated symmetry; it does not mean large.
Degenerate subspaces
Section titled “Degenerate subspaces”Within a degenerate manifold, individual vectors depend on the chosen basis. A physical preparation, field polarization, external splitting, or incoherent average selects the relevant combination. Summing over a complete unresolved degenerate manifold produces a basis-independent strength, while quoting one arbitrarily chosen component may not.
Length and velocity matrix elements
Section titled “Length and velocity matrix elements”For a local nonrelativistic Hamiltonian
the commutator
implies, for exact eigenstates,
This relation makes exact length- and velocity-form E1 amplitudes equivalent when all terms are treated consistently. Incomplete bases, nonlocal pseudopotentials, inconsistent energies, or omitted contact terms can spoil the numerical agreement. That disagreement is a convergence diagnostic, not a physical gauge dependence.
Validity Conditions
Section titled “Validity Conditions”Internal spatial variation
Section titled “Internal spatial variation”The first check is
for every appreciably occupied spectral component and for the transition distribution of interest. For a broadband pulse, use the largest relevant , not only the carrier wave number.
For a general field, also check
These estimates test the expansion, not the dominance of a particular transition.
Transition-specific amplitude test
Section titled “Transition-specific amplitude test”When the E1 amplitude is nonzero, the most useful diagnostic is
If the denominator vanishes or is accidentally small, this ratio is not a failure of arithmetic. It says that E1 is not the reference process. Compare each candidate amplitude with the largest retained physical amplitude instead.
Center-of-mass phase is a separate approximation
Section titled “Center-of-mass phase is a separate approximation”After making the internal dipole approximation, a plane-wave interaction still contains
The factor transfers photon momentum and couples center-of-mass motional states. Removing it requires a second approximation about the external wavepacket or trajectory.
For motional states ,
sets carrier and sideband strengths. In one harmonic direction with
the Lamb–Dicke parameter is
A useful state-dependent condition is
The effective wave vector is for a one-photon process and often a wave-vector difference for a Raman process. It is entirely possible to have
while not being in the Lamb–Dicke regime. The first statement concerns internal electronic or molecular structure; the second concerns external motion.
The recoil energy
and the Doppler shift also survive the internal dipole approximation.
Beam profile and structured light
Section titled “Beam profile and structured light”A paraxial beam introduces transverse and longitudinal scales in addition to . For waist , a localized internal system usually satisfies , but center-of-mass motion may sample the profile. Near a focus, interface, nanostructure, or optical vortex, longitudinal fields and steep gradients can matter.
At an electric-field node, the E1 amplitude can vanish because even though the E1 transition dipole is symmetry allowed. M1 or E2 coupling may then be the leading local interaction. This is a mode-geometry statement, not an intrinsic atomic selection rule.
Strong fields and changing length scales
Section titled “Strong fields and changing length scales”The dipole approximation does not require a weak field. Intense-field calculations often use a dipole Hamiltonian nonperturbatively. Nevertheless, the field can drive the wavefunction into Rydberg or continuum regions with a larger spatial extent, making the initial bound-state radius a poor value of .
Strong fields can independently invalidate:
- perturbation theory;
- a two-level reduction;
- the rotating-wave approximation;
- neglect of ionization or continuum states;
- a nonrelativistic treatment;
- a prescribed-field model that ignores depletion or backaction.
None of these failures follows solely from .
Temporal bandwidth
Section titled “Temporal bandwidth”Spatial and temporal variations are linked for freely propagating radiation through , but the approximation is still made in space. For a pulse with spectral support up to , a conservative test is
Using only the carrier can miss a high-frequency tail. A short pulse does not otherwise invalidate E1 coupling merely because it is short.
Near fields and material environments
Section titled “Near fields and material environments”Evanescent and quasistatic fields can have gradient scales set by distance to a surface, aperture size, or nanostructure geometry rather than by the free-space wavelength. The local test is then , together with direct M1 and E2 matrix-element comparisons.
In periodic systems, multiplying a wavefunction by position does not preserve its periodic boundary conditions. Current operators, Berry-phase polarization, and the chosen boundary conditions must be handled consistently. The localized atom-and-molecule construction here should not be transplanted unchanged into a crystal.
Worked Scale Estimates
Section titled “Worked Scale Estimates”Visible light on a compact atom
Section titled “Visible light on a compact atom”Take
Then
Internal spatial variation is very small. This does not prove that a chosen line is E1 allowed, but it makes the long-wavelength expansion strongly controlled for compact bound-state matrix elements.
Infrared light on a molecule
Section titled “Infrared light on a molecule”For
one finds
The field is nearly uniform over the molecular framework. Vibrational intensity still depends on the derivative of the molecular dipole surface along the normal coordinate.
X-rays and a localized core transition
Section titled “X-rays and a localized core transition”For a hard X-ray wavelength
and a geometric molecular scale ,
A molecular-size dipole approximation is uncontrolled. If the relevant core transition density has an effective extent , then
which is smaller but still not asymptotically small. A calculation may need the full spatial interaction rather than a low-order multipole truncation.
Internal dipole regime without Lamb–Dicke confinement
Section titled “Internal dipole regime without Lamb–Dicke confinement”Consider a trapped ion with
The internal parameter is
For the actual 729 nm clock transition, the E1 matrix element is forbidden and E2 coupling is leading. The small value of validates a rapidly ordered internal multipole expansion; it does not promote a vanishing E1 amplitude.
For mass and trap frequency ,
The internal long-wavelength expansion is excellent. At ,
so replacing the motional phase by unity is not well controlled. Internal and external long-wavelength statements give different answers because they address different coordinates.
An allowed transition at a standing-wave node
Section titled “An allowed transition at a standing-wave node”Let
and place the expansion point at . Although the internal parameter can satisfy ,
The local E1 term vanishes. The first electric gradient is
so an electric-quadrupole matrix element can become leading. The standing wave also has a magnetic field, so the M1 amplitude must be checked at the same order.
Breakdown and Multipole Corrections
Section titled “Breakdown and Multipole Corrections”First omitted interactions
Section titled “First omitted interactions”Introduce the primitive electric quadrupole tensor
With this convention, the first terms of a local multipolar interaction can be organized schematically as
The magnetic dipole contains orbital and, where relevant, spin contributions. The displayed formula fixes the primitive-quadrupole normalization used here. For an electrostatic external field, includes . A charged moving system also has center-of-mass vector-potential coupling, so is not exhausted by that scalar term.
Alternatively, define a traceless Cartesian quadrupole
In a source-free region where ,
Factors such as and cannot be compared across references without checking which quadrupole tensor has been defined.
Expected power counting
Section titled “Expected power counting”For a plane wave and typical nonvanishing moments,
For nonrelativistic internal motion, a typical M1-to-E1 amplitude can scale like . In atoms driven near internal resonances, these estimates may be of similar small order. Neither is universal: symmetry, cancellations, relativistic mixing, molecular geometry, and field structure can change the actual ratio.
The hierarchy is a statement about amplitudes. A correction of order in amplitude generally contributes at order through interference with E1 and at order through its own square, subject to symmetry and orientation averages.
Consistent expansion of probabilities
Section titled “Consistent expansion of probabilities”Write a transition amplitude as
where is nominally order . Through second order,
Keeping while omitting the interference is not a complete second-order calculation. Such partial truncations can spoil origin independence and sum rules.
When the series is not efficient
Section titled “When the series is not efficient”If , many multipole orders can contribute. In that regime, retaining the exact factor
or the full current–field interaction is often better than extending a slowly convergent multipole series. The exact spatial operator also avoids assigning physical significance to individual origin-dependent truncated pieces.
This can occur in X-ray processes, extended Rydberg or continuum motion, and strongly structured near fields; none of these labels alone proves that a given transition is outside the dipole regime. Relativistic corrections may also become important at high photon energy or for heavy elements; they are independent of the multipole truncation.
Forbidden E1 transitions
Section titled “Forbidden E1 transitions”If
then M1, E2, two-photon coupling, state mixing, or another mechanism may provide the leading transition. Calling the line “dipole forbidden” means only that its E1 amplitude vanishes under the stated model and symmetries.
Multipole Operators explains the rank and parity classification of E1, M1, E2, and higher operators. The next article, Multipole Expansion, develops their AMO interaction hierarchy and precision-spectroscopy applications.
Exercises
Section titled “Exercises”1. Bound the plane-wave error
Section titled “1. Bound the plane-wave error”Show that
Use the result to bound the pointwise internal field error for and .
Solution
For real ,
The triangle inequality and give
Set . If , then
Numerically,
The pointwise complex field differs from its uniform value by no more than about over the stated region. A transition amplitude can have a larger relative correction if its leading E1 integral nearly cancels.
2. Compute a carrier amplitude without expanding the phase
Section titled “2. Compute a carrier amplitude without expanding the phase”For a harmonic center-of-mass coordinate and , compute the carrier factors for . Explain why the internal condition does not make these factors unity.
Solution
Since , the Baker–Campbell–Hausdorff identity gives
For the vacuum, only the constant terms survive. For , one lowering and one raising also contribute, with product :
Thus the squared carrier matrix elements are and . At , the carrier vanishes by motional interference even for a perfectly E1-allowed internal transition. These are coupling strengths; an excitation probability also depends on pulse duration, detuning, and competing sidebands. The internal size and the trap length are different physical scales.
3. Estimate an E2-to-E1 ratio
Section titled “3. Estimate an E2-to-E1 ratio”Suppose a transition has representative matrix elements
For a plane wave at , estimate
Solution
Substitution gives
Using
one obtains
This is only a scale estimate. Angular factors, tensor components, and cancellations must be evaluated for the actual states. If , this ratio is not the appropriate comparison.
4. Test a broadband pulse
Section titled “4. Test a broadband pulse”A localized transition has . A pulse has a carrier wavelength of but appreciable spectral weight down to . Compare the carrier-only and conservative long-wavelength parameters.
Solution
At the carrier,
At the shortest appreciably occupied wavelength,
Both are smaller than unity, but the conservative correction estimate is ten times larger. Whether in amplitude is acceptable depends on the observable and required precision. The pulse duration alone was not the criterion; its spectral support supplied the relevant .
5. Transition charge and origin independence
Section titled “5. Transition charge and origin independence”Starting from
show that the transition charge vanishes for distinct states in one charge sector. Then prove directly from the integral definition that is invariant under .
Solution
Integrating the charge-density operator gives the total-charge operator:
Within a fixed charge sector,
Therefore
for .
After shifting the reference point,
The proof uses orthogonality and fixed total charge, not neutrality.
References
Section titled “References”- D. L. Andrews, G. A. Jones, A. Salam, and R. G. Woolley, “Perspective: Quantum Hamiltonians for optical interactions,” Journal of Chemical Physics 148, 040901 (2018), doi:10.1063/1.5018399.
- M. Babiker and R. Loudon, “Derivation of the Power–Zienau–Woolley Hamiltonian in quantum electrodynamics by gauge transformation,” Proceedings of the Royal Society A 385, 439–460 (1983), doi:10.1098/rspa.1983.0022.
- S. Bernadotte, A. J. Atkins, and C. R. Jacob, “Origin-independent calculation of quadrupole intensities in X-ray spectroscopy,” Journal of Chemical Physics 137, 204106 (2012), doi:10.1063/1.4766359.
- C. Cohen-Tannoudji, J. Dupont-Roc, and G. Grynberg, Photons and Atoms: Introduction to Quantum Electrodynamics, Wiley, 1989.
- C. Cohen-Tannoudji, J. Dupont-Roc, and G. Grynberg, Atom–Photon Interactions: Basic Processes and Applications, Wiley, 1992.
- D. P. Craig and T. Thirunamachandran, Molecular Quantum Electrodynamics, Dover, 1998.
- W. E. Lamb, R. R. Schlicher, and M. O. Scully, “Matter-field interaction in atomic physics and quantum optics,” Physical Review A 36, 2763–2772 (1987), doi:10.1103/PhysRevA.36.2763.
- P. J. Lestrange, F. Egidi, and X. Li, “The consequences of improperly describing oscillator strengths beyond the electric dipole approximation,” Journal of Chemical Physics 143, 234103 (2015), doi:10.1063/1.4937410.
- N. H. List, T. R. L. Melin, M. van Horn, and T. Saue, “Beyond the electric-dipole approximation in simulations of X-ray absorption spectroscopy: Lessons from relativistic theory,” Journal of Chemical Physics 152, 184110 (2020), doi:10.1063/5.0003103.
- E. A. Power and S. Zienau, “Coulomb gauge in non-relativistic quantum electro-dynamics and the shape of spectral lines,” Philosophical Transactions of the Royal Society A 251, 427–454 (1959), doi:10.1098/rsta.1959.0008.
- D. J. Wineland, C. Monroe, W. M. Itano, D. Leibfried, B. E. King, and D. M. Meekhof, “Experimental issues in coherent quantum-state manipulation of trapped atomic ions,” Journal of Research of the National Institute of Standards and Technology 103, 259–328 (1998), doi:10.6028/jres.103.019.