

Hund's Rules
Hund’s rules are an ordered set of approximate rules for predicting the lowest spectroscopic term and fine-structure level associated with a declared free-atom electron configuration in the -coupling regime. For one open subshell, their familiar form is:
- Among allowed terms of the configuration, the term with the largest total spin usually lies lowest.
- Among the terms with that largest , the term with the largest total orbital angular momentum usually lies lowest.
- Within the selected term, the smallest allowed usually lies lowest when the open subshell is less than half filled, while the largest usually lies lowest when it is more than half filled.
The ordering is lexicographic. Rule 2 is applied only among terms surviving Rule 1, and Rule 3 is applied only after an term has been selected. The rules do not replace antisymmetrization, determine the ground electron configuration, or provide exact energies.
Their best domain is a free atom or ion with one open subshell, a reasonably isolated configuration, and sufficiently good coupling. Outside that domain they remain clues, not verdicts.
Required background. Pauli antisymmetry determines the allowed terms; exchange and correlation supplies the fixed-orbital energy comparison. Use LS coupling and term symbols to interpret .
What the Rules Do and Do Not Order
Section titled “What the Rules Do and Do Not Order”Consider a central-field configuration
Several different objects can be associated with this one label:
- determinants with specified one-electron magnetic occupations;
- antisymmetric configuration-state functions with specified and ;
- terms labeled by ;
- fine-structure levels labeled by ;
- exact eigenstates that may mix several configurations and terms.
Hund’s first two rules compare terms within a declared configuration. The third compares levels within the selected term. A useful ideal hierarchy is
The first two rules summarize regularities in the electrostatic term splittings. The third uses the weaker spin–orbit ordering. If these scales overlap, the labels themselves can mix and the three-step procedure loses accuracy.
Four ideas that are often conflated
Section titled “Four ideas that are often conflated”- Pauli exclusion is a kinematic consequence of fermionic antisymmetry. It forbids repeated complete spin-orbitals and removes inadmissible terms.
- A configuration or filling rule proposes which subshell occupations should be compared for the ground state.
- Hund’s rules rank allowed terms and levels, approximately, after a configuration has been declared.
- Hamiltonian diagonalization supplies the actual energies and eigenvectors for the model being used.
The classroom instruction “put one electron in each degenerate orbital before pairing” is a determinant-level way to construct a maximum- representative of Rule 1. It is not the complete content of all three rules. It also assumes a degenerate or nearly degenerate orbital set; a sufficiently large orbital-energy splitting can favor pairing.
Free atoms, not arbitrary environments
Section titled “Free atoms, not arbitrary environments”The quantum numbers , , and refer to rotational symmetry of an isolated atom. A crystal field can split orbital components and quench ; a molecule has different spatial symmetry and often different equilibrium geometries for different spin states. Molecular Hund cases are angular-momentum coupling schemes, not these atomic term-ordering rules.
Rule 1: Maximum Spin
Section titled “Rule 1: Maximum Spin”Within a fixed configuration, the allowed term with the largest commonly has the lowest electrostatic energy. Its multiplicity is
“Parallel spins” is shorthand for the stretched component with . A rotationally invariant term also contains the other components. The rule does not assign classical arrows permanently pointing in one laboratory direction.
Two-electron fixed-orbital model
Section titled “Two-electron fixed-orbital model”For two orthonormal spatial orbitals , the singlet has symmetric spatial part and the triplet antisymmetric part . The direct and exchange calculation gives
The same one-electron orbitals are used in both trial states. The Coulomb exchange integral , so the triplet lies lower by in this restricted comparison. Its spatial wavefunction vanishes when the two positions coincide, expressing the same-spin exchange hole.
The important energy-accounting caveat
Section titled “The important energy-accounting caveat”The fixed-orbital derivation is exact for the trial functions just declared. It does not prove that a fully optimized high-spin atom has a smaller expectation value of electron–electron repulsion than the corresponding low-spin atom.
When each term is allowed to relax, its orbitals, kinetic energy, electron–nucleus attraction, direct energy, exchange energy, and correlation energy all change. Virial-theorem and multiconfiguration analyses of second- and third-row atoms find that the high-spin stabilization can be realized through stronger electron–nucleus attraction after radial contraction, even when the electron–electron repulsion and kinetic energy increase.
The trustworthy summary is:
Antisymmetry creates spin-dependent spatial structure. In a common fixed-orbital model this appears as a lowering by exchange integrals; in a relaxed atom, the final term separation is a total-energy balance.
This is why “electrons with parallel spins repel less” is too literal. The Coulomb operator is spin independent, and no additional exchange force has been added.
Why Pauli is not Rule 1
Section titled “Why Pauli is not Rule 1”Pauli antisymmetry determines the admissible Hilbert space. It can force two electrons in the same spatial orbital into a singlet, as in . For electrons in distinct orbitals, both singlet and triplet terms may be allowed. Hund’s first rule then predicts an energy ordering between allowed terms; it is not an exclusion law.
Rule 2: Maximum Orbital Angular Momentum
Section titled “Rule 2: Maximum Orbital Angular Momentum”After selecting the maximum- terms, the one with the largest commonly lies lowest. The ordering applies to total orbital angular momentum,
not to a sum of one-electron orbital energies.
For one equivalent open subshell, electrostatic term energies can be organized schematically as
Here is a configuration average, the are radial Slater–Condon integrals, and the dimensionless are angular coefficients fixed by the antisymmetric term. Rule 2 summarizes a frequent pattern of these angular coefficients. It is not a universal centrifugal-energy theorem proportional to .
Why the common orbital picture is limited
Section titled “Why the common orbital picture is limited”A stretched maximum- component places the parallel-spin electrons in the largest available values. This construction is excellent for finding the candidate term. Its angular density and exchange correlations often support efficient electron avoidance, but statements such as “the electrons orbit in the same direction and stay farther apart” are classical pictures, not derivations.
The actual comparison uses matrix elements of the Coulomb Hamiltonian. Different radial functions, several open subshells, configuration interaction, and unusually large can invalidate the simple ordering.
Equivalent-Subshell Construction
Section titled “Equivalent-Subshell Construction”Let electrons occupy one subshell. The number of spatial orbitals is
and the spin-orbital capacity is
Half filling occurs at .
Maximum spin
Section titled “Maximum spin”For , place the electrons in distinct orbitals with the same spin projection. Then
For , use the equivalent hole description with
The compact result is
The electron and hole configurations and support the same set of and terms, although their fine-structure ordering reverses across half filling.
Maximum orbital angular momentum
Section titled “Maximum orbital angular momentum”For the stretched maximum-spin component with , occupy
The largest projection is
A state with the largest possible projection must belong to a term with
Combining Rules 1 and 2 therefore predicts, for one equivalent open subshell,
The subscript marks the Hund-rule candidate, not an exact theorem about the full atomic ground state.
Quick checks
Section titled “Quick checks”- : , so and , giving .
- : , so and , giving .
- : , so and , giving .
- : , so and , giving .
- : , so and , giving .
These formulas identify the term selected by the first two rules. They do not enumerate all allowed terms, which still requires antisymmetry and state counting.
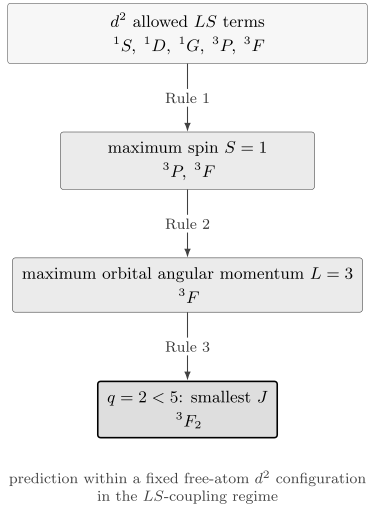
For a fixed configuration in coupling, Rule 1 retains the triplets, Rule 2 selects , and Rule 3 selects because the subshell is less than half filled. The diagram is a prediction workflow, not a proof of the level energies.
Rule 3: Ordering the J Levels
Section titled “Rule 3: Ordering the J Levels”Once Rules 1 and 2 have selected an term, total angular momentum can take the values
In an isolated term with weak fine structure, the leading effective interaction is often written
Here has energy units and the angular momenta have their usual units of . Its first-order level energy is
Adjacent levels obey
Therefore:
- if , energy increases with and the smallest lies lowest;
- if , energy decreases with and the largest lies lowest.
For a single equivalent open subshell in ordinary coupling, the effective sign commonly follows the filling:
At exact half filling, the maximum-spin term of one equivalent subshell has , so only occurs. There is no multiplet ordering left for Rule 3 to decide.
Particle–hole reversal
Section titled “Particle–hole reversal”Configurations and have the same allowed and terms. Within a fixed radial subshell and first-order one-electron spin–orbit theory, their electron and hole descriptions reverse the spin–orbit ordering within corresponding terms. Thus
share the Hund candidate , but Rule 3 predicts
This is the logic behind the less-than-half-filled and more-than-half-filled wording. It is not a statement that the magnitude of every fine-structure interval is the same for electron and hole partners.
Worked d² interval pattern
Section titled “Worked d² interval pattern”For , one has and , so . The effective shifts are
For , Rule 3 corresponds to , so lies lowest. The interval ratio is
This is the isolated-term Landé interval prediction. Real intervals need not obey it exactly because spin–spin, spin–other-orbit, configuration interaction, and intermediate coupling also contribute. LS coupling explains how departures from this ratio diagnose additional interactions.
Worked Ground-Term Examples
Section titled “Worked Ground-Term Examples”The following examples use closed cores with , so only the open subshell needs to be considered. NIST evaluated ground-level designations provide the empirical check.
Carbon, nitrogen, and oxygen
Section titled “Carbon, nitrogen, and oxygen”The neutral atoms have closed cores and open subshells. Carbon has , so , , and : the prediction is . Nitrogen has and a half-filled subshell: the stretched occupations give , , hence . The letter denotes total , not occupation of an orbital.
Oxygen’s is equivalent to two holes for the term count. It therefore has the same candidate as carbon, but the more-than-half-filled shell selects . All three predictions agree with the NIST ground-level designations. Parity is : odd for nitrogen, even for carbon and oxygen.
Titanium and nickel: d² and d⁸
Section titled “Titanium and nickel: d² and d⁸”The leading neutral-titanium ground configuration contains . The closed pair contributes zero angular momentum, and the open subshell gives
The NIST level compilation places the other members of this term at higher energy in the order
Neutral nickel’s leading ground configuration contains . The open subshell is the two-hole partner of , so it has the same term but the opposite third-rule ordering:
The evaluated Ti I and Ni I ground designations thus give a direct electron–hole comparison.
Limitations and Exceptions
Section titled “Limitations and Exceptions”Hund’s rules are remarkably successful for many free-atom ground configurations, but no variational theorem says that they must hold for every Hamiltonian or every configuration.
Configuration mixing
Section titled “Configuration mixing”The rules rank terms associated with a declared configuration. If two configurations of the same exact are close, the Hamiltonian can mix them:
Electrostatic configuration interaction can mix configurations while preserving and . Relativistic interactions can additionally mix different terms with the same . A dominant-component label can remain useful, but its weight should be reported.
Several open subshells
Section titled “Several open subshells”The compact formulas using apply to one equivalent open subshell. With two or more open subshells, one must first determine parent terms and their coupling. Kutzelnigg and Morgan’s review emphasizes that exceptions to simple first- and second-rule forms occur especially in configurations with multiple open subshells carrying nonzero orbital angular momentum and in some large- cases.
Relativistic and intermediate coupling
Section titled “Relativistic and intermediate coupling”The first two rules assume that and organize the electrostatic terms before fine structure is applied. In a heavy atom, strong one-electron spin–orbit splitting can favor a jj-coupled description. Then individual occupations and subshell parents can be more informative than one pure term.
Even when labels remain recognizable, same- mixing can move levels away from the third-rule pattern. A statement such as “more than half filled, therefore largest ” is not reliable unless the isolated-term and coupling assumptions have been checked.
Excited configurations and accidental near-degeneracy
Section titled “Excited configurations and accidental near-degeneracy”The rules are most dependable as ground-term guides. They do not globally sort terms from different configurations, and rare reversals occur among excited states. Small denominators amplify interactions that a broad energy-scale argument would otherwise treat as perturbations.
Explicit calculations along isoelectronic sequences provide documented examples of excited-state failures and changing assignments. Such cases are evidence against overextending the rules, not failures of antisymmetry or angular-momentum algebra.
Orbital relaxation and correlation
Section titled “Orbital relaxation and correlation”The fixed-orbital exchange formula explains an important tendency, but optimized terms use different radial functions and correlation patterns. The energy components
can each move in a direction that contradicts a one-sentence “less repulsion” story while their sum still obeys Rule 1. Only total energies computed with a consistent Hamiltonian decide the ordering.
External fields, solids, and molecules
Section titled “External fields, solids, and molecules”Electric, magnetic, crystal, and ligand fields can compete with the atomic electrostatic and spin–orbit scales. In a ligand field, the cost of occupying a higher crystal-field orbital can exceed the intra-atomic high-spin preference and produce a low-spin state. This is not a violation of an exact atomic law; it is a different Hamiltonian outside the free-atom assumptions.
Hund coupling in many-body models
Section titled “Hund coupling in many-body models”Multi-orbital Hubbard and Kanamori models often contain a parameter called Hund coupling or Hund exchange, usually written . It lowers selected high-spin local multiplets and may include spin-flip and pair-hopping terms required by rotational invariance.
That model parameter is related to the same atomic interaction physics, but it is not identical to the three spectroscopic rules. In particular, a local does not by itself reproduce free-atom Rule 2 or the less-than-half-filled versus more-than-half-filled fine-structure ordering. See the Hubbard Model for the lattice-model boundary.
Exercises
Section titled “Exercises”Exercise 1: Beyond the repeated p and d examples
Section titled “Exercise 1: Beyond the repeated p and d examples”Predict the Hund-rule ground levels of and , including parity.
Solution
An subshell has and capacity . Both configurations have , hence and , denoted . Their terms are : both contain an odd number of odd-parity electrons. The less-than-half-filled selects , while selects . These are isolated-subshell LS predictions; they do not determine which configuration is lowest for a specified atom.
Exercise 2: Exchange energy and its limit
Section titled “Exercise 2: Exchange energy and its limit”Two orthonormal fixed orbitals have exchange integral . What singlet–triplet splitting does the fixed-orbital model predict? Does this imply that the fully optimized triplet must have a lower electron–electron repulsion expectation than the fully optimized singlet?
Solution
The fixed-orbital result is
Thus the triplet is lower by in that restricted comparison.
The second conclusion does not follow. Once each term is optimized, its orbitals and correlation change. The triplet can have a larger electron–electron repulsion and kinetic energy but a sufficiently stronger electron–nucleus attraction to produce the lower total energy. Only a comparison of total energies under the same Hamiltonian determines the ordering.
Exercise 3: Diagnose when the rules are insufficient
Section titled “Exercise 3: Diagnose when the rules are insufficient”A heavy atom has two open subshells. A calculation finds three same- components with different labels and weights , , and in one orthonormal LS basis. The observed magnetic factor differs by from the pure-LS prediction. Is a Hund-rule ground-level assignment by itself trustworthy?
Solution
No. The system violates several assumptions behind a standalone Hund assignment:
- more than one open subshell makes the simple equivalent-subshell formulas inapplicable;
- no component dominates strongly;
- significant weights in different LS terms indicate intermediate coupling;
- the magnetic factor directly contradicts a pure-LS description.
Hund’s rules may still suggest useful basis components, but the assignment should come from diagonalizing the declared Hamiltonian in the full fixed- space and comparing energies, eigenvector compositions, magnetic factors, and transitions. Reporting only one pure term label would hide the relevant physics.
References
Section titled “References”- E. U. Condon and G. H. Shortley, The Theory of Atomic Spectra, Cambridge University Press, 1935.
- R. D. Cowan, The Theory of Atomic Structure and Spectra, University of California Press, 1981.
- G. Friesecke and B. D. Goddard, “Atomic Structure via Highly Charged Ions and Their Exact Quantum States”, Physical Review A 81, 032516 (2010).
- F. Hund, “Zur Deutung verwickelter Spektren, insbesondere der Elemente Scandium bis Nickel”, Zeitschrift für Physik 33, 345–371 (1925).
- W. R. Johnson, Atomic Structure Theory: Lectures on Atomic Physics, Springer, 2007.
- A. Kramida, Yu. Ralchenko, J. Reader, and the NIST ASD Team, NIST Atomic Spectra Database, version 5.12, National Institute of Standards and Technology, data content updated November 2024, DOI: 10.18434/T4W30F, accessed 2026-07-21.
- W. Kutzelnigg and J. D. Morgan III, “Hund’s Rules”, Zeitschrift für Physik D 36, 197–214 (1996).
- W. C. Martin and W. L. Wiese, “Atomic Spectroscopy: An Introduction”, in G. W. F. Drake, ed., Atomic, Molecular, and Optical Physics Handbook, AIP Press, 1996; NIST online revision updated 2025.
- J. D. Morgan III and W. Kutzelnigg, “Hund’s Rules, the Alternating Rule, and Symmetry Holes”, Journal of Physical Chemistry 97, 2425–2434 (1993).
- NIST Physical Measurement Laboratory, evaluated ground-state pages for C I, N I, O I, Ti I, and Ni I, accessed 2026-07-21.
- T. Oyamada, K. Hongo, Y. Kawazoe, and H. Yasuhara, “Unified Interpretation of Hund’s First and Second Rules for 2p and 3p Atoms”, Journal of Chemical Physics 133, 164113 (2010).
- J. C. Slater, Quantum Theory of Atomic Structure, Vols. I–II, McGraw–Hill, 1960.
- I. I. Sobelman, Atomic Spectra and Radiative Transitions, 2nd ed., Springer, 1992.