

Thermodynamic Potentials
Thermodynamic potentials are different coordinate descriptions of the same equilibrium thermodynamics. Each potential replaces one or more extensive variables by the intensive variables imposed by a reservoir. The replacement is a Legendre transform, and it determines both the differential identities and the equilibrium principle appropriate to the experimental controls.
For a simple one-component system, the central relations are
Their natural variables differ:
The subscript on distinguishes the grand potential from symbols sometimes used for microcanonical state counts. All four quantities have units of energy.
Assume positive temperature and equilibrium throughout the minimum and curvature statements below. The algebraic thermodynamic relations use a smooth macroscopic equation of state; exact finite-system partition sums require the qualifications given below. In the grand-canonical expressions, denotes the mean number. For a potential constrained by an order parameter, see Landau theory.
Helpful background. The canonical and grand-canonical ensembles supply the states whose energy, entropy, and number appear here.
Energy and entropy in an equilibrium family
Section titled “Energy and entropy in an equilibrium family”Let be the quantum Hamiltonian and a normalized density operator. The microscopic expectation value of energy is
The von Neumann entropy, with thermodynamic units, is
These formulas apply to finite quantum systems. Thermodynamic potentials become thermodynamic state functions when equilibrium macrostates can be parameterized by variables such as and . The Thermodynamic Limit page explains the scaling assumptions behind that macroscopic description.
The scaling assumptions behind “extensive” and “intensive,” including boundary corrections and failures of additivity, are developed in Extensive and Intensive Quantities.
Two distinctions matter from the beginning:
- is an operator; is an expectation value or thermodynamic state function.
- is defined for any density operator; thermodynamic entropy is its equilibrium use together with specified macroscopic constraints.
The thermodynamic relations below therefore require more than a formal density matrix. They assume an equilibrium family, specified controls, and enough regularity to define the stated derivatives.
Fundamental Thermodynamic Relation
Section titled “Fundamental Thermodynamic Relation”For a simple compressible, one-component system,
This differential defines the intensive variables in the internal-energy representation:
The minus sign in the pressure term follows the convention that positive pressure does positive work when the system expands. If additional work coordinates are present, the fundamental relation acquires terms of the form
with signs fixed by the work convention and by how the Hamiltonian depends on the external controls.
The entropy representation follows by solving locally for :
Thus entropy and internal energy contain the same equilibrium equation of state when the transformation is regular. The choice between them is a choice of natural variables and equilibrium principle, not a choice of different physics.
Why Change Potentials?
Section titled “Why Change Potentials?”The equilibrium entropy principle compares macrostates at fixed total energy, volume, and particle number. It does not itself prove that an isolated system dynamically approaches the maximizing state. A subsystem coupled to a large reservoir is controlled by different variables. The reservoir fixes an intensive parameter while the conjugate extensive quantity fluctuates in the subsystem.
For a reservoir,
Conservation across the system–reservoir boundary gives, for each exchanged extensive variable,
To first order in the subsystem exchanges,
Maximizing is therefore equivalent to minimizing the appropriate part of
Which terms appear depends on which exchanges are permitted:
| Environment and fixed controls | Fluctuating subsystem quantity | Equilibrium potential |
|---|---|---|
| isolated: fixed | internal redistribution | maximize |
| thermal: fixed | minimize | |
| thermal and mechanical: fixed | minimize | |
| thermal and particle: fixed | minimize |
The word “minimize” means minimize over states or macrostates compatible with the stated controls. It does not mean that the potential decreases under every arbitrary driven or nonequilibrium protocol.
Legendre Transforms
Section titled “Legendre Transforms”Suppose is differentiable. Replacing by its conjugate gives
evaluated where
The subtraction removes the old differential term and introduces the new term . A pressure transform has a plus sign because the variable conjugate to is :
Here denotes enthalpy. The notation prevents confusion with the Hamiltonian operator .
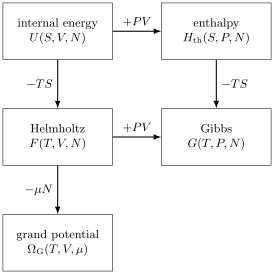
Each arrow replaces an extensive natural variable by its conjugate intensive variable. The sign of the added term follows the fundamental differential .
For the macroscopic equilibrium equation of state, it is often safer to formulate the transform variationally:
Likewise,
These are Legendre–Fenchel transforms of the thermodynamic functions. In particular, the transform over particle number is not an exact replacement for a finite grand-canonical sector sum; the section on exact transforms below makes the distinction explicit. When the relevant function is differentiable and has the required convexity, the infimum occurs at the familiar stationary condition and the construction reduces to the ordinary Legendre transform. At a first-order transition, a range of coexisting states can share the same supporting tangent, so derivatives may be discontinuous or set-valued.
The potential family
Section titled “The potential family”Each differential can be checked by differentiating the definition and substituting . For example, cancels the term and makes a natural variable.
| Potential | Definition | Differential |
|---|---|---|
| Internal energy | ||
| Enthalpy | ||
| Helmholtz free energy | ||
| Gibbs free energy | ||
| Grand potential |
The variables being differentiated in the last column are the natural variables. Thus is a function of , while is a function of . Although both are called free energies, at fixed volume gives . A pressure-controlled ensemble also sums over volumes.
Natural Variables and Equations of State
Section titled “Natural Variables and Equations of State”Once a potential is known as a function of its natural variables, its first derivatives recover the conjugate observables.
For ,
For ,
For ,
Every derivative must state what is held fixed. For example, differentiating with respect to at fixed is not the same operation as differentiating at fixed fugacity .
Quantum partition functions
Section titled “Quantum partition functions”The canonical ensemble gives with a trace over the fixed-number space. The grand-canonical ensemble gives with a sum over number sectors. Their logarithms are dimensionless Massieu functions; partition functions develops their cumulants.
Fixed-pressure partition function
Section titled “Fixed-pressure partition function”A formal isothermal–isobaric partition function for fixed is
where is a reference volume required to make the integral dimensionless. With this convention,
and
Changing to adds to and shifts its temperature-derived entropy by . It leaves pressure-derived unchanged when is constant. For a macroscopic system, the convention is subextensive and the saddle point yields the thermodynamic Gibbs free energy. For confined quantum systems, changing also changes the Hilbert space or boundary conditions, so the family must be specified.
Free-energy variational principles
Section titled “Free-energy variational principles”The state variational principles are exact even for a finite system. If is the Gibbs state on the fixed-number space, then
In the grand-canonical space the same identity reads
where . Nonnegativity of relative entropy makes the Gibbs state the unique minimizer when these quantities are finite. The first identity is proved in an exercise below. At a normalizable negative temperature the identities remain true, but the inequalities reverse.
The grand-state expression uses the entropy of its mixture over number sectors. It is generally not the canonical evaluated at the possibly noninteger mean . This distinction explains why the exact state minimum and the approximate macroscopic sector minimum are compatible.
Pressure
Section titled “Pressure”Pressure is the generalized force conjugate to volume. In the canonical representation,
If the Hamiltonian depends smoothly on a geometric parameter , the parameter-derivative identity gives
so
This compact formula hides geometric choices. One must specify how the boundary, metric, couplings, and ultraviolet regulator change with . In a lattice system, for example, changing the number of sites is not an infinitesimal deformation of a fixed Hilbert space.
In the grand-canonical representation,
For a homogeneous extensive phase in the thermodynamic limit,
Surface terms, traps, interfaces, long-range interactions, and finite-size corrections can invalidate the simple identity even though the derivative definition remains meaningful.
Finite-number derivatives
Section titled “Finite-number derivatives”In macroscopic thermodynamics . In a finite system, compare exact sectors through and . These can differ. The interval between them tests stability against neighboring sectors; global stability additionally requires comparison with every allowed sector, or discrete convexity. See chemical potential for the full condition and a counterexample.
Extensivity and Euler Relation
Section titled “Extensivity and Euler Relation”Suppose the internal energy is first-order homogeneous in its extensive variables:
Euler’s theorem then gives
Consequently,
These are Euler relations for a homogeneous, one-component, extensive phase. They are not definitions valid for every finite or inhomogeneous system.
For several conserved species,
and extensivity gives
Therefore
Gibbs–Duhem Relation
Section titled “Gibbs–Duhem Relation”Differentiate the Euler relation:
Subtracting the fundamental relation leaves
This is the Gibbs–Duhem relation. It shows that and are not independent for a homogeneous one-component equilibrium phase.
Writing entropy and volume per particle as
gives
For multiple species,
Maxwell Relations
Section titled “Maxwell Relations”Mixed partial derivatives of a smooth thermodynamic potential commute. From
one obtains
and
From ,
From ,
and
Maxwell relations are consistency conditions and computational tools. They are valid only where the potential is differentiable and the appropriate remaining natural variables are held fixed on each side. The differentiation variable itself changes sides; for example, .
Curvature and Stability
Section titled “Curvature and Stability”Thermodynamic stability constrains second derivatives. At fixed ,
where
Thus is concave in temperature for a stable canonical system.
The isothermal compressibility is
For a smooth mechanically stable branch with ,
Particle-number stability similarly requires
in a stable single phase.
Grand-canonically,
The final equality assumes . It links thermodynamic concavity to a nonnegative quantum fluctuation. More general sources require the Kubo–Mori treatment developed in Fluctuations and Susceptibilities.
Divergent compressibility or constrained zero response needs a limiting treatment rather than literal division by . With several coupled extensive variables, nonnegative diagonal curvatures alone are not sufficient: the full stability Hessian must be positive semidefinite. A potential can be concave in some natural variables and convex in others. Saying simply that “free energy is convex” is incomplete unless the variables are named.
Phase Coexistence
Section titled “Phase Coexistence”For a homogeneous one-component phase,
Two phases and coexist when their chemical potentials are equal at the same and :
For each phase,
Differentiating the coexistence condition along the coexistence curve gives the Clapeyron equation:
where is the latent heat per particle. The relation fails at points where the simple two-phase description or the required derivatives cease to apply.
At finite size, partition functions are typically analytic and the transition is rounded. Nondifferentiable thermodynamic potentials emerge only after an appropriate large-system limit. Finite-Temperature Phase Transitions develops this nonanalytic limit, the distinction between first-order and continuous transitions, and the finite-size signatures of each.
Example: A Spin in a Magnetic Field
Section titled “Example: A Spin in a Magnetic Field”Consider a two-level magnetic moment with
The energies are and , so
The Helmholtz free energy is
The magnetic moment conjugate to is
The internal energy is
and the entropy is
At , both levels are equally populated and . At fixed nonzero , the limit selects the lower level and .
The isothermal susceptibility is a curvature of the free energy:
This is the generalized-coordinate version of the relations between compressibility, number susceptibility, and potential curvature.
Finite Systems and Exact Transforms
Section titled “Finite Systems and Exact Transforms”Thermodynamic Legendre transforms often arise as large-system limits of exact Laplace transforms.
The grand partition function is exactly
Therefore
For a macroscopic system, the logarithm is often dominated by a narrow range of , giving
up to subextensive corrections. At finite size, the log-sum-exp need not equal the minimum. If there are allowed sectors and , then
Thus a subexponential number of sectors contributes at most a subextensive correction. For infinitely many sectors a separate bound on their convergent weight sum is needed. Analyticity requires a finite-dimensional trace or adequate uniform convergence, not merely the label “finite system.”
Similarly, using a density of states ,
becomes, when has exponential thermodynamic scaling,
The saddle point then gives
Here has units of inverse energy; the schematic exponential omits a fixed reference-energy factor, as explained in the canonical density-of-states calculation. If the entropy is not concave, transforming to the canonical potential and back recovers only its concave envelope. This is one route to ensemble nonequivalence in systems with long-range interactions or other failures of ordinary additivity. Such statements concern thermodynamic limits; they should not be inferred from a finite analytic partition function alone. See ensemble equivalence for supporting-line and large-deviation criteria.
Generalized Work Coordinates
Section titled “Generalized Work Coordinates”Let the Hamiltonian depend on a control parameter :
For a canonical Gibbs state,
If the generalized force is defined by
then
Examples include magnetization conjugate to magnetic field, polarization conjugate to electric field, and stress conjugate to strain. Electromagnetic and elastic work conventions differ across subfields, so the Hamiltonian coupling should determine the sign.
Exercises
Section titled “Exercises”Reconstruct the potential differentials
Section titled “Reconstruct the potential differentials”Starting from
derive the differentials of , , , and . Identify the natural variables of each potential.
Solution
For enthalpy,
so
Its natural variables are .
For Helmholtz free energy,
and therefore
Its natural variables are .
For Gibbs free energy,
which gives
Its natural variables are .
Finally,
so
Its natural variables are .
Prove the canonical variational identity
Section titled “Prove the canonical variational identity”Let
Show that
Solution
The Gibbs state satisfies
Hence
Multiply by and use
and
This gives
Since relative entropy is nonnegative, minimizes the free-energy functional.
Maxwell relation and thermal expansion
Section titled “Maxwell relation and thermal expansion”Use to prove
Express the right-hand side using the thermal expansion coefficient
Solution
From
we have
Commuting the mixed derivatives gives
Therefore
A material that expands on heating at fixed pressure has entropy that decreases with increasing pressure at fixed temperature.
Finite-sector transform
Section titled “Finite-sector transform”Suppose only the sectors and are available, with
Compute , , and . Compare the exact grand potential with the Legendre minimum of .
Solution
The sector sum is
Therefore
The mean occupation is
The two values of are
Their minimum is
At finite positive temperature, both weights are positive, so the exact lies strictly below this minimum. At the crossing it differs by , the mixing-entropy contribution. In the limit ,
while approaches a step function, with a degeneracy at .
References
Section titled “References”- R. Balian, From Microphysics to Macrophysics, Volume I, Springer (1991), chapters 4–7.
- H. B. Callen, Thermodynamics and an Introduction to Thermostatistics, 2nd ed., Wiley (1985), chapters 1–8.
- T. L. Hill, Thermodynamics of Small Systems, Dover (1994), parts I and II.
- M. Kardar, Statistical Physics of Particles, Cambridge University Press (2007), chapters 1, 4, and 6.
- L. D. Landau and E. M. Lifshitz, Statistical Physics, Part 1, 3rd ed., Butterworth–Heinemann (1980), sections 13–24 and 31–36.
- R. K. Pathria and P. D. Beale, Statistical Mechanics, 4th ed., Elsevier (2021), chapters 2–5.
- H. Touchette, “The Large Deviation Approach to Statistical Mechanics”, Physics Reports 478, 1–69 (2009).