

Hartree–Fock Approximation
The Hartree–Fock approximation is the variationally optimal single-Slater-determinant description of a fermionic ground state for a specified Hamiltonian and one-particle space. It replaces the interacting many-fermion eigenproblem by self-consistent one-body equations containing a direct field and a nonlocal exchange field.
For orthonormal occupied spin-orbitals , the trial state is the determinant
The approximation lies in restricting the many-body state to one determinant. Fermionic antisymmetry is not approximated: it is exact throughout the trial family. As a result, Hartree–Fock contains exchange, cancels one-particle self-interaction within the determinant, and produces an exchange hole. It does not contain the remaining dynamical or multireference correlation associated with superpositions or deformations beyond a single occupied subspace.
The defining equations are nonlinear because the Fock operator depends on the orbitals that solve it:
where is the occupied one-body projector. A self-consistent solution is an occupied subspace invariant under its own Fock operator.
Helpful background. Slater determinants supplies antisymmetry and normalization. The Hartree approximation derives the direct field; atomic Hartree–Fock develops radial applications.
Fermionic pair interactions and a determinant
Section titled “Fermionic pair interactions and a determinant”Consider identical fermions with Hamiltonian
The one-body operator may contain kinetic energy, external potentials, spin couplings, or lattice hopping. The pair interaction is symmetric under particle exchange,
Write a complete one-particle coordinate as when position and spin are both present. For the coordinate derivation, take to be local in position and independent of spin unless stated otherwise.
The same Hamiltonian can be written in an orthonormal spin-orbital basis as
where denotes antisymmetrized two-body matrix elements. The factor is in this convention; using non-antisymmetrized matrix elements instead gives a factor . Conventions must not be mixed.
Single-Determinant Trial Family
Section titled “Single-Determinant Trial Family”Choose orthonormal spin-orbitals,
Their determinant is an admissible fermionic state. The Hartree–Fock ground-state energy is the restricted minimum
The exact variational principle gives
This bound refers to the same Hamiltonian, boundary conditions, symmetry sector when one is imposed, and one-particle space. In a nested sequence of finite bases , the corresponding minima satisfy
A converged stationary determinant need not be the global minimum. It can be a local minimum, a saddle, or an excited self-consistent solution.
Direct and Exchange Integrals
Section titled “Direct and Exchange Integrals”Define the one-body matrix element
For two occupied spin-orbitals, define the direct integral
and the exchange integral
In compact two-particle notation,
The determinant expectation value is
The direct contribution is the interaction of one orbital density with another. Exchange arises from the crossed contraction required by antisymmetry. It has no separate classical pair-potential interpretation.
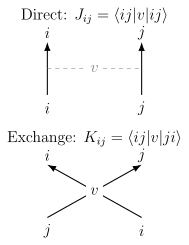
The two terms in a determinant pair expectation are . Direct and exchange differ by which ket orbitals connect to the two bra coordinates; the lines show index contractions, not literal particle trajectories. Their relative minus sign comes from fermionic antisymmetry.
Same-spin selection
Section titled “Same-spin selection”If the spin-orbitals factor as
with , then
for a spin-independent interaction. The direct integral can remain nonzero. Exchange therefore acts only between orbitals with overlapping spin components; in a collinear basis this means equal-spin pairs.
Self-interaction cancellation
Section titled “Self-interaction cancellation”For every occupied spin-orbital,
Hence its diagonal contribution cancels exactly:
Hartree–Fock does not make a fermion repel itself. This cancellation is exact for a one-particle system and for each occupied spin-orbital within the determinant. It does not mean that all many-electron self-interaction questions in other approximate frameworks are solved by the same mechanism.
Direct and Exchange Operators
Section titled “Direct and Exchange Operators”The direct operator generated by occupied orbital acts on a test spin-orbital as
For a coordinate-local pair interaction, is a multiplicative potential. By contrast, the exchange operator is
It is nonlocal: the value at depends on the test function at all . The matrix elements satisfy
and
The Fock operator is
Calling an effective one-particle Hamiltonian is useful, but it remains state dependent. Its eigenvalue problem is not equivalent to the original linear many-body Schrödinger equation.
Constrained Orbital Variation
Section titled “Constrained Orbital Variation”Minimize the determinant energy while enforcing all occupied-orbital overlaps. Introduce a Hermitian matrix of Lagrange multipliers :
Varying with respect to gives
The matrix, rather than scalar, multiplier is essential because occupied orbitals must remain mutually orthonormal. The equations state that maps the occupied space into itself.
Since is Hermitian, a unitary rotation among occupied orbitals diagonalizes it. In that canonical orbital basis,
One may complete the occupied set with virtual eigenfunctions of the same Fock operator,
Only the occupied subspace defines the determinant. Canonical virtual orbitals are a convenient completion, not occupied physical particles.
The detailed particle-hole matrix-element statement of stationarity is the Brillouin condition developed in Normal Ordering in Many-Body QM.
Occupied-Subspace Invariance
Section titled “Occupied-Subspace Invariance”Let
be the occupied one-body projector. If the occupied orbitals are rotated by a unitary matrix,
then
The determinant changes only by the phase , so all observables remain unchanged. The direct and exchange fields also remain unchanged because they depend on , not on a particular labeled set of occupied orbitals.
At a stationary point, the occupied space is invariant under . In projector form,
Equivalently, for a Hermitian Fock operator,
Thus the occupied subspace is invariant under the Fock operator built from it. Within a degenerate eigenspace, the projector may select any subspace; it need not include the entire degenerate eigenspace. It also explains why rotating localized occupied orbitals into canonical occupied orbitals does not change the Hartree–Fock energy.
One-Body Density-Matrix Formulation
Section titled “One-Body Density-Matrix Formulation”Take , with the creation coordinate second. The determinant kernel is
It obeys
Thus a zero-temperature number-conserving Slater determinant corresponds to a rank- orthogonal projector. The density is
For a local pair interaction, the energy functional becomes
The direct term depends only on the diagonal density. Exchange depends on the off-diagonal coherence of the one-body density matrix.
The corresponding Fock kernel is
This formula makes the local/nonlocal distinction explicit. Lattice coordinates replace the integrals by sums. A genuinely nonlocal two-body interaction requires its full four-index contraction, rather than inserting it unchanged into this two-coordinate formula.
Pair Density and the Exchange Hole
Section titled “Pair Density and the Exchange Hole”For a determinant, the ordered pair density factorizes as
The second term is the exchange deficit. For , define the exchange hole around a fermion at by
Projector idempotency implies
Therefore
Conditioning on one spin-coordinate removes one particle’s worth of density from the exchange neighborhood. For collinear spin orbitals the deficit is entirely in that spin component. The hole is a consequence of antisymmetry, not a literal repulsive force. Opposite-spin pairs have no exchange hole in a collinear spin basis, although interactions can generate a correlation hole beyond Hartree–Fock.
Orbital Energies and Total Energy
Section titled “Orbital Energies and Total Energy”For canonical occupied orbitals,
Summing occupied orbital energies counts each pair contribution twice:
Hence
Equivalently,
For molecular electronic structure, the classical nucleus-nucleus repulsion must be added separately to obtain the Born–Oppenheimer total energy.
Orbital eigenvalues are useful diagnostics and zeroth-order quantities, but the Hartree–Fock variational theorem applies to the total determinant energy, not separately to every .
Self-Consistent-Field Logic
Section titled “Self-Consistent-Field Logic”A conceptual Hartree–Fock iteration is:
- choose an initial rank- occupied projector ;
- build and ;
- form ;
- solve the one-body eigenproblem;
- construct a new occupied projector from a selected -dimensional eigenspace;
- repeat until the projector and orbital residuals are stationary.
For an orbital , define the residual
In projector form, a basis-independent occupied-virtual residual is
Energy changes can become small before this residual does. A robust calculation monitors the commutator , density or projector changes, orthonormality, and the energy reconstructed from both integrals and orbital eigenvalues.
Practical algorithms use damping, level shifts, extrapolation, direct inversion of the iterative subspace, fractional occupations, or trust-region orbital optimization. Fractional occupations may be an intermediate solver device, but a final zero-temperature single determinant must recover an idempotent rank- projector. Keeping fractional occupations defines an ensemble extension, and finite-temperature Hartree–Fock minimizes a free energy rather than the pure-state functional.
Molecular Electronic Hamiltonian
Section titled “Molecular Electronic Hamiltonian”After fixing nuclei in the Born–Oppenheimer approximation and using atomic units, the electronic Hamiltonian is
The nuclear repulsion is
Hartree–Fock minimizes over a chosen determinant family. The potential-energy surface uses
Relativistic effects, finite nuclear mass, effective core potentials, basis incompleteness, and electron correlation are distinct approximations or corrections. They should not be folded into a single undifferentiated “Hartree–Fock error.”
Closed-Shell Restricted Hartree–Fock
Section titled “Closed-Shell Restricted Hartree–Fock”For an even-electron closed shell, restricted Hartree–Fock uses the same spatial orbital for one and one spin-orbital. If spatial orbitals are doubly occupied, the spatial Fock operator is
The electronic energy is
The factor of two in the direct term counts both spin partners. Exchange remains only between equal-spin components, producing one spatial exchange term after spin summation.
RHF preserves the closed-shell spin symmetry and is often a useful molecular reference near equilibrium. Its restricted trial family can become qualitatively inadequate when bonds stretch, shells become nearly degenerate, or open-shell character develops.
Unrestricted and Generalized Forms
Section titled “Unrestricted and Generalized Forms”Unrestricted Hartree–Fock
Section titled “Unrestricted Hartree–Fock”Unrestricted Hartree–Fock allows different spatial orbitals for and electrons while preserving a fixed . Its variational space contains the RHF space, so
for the same basis and Hamiltonian. The lower energy may represent useful spin polarization, or it may arise through a determinant that is not an eigenstate of . The latter effect is commonly called spin contamination.
Restricted open-shell Hartree–Fock
Section titled “Restricted open-shell Hartree–Fock”Restricted open-shell constructions share spatial orbitals in selected closed shells while treating unpaired electrons in spin-adapted occupations. They preserve more spin structure but introduce choices in how open-shell Fock operators and canonical orbital energies are defined. The determinant energy and occupied space remain more fundamental than a particular canonicalization.
Generalized Hartree–Fock
Section titled “Generalized Hartree–Fock”Generalized Hartree–Fock allows each spin-orbital to mix spin components. It can describe noncollinear magnetism and broader symmetry breaking within a number-conserving determinant. Allowing anomalous particle-number-breaking densities leads further to Hartree–Fock–Bogoliubov or Bogoliubov–de Gennes families, which belong with pairing theory rather than ordinary Hartree–Fock.
Roothaan–Hall Matrix Equation
Section titled “Roothaan–Hall Matrix Equation”Expand molecular spin-orbitals or spatial orbitals in a finite nonorthogonal basis :
Define the overlap and Fock matrices
The canonical orbital equations become the generalized eigenproblem
The matrix depends on the occupied coefficients through the density matrix and two-electron integrals, so this is still a nonlinear self-consistency problem. Orthonormalizing the basis converts it to an ordinary Hermitian eigenproblem, provided is positive definite on the retained basis subspace.
This equation is the bridge from the continuum variational derivation to molecular calculations. Basis construction, integral evaluation, screening, convergence acceleration, and analytic derivatives belong to computational treatments.
Two-Electron Closed-Shell Example
Section titled “Two-Electron Closed-Shell Example”Let two electrons occupy the same normalized spatial orbital with opposite spins:
The self terms cancel, and opposite-spin exchange vanishes. The energy is
Thus antisymmetry does not remove the Coulomb repulsion between the two opposite-spin electrons. Improving the shared orbital accounts for average screening, as in the elementary helium variational estimate, but the single determinant still lacks explicit dependence on and cannot satisfy the full electron-electron cusp structure.
Bond Dissociation and Broken Symmetry
Section titled “Bond Dissociation and Broken Symmetry”The hydrogen molecule exposes the distinction between a low variational energy and a symmetry-faithful state. At large internuclear separation, the RHF bonding orbital is approximately
where and are orbitals localized on the two atoms. The closed-shell spatial factor expands as
The ionic terms and retain incorrect weight when the atoms separate. UHF can lower the energy by localizing opposite spins on opposite atoms, but a single localized determinant is not a pure total-spin singlet. The exact separated singlet requires a symmetry-adapted superposition of determinants.
This is static correlation or multireference failure: several determinants become comparably important. More elaborate iteration of one determinant cannot repair the missing state-space dimension. Valence Bond Theory expresses the same dissociation problem in localized covalent and ionic structures and explains how a spin-adapted structure expansion restores the required state space.
Uniform Electron Gas
Section titled “Uniform Electron Gas”For a homogeneous spin- electron gas in volume , translational symmetry makes plane-wave spin-orbitals natural:
At zero temperature, the unpolarized determinant fills for both spins, with
In jellium, the uniform electronic Hartree term is canceled by the positive background and its background self-energy. The remaining Hartree–Fock correction is exchange.
For the three-dimensional unpolarized gas, the exchange energy per particle is
Equivalently,
Thus exchange scales as per particle and as per volume. If
is the spin polarization, then
Because , exchange favors spin polarization, while the kinetic energy opposes it. Comparing homogeneous polarized Fermi seas can therefore predict a change of preferred polarization at sufficiently low density. Correlation and screening strongly affect that competition, so the Hartree–Fock transition is not a quantitatively settled prediction for the physical electron liquid.
This calculation is restricted to homogeneous Fermi-sea determinants. It is not the unrestricted Hartree–Fock ground-state solution: Overhauser showed that spatially varying spin-density-wave determinants can lower the unpolarized sea’s energy, including at high density. A homogeneous polarization comparison alone misses that instability.
Unscreened Fock exchange also gives a nonanalytic slope in the single-particle dispersion at the Fermi surface. This pathology is a warning that static exchange alone does not describe metallic screening or the full quasiparticle spectrum.
Koopmans’ Frozen-Orbital Relation
Section titled “Koopmans’ Frozen-Orbital Relation”Remove an electron from occupied canonical orbital while freezing every remaining orbital. The energy difference is
Thus the frozen-orbital ionization energy is approximately
The equality is exact only for the frozen Hartree–Fock energy expressions. A physical ionization energy also contains orbital relaxation, electron correlation, possible symmetry changes, relativistic effects, and nuclear response depending on whether the process is vertical or adiabatic.
Virtual orbital energies provide an even less direct estimate of electron affinities because the added electron changes the self-consistent field and may not be bound in the -electron Fock potential. Koopmans’ relation is a useful organizing approximation, not a general identification of all Fock eigenvalues with exact addition and removal energies.
Correlation Beyond Hartree–Fock
Section titled “Correlation Beyond Hartree–Fock”For the same Hamiltonian and finite one-particle space, define
This quantum-chemistry convention excludes exchange because exchange is already present in the optimized determinant. Correlation methods recover effects of the residual interaction through determinant superpositions, perturbative excitations, coupled-cluster amplitudes, Green functions, tensor networks, quantum Monte Carlo, or other richer state families. Variational many-body states compares these trial families.
Two broad failure patterns are useful:
- dynamical correlation: many determinants have individually small amplitudes but collectively describe short-range avoidance and fluctuations;
- static correlation: a few determinants become nearly degenerate and no single reference dominates.
The numerical value of is not an entanglement measure, and a small correlation energy does not guarantee that every correlation-sensitive observable is accurate.
What It Misses
Section titled “What It Misses”Short-range correlation
Section titled “Short-range correlation”The pair density is fixed by the one-body projector. It cannot independently optimize opposite-spin avoidance or satisfy general interaction cusps.
Dispersion
Section titled “Dispersion”Separated neutral closed-shell fragments have no London dispersion interaction at ordinary Hartree–Fock level because correlated instantaneous dipole fluctuations require excitations beyond one determinant.
Static multireference structure
Section titled “Static multireference structure”Bond breaking, near-degenerate shells, frustrated magnets, and strongly localized electrons may require several determinants with comparable weights.
Screening and collective modes
Section titled “Screening and collective modes”Bare exchange is not dynamically screened. Random Phase Approximation develops the direct density-response bubble chain, screened interaction, and collective poles; its exchange-inclusive particle–hole variant is time-dependent Hartree–Fock. Neither construction supplies all short-range or multireference correlation.
Symmetry restoration
Section titled “Symmetry restoration”An unrestricted determinant may imitate correlation by breaking spin or spatial symmetry. The energy can improve while exact quantum numbers and finite-system superpositions are lost.
Exact spectra
Section titled “Exact spectra”Fock orbital energies are not generally exact excitation, addition, or removal energies. Total-energy differences and many-body Green-function poles answer different questions.
Exercises
Section titled “Exercises”Two-orbital determinant energy
Section titled “Two-orbital determinant energy”For two orthonormal spin-orbitals and , evaluate the expectation value of
in their normalized determinant.
Solution
The determinant is
Orthonormality removes one-body cross terms, giving . Expanding the two-body expectation yields two direct terms and two crossed terms. The normalization factor gives
Therefore
Electron-gas exchange scaling
Section titled “Electron-gas exchange scaling”Use to determine how the three-dimensional exchange energy per particle and exchange energy density scale with . What happens to the magnitude under full spin polarization?
Solution
Since
the exchange energy per particle scales as . Multiplication by particle density gives
For , the polarization factor is
The exchange energy is therefore more negative by a factor than in the unpolarized gas at the same total density. The kinetic energy also rises, so this result alone does not decide the stable polarization.
Derive the frozen-orbital removal energy
Section titled “Derive the frozen-orbital removal energy”Remove occupied canonical orbital while leaving every other orbital fixed. Show that the Hartree–Fock energy difference is .
Solution
Removing subtracts its one-body contribution and every pair containing it:
The self term is zero because . The canonical orbital energy is
so
Allowing the remaining orbitals to relax changes the result and moves beyond the frozen-orbital statement.
When the exchange-hole integral changes
Section titled “When the exchange-hole integral changes”A number-conserving Gaussian mixed state has one-body eigenvalues . Does its exchange deficit still integrate to ?
Solution
Wick factorization still gives the deficit , but . Its integral is
which generally lies between and , rather than equaling . The determinant sum rule uses exact occupations zero or one. A fixed-number thermal state generally has additional correlations and is not the unrestricted Gaussian ensemble used in this calculation.
References
Section titled “References”- V. Bach, E. H. Lieb, and J. P. Solovej, “Generalized Hartree–Fock Theory and the Hubbard Model,” Journal of Statistical Physics 76, 3–89 (1994), doi:10.1007/BF02188656.
- P. A. M. Dirac, “Note on Exchange Phenomena in the Thomas Atom,” Proceedings of the Cambridge Philosophical Society 26, 376–385 (1930), doi:10.1017/S0305004100016108.
- A. L. Fetter and J. D. Walecka, Quantum Theory of Many-Particle Systems, Dover (2003).
- V. Fock, “Näherungsmethode zur Lösung des quantenmechanischen Mehrkörperproblems,” Zeitschrift für Physik 61, 126–148 (1930), doi:10.1007/BF01340294.
- G. F. Giuliani and G. Vignale, Quantum Theory of the Electron Liquid, Cambridge University Press (2005), doi:10.1017/CBO9780511619915.
- T. Koopmans, “Über die Zuordnung von Wellenfunktionen und Eigenwerten zu den Einzelnen Elektronen Eines Atoms,” Physica 1, 104–113 (1934), doi:10.1016/S0031-8914(34)90011-2.
- E. H. Lieb and B. Simon, “The Hartree–Fock Theory for Coulomb Systems,” Communications in Mathematical Physics 53, 185–194 (1977), doi:10.1007/BF01609845.
- R. McWeeny, Methods of Molecular Quantum Mechanics, 2nd ed., Academic Press (1992).
- A. W. Overhauser, “Spin Density Waves in an Electron Gas,” Physical Review 128, 1437–1452 (1962), doi:10.1103/PhysRev.128.1437.
- J. A. Pople and R. K. Nesbet, “Self-Consistent Orbitals for Radicals,” Journal of Chemical Physics 22, 571–572 (1954), doi:10.1063/1.1740120.
- P. Ring and P. Schuck, The Nuclear Many-Body Problem, Springer (1980), doi:10.1007/978-3-642-61852-9.
- C. C. J. Roothaan, “New Developments in Molecular Orbital Theory,” Reviews of Modern Physics 23, 69–89 (1951), doi:10.1103/RevModPhys.23.69.
- J. C. Slater, “Note on Hartree’s Method,” Physical Review 35, 210–211 (1930), doi:10.1103/PhysRev.35.210.2.
- A. Szabo and N. S. Ostlund, Modern Quantum Chemistry: Introduction to Advanced Electronic Structure Theory, Dover (1996).