

Central-Field Approximation
The central-field approximation replaces the coupled motion of many atomic electrons by one-electron motion in an effective spherically symmetric potential. The potential contains the nuclear attraction and an averaged description of the other electrons. Its eigenfunctions provide orbitals, subshells, and a reference configuration; residual interactions then refine or mix that reference.
Required background. Hydrogenic states supply the radial and angular labels; Pauli exclusion and Slater determinants explain how orbitals form fermionic states.
The Many-Electron Problem
Section titled “The Many-Electron Problem”For nonrelativistic electrons and a point nucleus of charge , the fixed-nucleus Hamiltonian in Hartree atomic units is
The nuclear term is central, but the electron–electron term depends on the angle between and as well as their radii. It couples electron coordinates and prevents the wavefunction from factorizing into exact independent orbitals.
The central-field strategy chooses a reference Hamiltonian
with
The difference
contains whatever two-electron interaction and one-electron counterterms were not absorbed into . Calling “small” is a hypothesis to test, not a consequence of writing this decomposition.
What Central Means
Section titled “What Central Means”A local central potential depends only on . Therefore
One-electron orbitals can be written
where is a spin function. For a bound orbital, impose , decay at infinity, and . The radial orbital satisfies
Spherical symmetry guarantees the degeneracy in before external fields or nonspherical interactions are included. Unlike the pure Coulomb problem, a generic central potential does not make different values at the same principal label degenerate. The orbital energy normally depends on both and .
A central field is a model, not a literal force from a frozen cloud
Section titled “A central field is a model, not a literal force from a frozen cloud”The phrase “each electron feels the average field of the others” is useful but incomplete. In a self-consistent theory, the orbitals create a density, the density creates an effective operator, and that operator changes the orbitals. For fermions, exchange is generally nonlocal; correlation is not captured by a simple average density. A fitted local potential may reproduce selected energies without being the Hartree field of any unique density.
Screening and Effective Charge
Section titled “Screening and Effective Charge”Write a local effective potential schematically as
The screening term is positive for the direct electrostatic repulsion from other electrons. It weakens the nuclear attraction experienced by an electron, but it is not generally equivalent to replacing by one constant.
For a spherical number density of other electrons, normalized to their number, the direct Hartree potential in atomic units is
Charge inside radius contributes as if concentrated at the origin; charge outside contributes a spatially constant potential within each spherical shell. The result follows from the spherical average of the Coulomb kernel, not from assuming every electron occupies a sharp shell.
Near and far from the nucleus
Section titled “Near and far from the nucleus”At small , the singular nuclear attraction dominates:
Far outside all other electrons, an electron sees the net charge of the nucleus plus the remaining electrons. If the ion has charge , then
For a neutral atom, , so a distant valence electron sees an asymptotic tail. For a singly positive ion, the tail is . This is the physical active-electron tail, and the tail of a self-excluded direct field. A direct potential constructed from all electrons instead tends to unless its self-interaction is canceled. Thus an arbitrary mean-field or approximate density-functional potential need not have the correct Rydberg tail. Nonlocal exchange cannot in general be summarized by one local function .
When the potential is local, one may define a radius-dependent effective charge by
It approaches near the nucleus and far away under the ideal limits above. A single tabulated “effective nuclear charge” is therefore an orbital-dependent approximation to a function, not an exact property of the atom.
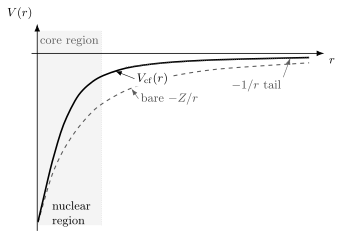
Schematic local potentials for a neutral atom. Near the nucleus, both retain the singularity. Through the core region, electron screening weakens the attraction, and the effective valence potential approaches at large . A real self-consistent or fitted potential need not follow the particular interpolation drawn here.
Penetration and Orbital-Dependent Screening
Section titled “Penetration and Orbital-Dependent Screening”Orbitals with different sample the core differently. The centrifugal term
suppresses low-radius amplitude increasingly as grows. At comparable principal excitation:
- orbitals penetrate the core most strongly;
- orbitals generally penetrate less;
- and orbitals are often concentrated farther outside the core;
- stronger penetration exposes an electron to a larger effective nuclear charge and usually increases its binding.
This ordering is qualitative, not a universal theorem about every level. Radial nodes, core relaxation, relativistic contraction, near-degeneracy, and configuration mixing can change simple expectations.
Screening is not shielding by rigid shells
Section titled “Screening is not shielding by rigid shells”Electron density is continuous and orbitals overlap. Inner electrons do not create a perfectly opaque barrier, and outer electrons still contribute to self-consistent fields. Shell language organizes the density and angular momenta; it should not be converted into a classical picture of nested charged surfaces.
Shells, Subshells, and Configurations
Section titled “Shells, Subshells, and Configurations”The central-field orbitals are labeled . Pauli exclusion allows at most one electron per spin-orbital. A spatial subshell of given therefore has capacity
| Subshell | Spatial orbitals | Maximum electrons | |
|---|---|---|---|
An electron configuration records occupation of these subshells, for example . It is a basis-level description of a reference state. An exact many-electron eigenstate can be a superposition of configurations, and open-shell levels require coupling the individual orbital and spin angular momenta into total , , and .
See Electron Configurations for occupation notation and the limits of Aufbau ordering.
Orbital energies are not the many-electron spectrum
Section titled “Orbital energies are not the many-electron spectrum”The eigenvalue belongs to an effective one-electron equation. The total energy includes interaction double counting and depends on the chosen approximation. Excitation energies are differences of many-electron state energies, not generally differences of frozen central-field orbital eigenvalues. Koopmans-type relations add assumptions such as a single determinant and frozen relaxation; they are not exact identities for measured spectra.
Quantum Defects
Section titled “Quantum Defects”A valence or Rydberg electron in a neutral alkali atom sees an asymptotic potential but departs from a pure Coulomb potential in the core. A Rydberg series is often represented by
where is the residual core charge, is the reduced-mass Rydberg constant in inverse-length units, and
is the effective principal quantum number. The quantum defect summarizes the short-range phase accumulated when the electron penetrates and polarizes the core.
Low- series usually have larger defects because they penetrate more strongly. High- Rydberg states remain mostly outside the core and become nearly hydrogenic. Fine structure can make the defect dependent.
What a quantum defect is not
Section titled “What a quantum defect is not”A defect is not a missing fraction of an electron, a literal change in , or a universal constant of an element. It can depend on energy, isotope, ionization threshold, channel coupling, and the fitting convention. Near perturbing configurations, a constant-defect model may fail and multichannel quantum-defect methods or explicit configuration mixing may be required.
The NIST atomic-spectroscopy compendium gives the evaluated spectroscopic convention for term series and quantum defects.
Rydberg Atoms Basics uses to organize the large-orbit, polarizability, lifetime, and interaction scalings that follow from this effective one-electron picture.
How Central Fields Are Constructed
Section titled “How Central Fields Are Constructed”Different constructions answer different questions.
| Construction | Effective operator | What it includes | Characteristic limitation |
|---|---|---|---|
| screened Coulomb model | chosen local | qualitative penetration and shell ordering | fitted parameters may not transfer between observables |
| Hartree | local direct field | self-consistent density response | raw orbital product lacks antisymmetry; no exchange, even when the direct field excludes self-interaction |
| Hartree–Fock | direct plus nonlocal exchange | determinant antisymmetry and exchange | no correlation beyond one determinant |
| central-field Hartree–Fock | spherical average of open shells | atomic subshell structure with exchange | nonspherical and configuration effects are averaged |
| local density-functional model | local effective potential | exchange-correlation approximation through the density | orbital energies and functional errors require interpretation |
| model potential fitted to data | local or nonlocal effective operator | selected observed energies or phase shifts | may lack a unique variational or microscopic meaning |
The variational derivations are given in Hartree Approximation and Hartree–Fock Approximation. Hartree–Fock for Atoms develops the spherical closed-shell reduction, open-shell averages, and radial exchange multipoles. A local central potential cannot exactly reproduce a general nonlocal exchange operator for every orbital simultaneously.
A self-consistent calculation alternates between constructing the field from occupied orbitals and solving its eigenvalue problem; mixing successive fields can stabilize iteration. Convergence of energy, density, and equation residuals establishes a numerical fixed point. Testing other occupations and larger radial bases is still needed to distinguish a stable solution from a converged but inaccurate approximation. Hartree Method gives the radial iteration explicitly.
Residual Interactions and Configuration Mixing
Section titled “Residual Interactions and Configuration Mixing”Once is chosen, the residual operator can be treated perturbatively or by diagonalization in a configuration basis. Schematically,
where are antisymmetrized configuration-state functions with the required total symmetry. The coefficients encode mixing that a single configuration cannot represent.
Residual electrostatic interactions split terms, spin–orbit interactions mix or split angular-momentum couplings, and near-degenerate configurations can invalidate low-order perturbation theory. The central field remains useful as a basis generator even when it is not itself an accurate final-state description.
Assessing the approximation
Section titled “Assessing the approximation”The central field is a useful starting point when a compact core and a dominant configuration describe the states of interest. Near-degenerate configurations, open-shell multipoles, core excitation, and continuum coupling can instead require a multiconfiguration treatment. An accurate energy is not sufficient evidence for an accurate transition amplitude: matrix elements may be sensitive to small admixtures, radial tails, or cancellation between contributions.
Test the quantity the model is intended to predict. For spectroscopy, compare ionization energies and term intervals; for dipole transitions, compare lifetimes and branching ratios; for response, compare polarizabilities and oscillator-strength sums. Basis and radial-box convergence must precede that comparison.
Length and velocity forms provide a useful consistency check only when the velocity operator matches the Hamiltonian:
For a local nonrelativistic potential, . A nonlocal exchange or model operator contributes an additional commutator, so comparing a length-form result with an uncorrected matrix element need not test the same model. Even agreement of consistent forms does not prove that every individual line is accurate.
Validate the tail separately
Section titled “Validate the tail separately”Weakly bound states, Rydberg series, polarizabilities, tunneling rates, and threshold scattering are especially sensitive to the long-range potential. A model fitted to compact low-lying orbitals may have the wrong asymptotic charge or omit the polarization tail
where is the static dipole polarizability of the residual core in atomic units. This term is weaker than the Coulomb tail but can matter once the leading Coulomb behavior is fixed. It must not be inserted twice if core polarization is already represented by the underlying many-body method or fitted potential.
Exercises
Section titled “Exercises”1. Asymptotic charge seen by an outer electron
Section titled “1. Asymptotic charge seen by an outer electron”An atom or ion has nuclear charge and electrons. Let be its net charge in units of . Place one electron far outside the other electrons. Show that its leading potential energy is
What does this become for a neutral atom?
Solution
At distances much larger than the core, only its total charge enters the monopole field. The nucleus contributes , while the other electrons contribute . The residual core charge is therefore
The active electron has negative charge, so its potential energy is . For a neutral atom, and the tail is : far away, the electron sees the singly charged ion left after its removal. Multipole and polarization terms fall faster with .
2. A continuous screening cloud
Section titled “2. A continuous screening cloud”Let the other electrons have uniform spherical number density inside radius . Derive inside and outside that sphere, and check continuity of the potential and its derivative at .
Solution
The density is . Substitution in the spherical integral gives
Both expressions give and derivative at . Inside the cloud, screening is a constant plus a quadratic term, not another Coulomb singularity. Adding therefore preserves the nuclear singularity while producing the correct residual charge outside the cloud.
3. Penetration from quantum defects
Section titled “3. Penetration from quantum defects”Two Rydberg series converge to the same threshold and share the same mass-corrected Rydberg constant. At , suppose an series has and a series has . Compute their effective principal quantum numbers and the ratio of their binding-energy magnitudes in the constant-defect model.
Solution
The effective principal quantum numbers are
Because ,
The larger -wave defect signals stronger penetration into the core and a substantially larger binding magnitude at the same nominal . The numerical defects are model inputs here; near perturbers, an energy-independent defect may not be adequate.
4. Which degeneracy survives?
Section titled “4. Which degeneracy survives?”Explain why a generic spin-independent central potential keeps the degeneracy for fixed but does not preserve the hydrogenic degeneracy between different values at fixed .
Solution
A central Hamiltonian commutes with and . Rotational invariance makes all orientations of a given orbital equivalent, so the energy is independent of . Different values, however, obey radial equations with different centrifugal terms,
and they penetrate a screened core differently. Rotational symmetry does not connect them. The pure Coulomb problem has an additional conserved Runge–Lenz vector and an enlarged dynamical symmetry, which produces the accidental degeneracy. A generic lacks that symmetry.
5. Accurate energies, poor oscillator strengths
Section titled “5. Accurate energies, poor oscillator strengths”A one-configuration central-field calculation reproduces several level energies to within one percent, but its electric-dipole oscillator strengths disagree strongly with measured branching ratios. Give at least three plausible reasons and outline a useful response.
Solution
Plausible causes include configuration mixing that changes transition amplitudes more than energies, core polarization, inconsistent treatment of the transition operator, incorrect radial tails, and cancellation between several amplitude contributions. In variational calculations, an energy can also be less sensitive to a small wavefunction error than a matrix element is.
A useful response is to inspect the dominant configuration-state amplitudes, compare length and velocity forms using a consistent Hamiltonian, enlarge the configuration space, add core-polarization or response corrections without double counting, and benchmark lifetimes or several related lines rather than one fitted transition. The appropriate remedy follows the observable-level diagnosis; refitting a scalar screening charge to the energies alone need not improve the amplitudes.
References
Section titled “References”- B. H. Bransden and C. J. Joachain, Physics of Atoms and Molecules, 2nd ed. (Pearson, 2003), Chapters 6–8.
- E. U. Condon and G. H. Shortley, The Theory of Atomic Spectra (Cambridge University Press, 1935), Chapters 6–9.
- R. D. Cowan, The Theory of Atomic Structure and Spectra (University of California Press, 1981), especially Chapters 3–8.
- V. Fock, “Näherungsmethode zur Lösung des quantenmechanischen Mehrkörperproblems,” Zeitschrift für Physik 61, 126–148 (1930), doi:10.1007/BF01340294.
- C. J. Foot, Atomic Physics (Oxford University Press, 2005), Chapters 3–5.
- D. R. Hartree, “The Wave Mechanics of an Atom with a Non-Coulomb Central Field. Part I. Theory and Methods,” Proceedings of the Cambridge Philosophical Society 24, 89–110 (1928), doi:10.1017/S0305004100011919.
- W. R. Johnson, Atomic Structure Theory: Lectures on Atomic Physics (Springer, 2007), doi:10.1007/978-3-540-68013-0.
- A. Kramida, Y. Ralchenko, J. Reader, and the NIST ASD Team, NIST Atomic Spectra Database, National Institute of Standards and Technology, doi:10.18434/T4W30F.
- National Institute of Standards and Technology, Atomic Spectroscopy: Term Series, Quantum Defects, and Spectral-Line Conventions.