

Atomic Orbitals
An atomic orbital is a one-electron wavefunction used to describe the spatial state of an electron bound to a nucleus. In the exact nonrelativistic hydrogenic problem, orbitals are stationary eigenfunctions of the point-Coulomb Hamiltonian. In many-electron atoms and molecules, the same word is also used for approximate one-electron functions inside a larger many-body description.
This page treats the clean hydrogenic meaning. It explains the notation, probability-density interpretation, real and complex orbital bases, nodal surfaces, and common visualization traps. Many-electron and chemical-orbital uses should be read as later applications of this one-electron prototype, not as part of the exact hydrogen solution.
Helpful background. Hydrogen Atom to use the Coulomb quantum numbers and bound-state energies.
Definition in the Hydrogenic Problem
Section titled “Definition in the Hydrogenic Problem”For a spinless one-electron Coulomb problem, the bound-state wavefunctions separate as
The quantum numbers obey
The radial factor is described in Radial Wavefunctions. The angular factor is a spherical harmonic. Together they define a spatial orbital.
The shell-level angular organization of these labels is summarized in Hydrogen Atom Angular Structure.
An orbital is not a classical path. It is a wavefunction, and its physical probability density is
For the ideal Coulomb Hamiltonian, is stationary up to the phase factor . The density is then time-independent.
s, p, d, f Notation
Section titled “s, p, d, f Notation”The spectroscopic letter names the orbital angular-momentum quantum number .
| Letter | Number of values | Angular character | |
|---|---|---|---|
| spherical angular factor | |||
| one angular nodal surface in the usual real separated basis | |||
| two angular nodal surfaces in the usual real separated basis | |||
| three angular nodal surfaces in the usual real separated basis |
The shell label combines with the letter. Thus means , means , and means . The label still matters: for example, the subshell contains three independent spatial orbitals.
The number of spatial orbitals in a fixed subshell is
If electron spin is included without spin-dependent interactions, each spatial orbital can be paired with two spin states. The word orbital by itself usually refers only to the spatial wavefunction unless a spin-orbital convention has been explicitly introduced.
Probability Densities and Radial Probability
Section titled “Probability Densities and Radial Probability”The three-dimensional probability for the electron to lie in a region is
In spherical coordinates,
After integrating over angles, the radial probability density is
This radial density is not the same object as the full three-dimensional density. For example, the hydrogen wavefunction is largest at , but the radial probability density is maximal at one Bohr radius because of the volume factor.
Orbital pictures usually show an isosurface of or a surface enclosing a chosen probability such as 90 or 95 percent. The boundary in such a picture is a visualization convention, not a hard edge of the atom.
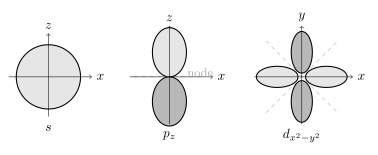
Schematic real-orbital shapes and nodal surfaces. The shaded lobes represent relative sign or phase of the wavefunction, not separate charged objects; probabilities come from .
Real and Complex Orbitals
Section titled “Real and Complex Orbitals”The separated functions are complex angular-momentum eigenfunctions. They are simultaneous eigenfunctions of and :
Complex orbitals are therefore natural when the component of angular momentum is being measured or when axial symmetry selects a preferred axis.
Real orbital pictures are different bases in the same degenerate subspace. For the orbitals, one common convention is
Overall signs and phases depend on the spherical-harmonic convention, but the three-dimensional subspace is the same. The real orbitals are convenient for visualizing nodal planes and directional bonding. They are not all eigenstates of .
This distinction is harmless in the ideal hydrogenic problem because all states with fixed and are degenerate. A linear combination inside the degenerate subspace is still a stationary state. If an external magnetic field selects a axis, however, the complex eigenstates often become the better basis.
Nodal Surfaces
Section titled “Nodal Surfaces”A node is a place where the wavefunction vanishes. Hydrogenic orbitals have radial nodes from and angular nodes from or from real linear combinations of spherical harmonics.
For hydrogenic bound states,
For the conventional real separated angular basis (polar factors times or ), one counts polar nodal cones and azimuthal nodal planes. Their sum is , giving the familiar textbook count of radial-plus-angular nodes in that basis.
This is not a basis-independent count of nodal surfaces. For example, vanishes in the equatorial plane, whereas has zeros on the polar axis and no nodal plane. The complex phase never vanishes. General linear combinations can change the geometry further. Always specify the actual orbital before counting its zero sets.
| Real orbital | Radial spheres | Angular zero surfaces |
|---|---|---|
| none | ||
| none | ||
| equatorial plane | ||
| equatorial plane | ||
| planes and |
Visualization Cautions
Section titled “Visualization Cautions”Orbital diagrams are useful, but they are compressed representations of a wavefunction. Several conventions are usually hidden:
- The plotted surface is an isosurface or probability-containing surface chosen by the author.
- Color or shading usually indicates the sign or phase of , not electric charge.
- The density is , so a sign change across a node is not itself visible in the probability density.
- Real-orbital shapes depend on a chosen basis in a degenerate subspace.
- Rotating the coordinate axes rotates the orbital labels.
- Complex orbitals carry phase variation that cannot be fully shown with simple positive and negative lobes.
- Hydrogenic orbitals are exact one-electron stationary states only in the ideal central Coulomb model.
The phrase electron cloud is acceptable as a qualitative image for probability density, but it should not be read as a classical smear of charge following a hidden orbit. In nonrelativistic wave mechanics, the orbital is the state assignment, and measured positions are sampled according to .
Relation to Many-Electron Orbitals
Section titled “Relation to Many-Electron Orbitals”In many-electron atoms, an orbital usually means a one-electron function appearing in an approximation such as Hartree, Hartree-Fock, or density-functional theory. Those orbitals are not exact one-electron eigenfunctions of a two-body Coulomb problem, because the electrons interact with one another.
The hydrogenic notation remains influential because it supplies a basis and a vocabulary: label angular behavior, radial nodes organize shells, and real combinations help visualize directional structure. But the interpretation changes. In a many-electron system, the many-body state and its antisymmetry carry the physical content, while orbitals are basis functions, variational objects, or effective one-particle quantities depending on the method.
Common Mistakes
Section titled “Common Mistakes”- Treating orbitals as classical trajectories.
- Thinking an orbital picture shows a hard surface where the atom ends.
- Confusing with the radial probability density .
- Assuming real orbitals are eigenstates.
- Forgetting that different real orbital pictures can be different bases for the same degenerate subspace.
- Interpreting positive and negative lobes as positive and negative charge.
- Applying exact hydrogenic formulas unchanged to many-electron atoms.
Exercises
Section titled “Exercises”- Count the number of spatial orbitals in a subshell.
Solution
A subshell has . The number of magnetic quantum numbers is
So there are five independent spatial orbitals.
- Determine the node counts for the real hydrogenic orbital.
Solution
For , and . The radial node count is
The angular node count is
Thus has one radial nodal sphere and the angular nodal plane . The same surface count need not hold for a complex orbital.
- Explain why is not an eigenstate of .
Solution
In a common convention,
The two terms have different eigenvalues:
Since is a superposition of two different values, applying does not return a single constant times . Thus is not an eigenstate, even though it is a valid stationary state in the ideal degenerate hydrogenic subspace.
- Why can two drawings of a orbital look rotated but describe the same physics in a central potential?
Solution
For a central potential, no direction in space is preferred. The three states span one angular-momentum subspace. Choosing , , and is choosing a real basis tied to chosen coordinate axes. Rotating the axes rotates the basis, but it does not change the subspace or the energy in the ideal central potential.
References
Section titled “References”- H. A. Bethe and E. E. Salpeter, Quantum Mechanics of One- and Two-Electron Atoms, Springer, 1957.
- C. Cohen-Tannoudji, B. Diu, and F. Laloë, Quantum Mechanics, Wiley, 1977.
- D. J. Griffiths and D. F. Schroeter, Introduction to Quantum Mechanics, 3rd ed., Cambridge University Press, 2018.
- J. J. Sakurai and J. Napolitano, Modern Quantum Mechanics, 3rd ed., Cambridge University Press, 2020.
- R. Shankar, Principles of Quantum Mechanics, 2nd ed., Springer, 1994.